Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.546326 |
| Energy at 298.15K | -133.551814 |
| HF Energy | -133.082581 |
| Nuclear repulsion energy | 70.891546 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3528 |
3297 |
2.65 |
|
|
|
| 2 |
A' |
3260 |
3047 |
8.17 |
|
|
|
| 3 |
A' |
3132 |
2926 |
13.07 |
|
|
|
| 4 |
A' |
3119 |
2914 |
53.26 |
|
|
|
| 5 |
A' |
1733 |
1619 |
38.11 |
|
|
|
| 6 |
A' |
1536 |
1435 |
8.76 |
|
|
|
| 7 |
A' |
1471 |
1374 |
14.73 |
|
|
|
| 8 |
A' |
1435 |
1341 |
9.65 |
|
|
|
| 9 |
A' |
1306 |
1220 |
36.16 |
|
|
|
| 10 |
A' |
1090 |
1019 |
29.06 |
|
|
|
| 11 |
A' |
959 |
896 |
4.97 |
|
|
|
| 12 |
A' |
495 |
462 |
16.89 |
|
|
|
| 13 |
A" |
3215 |
3004 |
13.81 |
|
|
|
| 14 |
A" |
1541 |
1440 |
7.60 |
|
|
|
| 15 |
A" |
1148 |
1073 |
5.93 |
|
|
|
| 16 |
A" |
1111 |
1038 |
14.69 |
|
|
|
| 17 |
A" |
698 |
652 |
54.13 |
|
|
|
| 18 |
A" |
188 |
176 |
1.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15482.0 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 14466.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.040 |
-0.617 |
0.000 |
| C2 |
0.000 |
0.454 |
0.000 |
| N3 |
1.248 |
0.160 |
0.000 |
| H4 |
-0.559 |
-1.590 |
0.000 |
| H5 |
-1.680 |
-0.528 |
0.877 |
| H6 |
-1.680 |
-0.528 |
-0.877 |
| H7 |
-0.373 |
1.483 |
0.000 |
| H8 |
1.799 |
1.020 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4936 | 2.4165 | 1.0855 | 1.0894 | 1.0894 | 2.2032 | 3.2776 |
C2 | 1.4936 | | 1.2819 | 2.1193 | 2.1347 | 2.1347 | 1.0937 | 1.8860 | N3 | 2.4165 | 1.2819 | | 2.5150 | 3.1327 | 3.1327 | 2.0920 | 1.0213 | H4 | 1.0855 | 2.1193 | 2.5150 | | 1.7761 | 1.7761 | 3.0781 | 3.5170 | H5 | 1.0894 | 2.1347 | 3.1327 | 1.7761 | | 1.7549 | 2.5533 | 3.9076 | H6 | 1.0894 | 2.1347 | 3.1327 | 1.7761 | 1.7549 | | 2.5533 | 3.9076 | H7 | 2.2032 | 1.0937 | 2.0920 | 3.0781 | 2.5533 | 2.5533 | | 2.2214 | H8 | 3.2776 | 1.8860 | 1.0213 | 3.5170 | 3.9076 | 3.9076 | 2.2214 | |
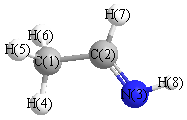 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.875 |
|
C1 |
C2 |
H7 |
115.895 |
| C2 |
C1 |
H4 |
109.502 |
|
C2 |
C1 |
H5 |
110.497 |
| C2 |
C1 |
H6 |
110.497 |
|
C2 |
N3 |
H8 |
109.418 |
| N3 |
C2 |
H7 |
123.229 |
|
H4 |
C1 |
H5 |
109.502 |
| H4 |
C1 |
H6 |
109.502 |
|
H5 |
C1 |
H6 |
107.308 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.545254 |
| Energy at 298.15K | -133.550705 |
| HF Energy | -133.081577 |
| Nuclear repulsion energy | 70.798665 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3493 |
3264 |
6.47 |
|
|
|
| 2 |
A' |
3235 |
3023 |
21.74 |
|
|
|
| 3 |
A' |
3189 |
2980 |
27.62 |
|
|
|
| 4 |
A' |
3125 |
2920 |
7.45 |
|
|
|
| 5 |
A' |
1725 |
1612 |
36.18 |
|
|
|
| 6 |
A' |
1542 |
1440 |
20.27 |
|
|
|
| 7 |
A' |
1465 |
1369 |
19.41 |
|
|
|
| 8 |
A' |
1447 |
1352 |
3.78 |
|
|
|
| 9 |
A' |
1306 |
1220 |
52.86 |
|
|
|
| 10 |
A' |
1095 |
1023 |
24.30 |
|
|
|
| 11 |
A' |
939 |
878 |
2.68 |
|
|
|
| 12 |
A' |
498 |
465 |
8.59 |
|
|
|
| 13 |
A" |
3218 |
3007 |
10.22 |
|
|
|
| 14 |
A" |
1534 |
1433 |
7.04 |
|
|
|
| 15 |
A" |
1179 |
1102 |
42.78 |
|
|
|
| 16 |
A" |
1103 |
1031 |
11.13 |
|
|
|
| 17 |
A" |
696 |
650 |
11.27 |
|
|
|
| 18 |
A" |
164 |
153 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15475.8 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 14460.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.028 |
-0.610 |
0.000 |
| C2 |
0.000 |
0.482 |
0.000 |
| N3 |
1.276 |
0.364 |
0.000 |
| H4 |
-0.557 |
-1.591 |
0.000 |
| H5 |
-1.669 |
-0.527 |
0.877 |
| H6 |
-1.669 |
-0.527 |
-0.877 |
| H7 |
-0.390 |
1.498 |
0.000 |
| H8 |
1.521 |
-0.630 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4994 | 2.5011 | 1.0880 | 1.0892 | 1.0892 | 2.2026 | 2.5493 |
C2 | 1.4994 | | 1.2812 | 2.1464 | 2.1380 | 2.1380 | 1.0890 | 1.8842 | N3 | 2.5011 | 1.2812 | | 2.6801 | 3.1988 | 3.1988 | 2.0156 | 1.0237 | H4 | 1.0880 | 2.1464 | 2.6801 | | 1.7707 | 1.7707 | 3.0939 | 2.2902 | H5 | 1.0892 | 2.1380 | 3.1988 | 1.7707 | | 1.7540 | 2.5506 | 3.3098 | H6 | 1.0892 | 2.1380 | 3.1988 | 1.7707 | 1.7540 | | 2.5506 | 3.3098 | H7 | 2.2026 | 1.0890 | 2.0156 | 3.0939 | 2.5506 | 2.5506 | | 2.8607 | H8 | 2.5493 | 1.8842 | 1.0237 | 2.2902 | 3.3098 | 3.3098 | 2.8607 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.003 |
|
C1 |
C2 |
H7 |
115.721 |
| C2 |
C1 |
H4 |
111.107 |
|
C2 |
C1 |
H5 |
110.356 |
| C2 |
C1 |
H6 |
110.356 |
|
C2 |
N3 |
H8 |
109.148 |
| N3 |
C2 |
H7 |
116.276 |
|
H4 |
C1 |
H5 |
108.836 |
| H4 |
C1 |
H6 |
108.836 |
|
H5 |
C1 |
H6 |
107.250 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
