Jump to
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.561893 |
| Energy at 298.15K | -133.567378 |
| HF Energy | -133.083006 |
| Nuclear repulsion energy | 70.846939 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3530 |
3309 |
4.92 |
|
|
|
| 2 |
A' |
3237 |
3035 |
12.39 |
|
|
|
| 3 |
A' |
3122 |
2927 |
35.11 |
|
|
|
| 4 |
A' |
3112 |
2918 |
31.31 |
|
|
|
| 5 |
A' |
1781 |
1670 |
39.50 |
|
|
|
| 6 |
A' |
1540 |
1444 |
6.82 |
|
|
|
| 7 |
A' |
1490 |
1397 |
18.67 |
|
|
|
| 8 |
A' |
1449 |
1359 |
6.33 |
|
|
|
| 9 |
A' |
1317 |
1235 |
33.60 |
|
|
|
| 10 |
A' |
1096 |
1028 |
26.29 |
|
|
|
| 11 |
A' |
958 |
898 |
5.00 |
|
|
|
| 12 |
A' |
496 |
465 |
16.91 |
|
|
|
| 13 |
A" |
3189 |
2990 |
19.58 |
|
|
|
| 14 |
A" |
1542 |
1445 |
6.60 |
|
|
|
| 15 |
A" |
1157 |
1085 |
6.49 |
|
|
|
| 16 |
A" |
1115 |
1046 |
14.91 |
|
|
|
| 17 |
A" |
697 |
653 |
52.35 |
|
|
|
| 18 |
A" |
182 |
171 |
1.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15504.3 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 14536.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.038 |
-0.629 |
0.000 |
| C2 |
0.000 |
0.454 |
0.000 |
| N3 |
1.246 |
0.175 |
0.000 |
| H4 |
-0.550 |
-1.601 |
0.000 |
| H5 |
-1.678 |
-0.545 |
0.879 |
| H6 |
-1.678 |
-0.545 |
-0.879 |
| H7 |
-0.381 |
1.480 |
0.000 |
| H8 |
1.792 |
1.038 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5000 | 2.4210 | 1.0872 | 1.0913 | 1.0913 | 2.2089 | 3.2843 |
C2 | 1.5000 | | 1.2767 | 2.1278 | 2.1418 | 2.1418 | 1.0943 | 1.8848 | N3 | 2.4210 | 1.2767 | | 2.5265 | 3.1372 | 3.1372 | 2.0854 | 1.0210 | H4 | 1.0872 | 2.1278 | 2.5265 | | 1.7780 | 1.7780 | 3.0858 | 3.5289 | H5 | 1.0913 | 2.1418 | 3.1372 | 1.7780 | | 1.7589 | 2.5603 | 3.9143 | H6 | 1.0913 | 2.1418 | 3.1372 | 1.7780 | 1.7589 | | 2.5603 | 3.9143 | H7 | 2.2089 | 1.0943 | 2.0854 | 3.0858 | 2.5603 | 2.5603 | | 2.2177 | H8 | 3.2843 | 1.8848 | 1.0210 | 3.5289 | 3.9143 | 3.9143 | 2.2177 | |
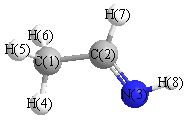 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.150 |
|
C1 |
C2 |
H7 |
115.856 |
| C2 |
C1 |
H4 |
109.622 |
|
C2 |
C1 |
H5 |
110.495 |
| C2 |
C1 |
H6 |
110.495 |
|
C2 |
N3 |
H8 |
109.733 |
| N3 |
C2 |
H7 |
122.994 |
|
H4 |
C1 |
H5 |
109.403 |
| H4 |
C1 |
H6 |
109.403 |
|
H5 |
C1 |
H6 |
107.384 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -133.560916 |
| Energy at 298.15K | -133.566376 |
| HF Energy | -133.081965 |
| Nuclear repulsion energy | 70.783441 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3492 |
3274 |
8.85 |
|
|
|
| 2 |
A' |
3211 |
3011 |
30.96 |
|
|
|
| 3 |
A' |
3178 |
2979 |
23.65 |
|
|
|
| 4 |
A' |
3112 |
2918 |
10.01 |
|
|
|
| 5 |
A' |
1778 |
1667 |
39.47 |
|
|
|
| 6 |
A' |
1546 |
1449 |
17.22 |
|
|
|
| 7 |
A' |
1476 |
1384 |
19.62 |
|
|
|
| 8 |
A' |
1456 |
1365 |
4.49 |
|
|
|
| 9 |
A' |
1321 |
1239 |
51.79 |
|
|
|
| 10 |
A' |
1100 |
1031 |
21.17 |
|
|
|
| 11 |
A' |
942 |
884 |
1.97 |
|
|
|
| 12 |
A' |
502 |
471 |
8.81 |
|
|
|
| 13 |
A" |
3192 |
2993 |
15.17 |
|
|
|
| 14 |
A" |
1535 |
1439 |
6.09 |
|
|
|
| 15 |
A" |
1181 |
1108 |
44.59 |
|
|
|
| 16 |
A" |
1110 |
1041 |
9.12 |
|
|
|
| 17 |
A" |
701 |
657 |
11.96 |
|
|
|
| 18 |
A" |
162 |
152 |
0.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15498.5 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 14531.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.024 |
-0.618 |
0.000 |
| C2 |
0.000 |
0.485 |
0.000 |
| N3 |
1.271 |
0.371 |
0.000 |
| H4 |
-0.547 |
-1.598 |
0.000 |
| H5 |
-1.666 |
-0.540 |
0.879 |
| H6 |
-1.666 |
-0.540 |
-0.879 |
| H7 |
-0.399 |
1.500 |
0.000 |
| H8 |
1.528 |
-0.620 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5054 | 2.4995 | 1.0898 | 1.0911 | 1.0911 | 2.2082 | 2.5520 |
C2 | 1.5054 | | 1.2761 | 2.1532 | 2.1449 | 2.1449 | 1.0900 | 1.8853 | N3 | 2.4995 | 1.2761 | | 2.6796 | 3.1986 | 3.1986 | 2.0151 | 1.0237 | H4 | 1.0898 | 2.1532 | 2.6796 | | 1.7730 | 1.7730 | 3.1007 | 2.2933 | H5 | 1.0911 | 2.1449 | 3.1986 | 1.7730 | | 1.7579 | 2.5576 | 3.3136 | H6 | 1.0911 | 2.1449 | 3.1986 | 1.7730 | 1.7579 | | 2.5576 | 3.3136 | H7 | 2.2082 | 1.0900 | 2.0151 | 3.1007 | 2.5576 | 2.5576 | | 2.8639 | H8 | 2.5520 | 1.8853 | 1.0237 | 2.2933 | 3.3136 | 3.3136 | 2.8639 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.763 |
|
C1 |
C2 |
H7 |
115.668 |
| C2 |
C1 |
H4 |
111.121 |
|
C2 |
C1 |
H5 |
110.374 |
| C2 |
C1 |
H6 |
110.374 |
|
C2 |
N3 |
H8 |
109.636 |
| N3 |
C2 |
H7 |
116.569 |
|
H4 |
C1 |
H5 |
108.772 |
| H4 |
C1 |
H6 |
108.772 |
|
H5 |
C1 |
H6 |
107.327 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
