Jump to
S2C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -498.540777 |
| Energy at 298.15K | -498.540645 |
| HF Energy | -498.540777 |
| Nuclear repulsion energy | 38.505295 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.046 |
1.200 |
0.000 |
| Cl2 |
0.046 |
-0.507 |
0.000 |
| H3 |
-1.048 |
1.428 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.7072 | 1.1173 |
Cl2 | 1.7072 | | 2.2233 | H3 | 1.1173 | 2.2233 | |
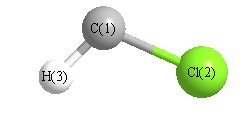 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
101.803 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.250 |
|
|
|
| 2 |
Cl |
0.068 |
|
|
|
| 3 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.528 |
-0.527 |
0.000 |
1.616 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.354 |
-2.570 |
0.000 |
| y |
-2.570 |
-19.987 |
0.000 |
| z |
0.000 |
0.000 |
-17.831 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.446 |
-2.570 |
0.000 |
| y |
-2.570 |
-1.394 |
0.000 |
| z |
0.000 |
0.000 |
1.840 |
|
| Polar |
|---|
| 3z2-r2 | 3.679 |
|---|
| x2-y2 | 0.632 |
|---|
| xy | -2.570 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.335 |
-0.385 |
0.000 |
| y |
-0.385 |
4.512 |
0.000 |
| z |
0.000 |
0.000 |
1.831 |
<r2> (average value of r
2) Å
2
| <r2> |
28.103 |
| (<r2>)1/2 |
5.301 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -498.545038 |
| Energy at 298.15K | -498.544897 |
| HF Energy | -498.545038 |
| Nuclear repulsion energy | 38.963875 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.037 |
1.154 |
0.000 |
| Cl2 |
0.037 |
-0.513 |
0.000 |
| H3 |
-0.841 |
1.793 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.6663 | 1.0854 |
Cl2 | 1.6663 | | 2.4667 | H3 | 1.0854 | 2.4667 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
126.071 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.284 |
|
|
|
| 2 |
Cl |
0.064 |
|
|
|
| 3 |
H |
0.220 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.796 |
0.478 |
0.000 |
0.929 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.225 |
-1.493 |
0.000 |
| y |
-1.493 |
-16.885 |
0.000 |
| z |
0.000 |
0.000 |
-19.351 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.107 |
-1.493 |
0.000 |
| y |
-1.493 |
1.903 |
0.000 |
| z |
0.000 |
0.000 |
-1.796 |
|
| Polar |
|---|
| 3z2-r2 | -3.592 |
|---|
| x2-y2 | -1.340 |
|---|
| xy | -1.493 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.876 |
-0.231 |
0.000 |
| y |
-0.231 |
4.187 |
0.000 |
| z |
0.000 |
0.000 |
1.479 |
<r2> (average value of r
2) Å
2
| <r2> |
27.744 |
| (<r2>)1/2 |
5.267 |
