Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -193.226555 |
| Energy at 298.15K | |
| HF Energy | -193.226555 |
| Nuclear repulsion energy | 148.932062 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3293 |
3138 |
0.13 |
|
|
|
| 2 |
A1 |
3285 |
3131 |
9.57 |
|
|
|
| 3 |
A1 |
3252 |
3099 |
1.42 |
|
|
|
| 4 |
A1 |
1615 |
1539 |
0.08 |
|
|
|
| 5 |
A1 |
1492 |
1422 |
1.76 |
|
|
|
| 6 |
A1 |
1161 |
1107 |
0.00 |
|
|
|
| 7 |
A1 |
1101 |
1049 |
9.22 |
|
|
|
| 8 |
A1 |
1061 |
1011 |
9.86 |
|
|
|
| 9 |
A1 |
843 |
804 |
0.30 |
|
|
|
| 10 |
A2 |
909 |
866 |
0.00 |
|
|
|
| 11 |
A2 |
823 |
784 |
0.00 |
|
|
|
| 12 |
A2 |
529 |
504 |
0.00 |
|
|
|
| 13 |
B1 |
882 |
840 |
6.73 |
|
|
|
| 14 |
B1 |
723 |
689 |
0.84 |
|
|
|
| 15 |
B1 |
688 |
656 |
89.60 |
|
|
|
| 16 |
B1 |
490 |
467 |
11.52 |
|
|
|
| 17 |
B2 |
3268 |
3115 |
16.27 |
|
|
|
| 18 |
B2 |
3253 |
3100 |
8.75 |
|
|
|
| 19 |
B2 |
1423 |
1356 |
42.74 |
|
|
|
| 20 |
B2 |
1308 |
1246 |
0.00 |
|
|
|
| 21 |
B2 |
1231 |
1173 |
1.72 |
|
|
|
| 22 |
B2 |
987 |
940 |
0.10 |
|
|
|
| 23 |
B2 |
934 |
890 |
2.64 |
|
|
|
| 24 |
B2 |
36i |
34i |
3.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17255.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 16446.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.223 |
| C2 |
0.000 |
1.121 |
0.392 |
| C3 |
0.000 |
0.678 |
-1.004 |
| C4 |
0.000 |
-0.678 |
-1.004 |
| C5 |
0.000 |
-1.121 |
0.392 |
| H6 |
0.000 |
0.000 |
2.305 |
| H7 |
0.000 |
2.157 |
0.714 |
| H8 |
0.000 |
1.335 |
-1.865 |
| H9 |
0.000 |
-1.335 |
-1.865 |
| H10 |
0.000 |
-2.157 |
0.714 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3960 | 2.3281 | 2.3281 | 1.3960 | 1.0820 | 2.2167 | 3.3646 | 3.3646 | 2.2167 |
C2 | 1.3960 | | 1.4642 | 2.2774 | 2.2427 | 2.2178 | 1.0850 | 2.2670 | 3.3358 | 3.2945 | C3 | 2.3281 | 1.4642 | | 1.3567 | 2.2774 | 3.3779 | 2.2668 | 1.0831 | 2.1899 | 3.3154 | C4 | 2.3281 | 2.2774 | 1.3567 | | 1.4642 | 3.3779 | 3.3154 | 2.1899 | 1.0831 | 2.2668 | C5 | 1.3960 | 2.2427 | 2.2774 | 1.4642 | | 2.2178 | 3.2945 | 3.3358 | 2.2670 | 1.0850 | H6 | 1.0820 | 2.2178 | 3.3779 | 3.3779 | 2.2178 | | 2.6808 | 4.3788 | 4.3788 | 2.6808 | H7 | 2.2167 | 1.0850 | 2.2668 | 3.3154 | 3.2945 | 2.6808 | | 2.7069 | 4.3415 | 4.3149 | H8 | 3.3646 | 2.2670 | 1.0831 | 2.1899 | 3.3358 | 4.3788 | 2.7069 | | 2.6702 | 4.3415 | H9 | 3.3646 | 3.3358 | 2.1899 | 1.0831 | 2.2670 | 4.3788 | 4.3415 | 2.6702 | | 2.7069 | H10 | 2.2167 | 3.2945 | 3.3154 | 2.2668 | 1.0850 | 2.6808 | 4.3149 | 4.3415 | 2.7069 | |
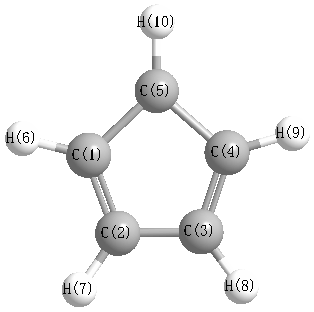 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.946 |
|
C1 |
C2 |
H7 |
126.174 |
| C1 |
C5 |
C4 |
108.946 |
|
C1 |
C5 |
H10 |
126.174 |
| C2 |
C1 |
C5 |
106.886 |
|
C2 |
C1 |
H6 |
126.557 |
| C2 |
C3 |
C4 |
107.611 |
|
C2 |
C3 |
H8 |
125.061 |
| C3 |
C2 |
H7 |
124.879 |
|
C3 |
C4 |
C5 |
107.611 |
| C3 |
C4 |
H9 |
127.329 |
|
C4 |
C3 |
H8 |
127.329 |
| C4 |
C5 |
H10 |
124.879 |
|
C5 |
C1 |
H6 |
126.557 |
| C5 |
C4 |
H9 |
125.061 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.197 |
|
|
|
| 2 |
C |
-0.152 |
|
|
|
| 3 |
C |
-0.181 |
|
|
|
| 4 |
C |
-0.181 |
|
|
|
| 5 |
C |
-0.152 |
|
|
|
| 6 |
H |
0.168 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.031 |
0.031 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.491 |
0.000 |
0.000 |
| y |
0.000 |
-25.022 |
0.000 |
| z |
0.000 |
0.000 |
-26.906 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.527 |
0.000 |
0.000 |
| y |
0.000 |
4.676 |
0.000 |
| z |
0.000 |
0.000 |
1.851 |
|
| Polar |
|---|
| 3z2-r2 | 3.701 |
|---|
| x2-y2 | -7.469 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.727 |
0.000 |
0.000 |
| y |
0.000 |
8.665 |
0.000 |
| z |
0.000 |
0.000 |
7.797 |
<r2> (average value of r
2) Å
2
| <r2> |
87.263 |
| (<r2>)1/2 |
9.341 |
