Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -193.250543 |
| Energy at 298.15K | -193.254505 |
| HF Energy | -193.250543 |
| Nuclear repulsion energy | 148.966490 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3288 |
3138 |
0.37 |
|
|
|
| 2 |
A1 |
3280 |
3130 |
8.34 |
|
|
|
| 3 |
A1 |
3244 |
3096 |
1.56 |
|
|
|
| 4 |
A1 |
1611 |
1538 |
0.06 |
|
|
|
| 5 |
A1 |
1487 |
1419 |
1.84 |
|
|
|
| 6 |
A1 |
1157 |
1104 |
0.00 |
|
|
|
| 7 |
A1 |
1097 |
1047 |
8.75 |
|
|
|
| 8 |
A1 |
1057 |
1009 |
9.97 |
|
|
|
| 9 |
A1 |
843 |
804 |
0.32 |
|
|
|
| 10 |
A2 |
912 |
871 |
0.00 |
|
|
|
| 11 |
A2 |
823 |
786 |
0.00 |
|
|
|
| 12 |
A2 |
528 |
504 |
0.00 |
|
|
|
| 13 |
B1 |
884 |
844 |
6.54 |
|
|
|
| 14 |
B1 |
725 |
692 |
0.47 |
|
|
|
| 15 |
B1 |
689 |
657 |
84.81 |
|
|
|
| 16 |
B1 |
490 |
468 |
10.99 |
|
|
|
| 17 |
B2 |
3262 |
3112 |
13.55 |
|
|
|
| 18 |
B2 |
3245 |
3097 |
9.53 |
|
|
|
| 19 |
B2 |
1417 |
1352 |
42.06 |
|
|
|
| 20 |
B2 |
1303 |
1243 |
0.00 |
|
|
|
| 21 |
B2 |
1228 |
1172 |
1.60 |
|
|
|
| 22 |
B2 |
983 |
938 |
0.10 |
|
|
|
| 23 |
B2 |
931 |
889 |
2.65 |
|
|
|
| 24 |
B2 |
29 |
28 |
2.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17257.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 16468.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.223 |
| C2 |
0.000 |
1.121 |
0.392 |
| C3 |
0.000 |
0.678 |
-1.004 |
| C4 |
0.000 |
-0.678 |
-1.004 |
| C5 |
0.000 |
-1.121 |
0.392 |
| H6 |
0.000 |
0.000 |
2.304 |
| H7 |
0.000 |
2.157 |
0.714 |
| H8 |
0.000 |
1.334 |
-1.865 |
| H9 |
0.000 |
-1.334 |
-1.865 |
| H10 |
0.000 |
-2.157 |
0.714 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3958 | 2.3276 | 2.3276 | 1.3958 | 1.0811 | 2.2159 | 3.3635 | 3.3635 | 2.2159 |
C2 | 1.3958 | | 1.4641 | 2.2771 | 2.2426 | 2.2168 | 1.0843 | 2.2664 | 3.3346 | 3.2937 | C3 | 2.3276 | 1.4641 | | 1.3564 | 2.2771 | 3.3765 | 2.2663 | 1.0823 | 2.1887 | 3.3146 | C4 | 2.3276 | 2.2771 | 1.3564 | | 1.4641 | 3.3765 | 3.3146 | 2.1887 | 1.0823 | 2.2663 | C5 | 1.3958 | 2.2426 | 2.2771 | 1.4641 | | 2.2168 | 3.2937 | 3.3346 | 2.2664 | 1.0843 | H6 | 1.0811 | 2.2168 | 3.3765 | 3.3765 | 2.2168 | | 2.6794 | 4.3769 | 4.3769 | 2.6794 | H7 | 2.2159 | 1.0843 | 2.2663 | 3.3146 | 3.2937 | 2.6794 | | 2.7067 | 4.3398 | 4.3132 | H8 | 3.3635 | 2.2664 | 1.0823 | 2.1887 | 3.3346 | 4.3769 | 2.7067 | | 2.6680 | 4.3398 | H9 | 3.3635 | 3.3346 | 2.1887 | 1.0823 | 2.2664 | 4.3769 | 4.3398 | 2.6680 | | 2.7067 | H10 | 2.2159 | 3.2937 | 3.3146 | 2.2663 | 1.0843 | 2.6794 | 4.3132 | 4.3398 | 2.7067 | |
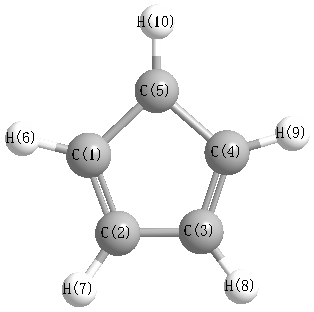 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.935 |
|
C1 |
C2 |
H7 |
126.161 |
| C1 |
C5 |
C4 |
108.935 |
|
C1 |
C5 |
H10 |
126.161 |
| C2 |
C1 |
C5 |
106.899 |
|
C2 |
C1 |
H6 |
126.550 |
| C2 |
C3 |
C4 |
107.616 |
|
C2 |
C3 |
H8 |
125.090 |
| C3 |
C2 |
H7 |
124.905 |
|
C3 |
C4 |
C5 |
107.616 |
| C3 |
C4 |
H9 |
127.295 |
|
C4 |
C3 |
H8 |
127.295 |
| C4 |
C5 |
H10 |
124.905 |
|
C5 |
C1 |
H6 |
126.550 |
| C5 |
C4 |
H9 |
125.090 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.149 |
|
|
|
| 2 |
C |
-0.105 |
|
|
|
| 3 |
C |
-0.133 |
|
|
|
| 4 |
C |
-0.133 |
|
|
|
| 5 |
C |
-0.105 |
|
|
|
| 6 |
H |
0.120 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.123 |
|
|
|
| 9 |
H |
0.123 |
|
|
|
| 10 |
H |
0.130 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.030 |
0.030 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.397 |
0.000 |
0.000 |
| y |
0.000 |
-25.108 |
0.000 |
| z |
0.000 |
0.000 |
-26.979 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.354 |
0.000 |
0.000 |
| y |
0.000 |
4.581 |
0.000 |
| z |
0.000 |
0.000 |
1.773 |
|
| Polar |
|---|
| 3z2-r2 | 3.546 |
|---|
| x2-y2 | -7.290 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.779 |
0.000 |
0.000 |
| y |
0.000 |
8.745 |
0.000 |
| z |
0.000 |
0.000 |
7.877 |
<r2> (average value of r
2) Å
2
| <r2> |
87.243 |
| (<r2>)1/2 |
9.340 |
