Jump to
S2C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -77.010328 |
| Energy at 298.15K | -77.009995 |
| HF Energy | -76.767077 |
| Nuclear repulsion energy | 23.789984 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3236 |
3023 |
44.60 |
|
|
|
| 2 |
A1 |
1716 |
1603 |
78.76 |
|
|
|
| 3 |
A1 |
1268 |
1185 |
17.00 |
|
|
|
| 4 |
B1 |
760 |
711 |
93.49 |
|
|
|
| 5 |
B2 |
3336 |
3117 |
18.77 |
|
|
|
| 6 |
B2 |
317 |
296 |
4.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5316.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 4967.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.482 |
| C2 |
0.000 |
0.000 |
0.825 |
| H3 |
0.000 |
0.935 |
-1.027 |
| H4 |
0.000 |
-0.935 |
-1.027 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3067 | 1.0823 | 1.0823 |
C2 | 1.3067 | | 2.0745 | 2.0745 | H3 | 1.0823 | 2.0745 | | 1.8701 | H4 | 1.0823 | 2.0745 | 1.8701 | |
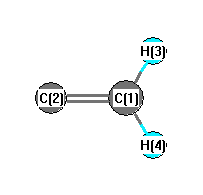 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.238 |
|
C2 |
C1 |
H4 |
120.238 |
| H3 |
C1 |
H4 |
119.524 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -76.937308 |
| Energy at 298.15K | -76.937433 |
| HF Energy | -76.736791 |
| Nuclear repulsion energy | 23.785987 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3195 |
2985 |
2.93 |
|
|
|
| 2 |
A1 |
1647 |
1539 |
3.01 |
|
|
|
| 3 |
A1 |
1471 |
1374 |
4.52 |
|
|
|
| 4 |
B1 |
953 |
891 |
32.34 |
|
|
|
| 5 |
B2 |
3288 |
3073 |
5.82 |
|
|
|
| 6 |
B2 |
1047 |
979 |
2.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5800.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 5420.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.479 |
| C2 |
0.000 |
0.000 |
0.826 |
| H3 |
0.000 |
0.926 |
-1.042 |
| H4 |
0.000 |
-0.926 |
-1.042 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3053 | 1.0836 | 1.0836 |
C2 | 1.3053 | | 2.0853 | 2.0853 | H3 | 1.0836 | 2.0853 | | 1.8513 | H4 | 1.0836 | 2.0853 | 1.8513 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
