return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5 (phenyl)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-230.818202
Energy at 298.15K-230.823328
HF Energy-230.068458
Nuclear repulsion energy197.781038
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3303 3086 1.38      
2 A1 3297 3081 9.54      
3 A1 3267 3053 0.55      
4 A1 1845 1724 2.76      
5 A1 1600 1495 12.94      
6 A1 1252 1170 0.05      
7 A1 1143 1068 14.52      
8 A1 1091 1019 0.02      
9 A1 988 923 0.50      
10 A1 658 614 1.75      
11 A2 1124 1050 0.00      
12 A2 985 920 0.00      
13 A2 470 440 0.00      
14 B1 1108 1035 0.18      
15 B1 1039 971 0.70      
16 B1 823 769 86.57      
17 B1 721 674 1.04      
18 B1 494 462 5.08      
19 B2 3302 3086 8.12      
20 B2 3276 3061 9.79      
21 B2 1900 1775 8.09      
22 B2 1587 1483 7.55      
23 B2 1376 1286 0.26      
24 B2 1277 1193 0.07      
25 B2 1200 1122 2.52      
26 B2 1122 1049 5.85      
27 B2 635 593 0.64      

Unscaled Zero Point Vibrational Energy (zpe) 20441.4 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 19100.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.21744 0.19380 0.10247

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.373
C2 0.000 1.205 0.755
C3 0.000 -1.205 0.755
C4 0.000 1.188 -0.617
C5 0.000 -1.188 -0.617
C6 0.000 0.000 -1.299
H7 0.000 2.134 1.306
H8 0.000 -2.134 1.306
H9 0.000 2.123 -1.162
H10 0.000 -2.123 -1.162
H11 0.000 0.000 -2.380

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11
C11.35441.35442.31842.31842.67282.13492.13493.30693.30693.7531
C21.35442.40981.37242.75852.38151.08003.38392.12543.84043.3582
C31.35442.40982.75851.37242.38153.38391.08003.84042.12543.3582
C42.31841.37242.75852.37621.36992.14303.83841.08193.35562.1254
C52.31842.75851.37242.37621.36993.83842.14303.35561.08192.1254
C62.67282.38152.38151.36991.36993.36743.36742.12742.12741.0804
H72.13491.08003.38392.14303.83843.36744.26772.46774.92034.2585
H82.13493.38391.08003.83842.14303.36744.26774.92032.46774.2585
H93.30692.12543.84041.08193.35562.12742.46774.92034.24592.4474
H103.30693.84042.12543.35561.08192.12744.92032.46774.24592.4474
H113.75313.35823.35822.12542.12541.08044.25854.25852.44742.4474

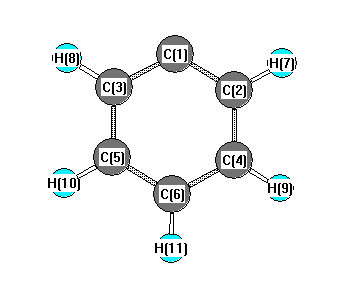
picture of phenyl state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 116.474 C1 C2 H7 122.165
C1 C3 C5 116.474 C1 C3 H8 122.165
C2 C1 C3 125.651 C2 C4 C6 120.556
C2 C4 H9 119.526 C3 C5 C6 120.556
C3 C5 H10 119.526 C4 C2 H7 121.361
C4 C6 C5 120.288 C4 C6 H11 119.856
C5 C3 H8 121.361 C5 C6 H11 119.856
C6 C4 H9 119.918 C6 C5 H10 119.918
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability