Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.551912 |
| Energy at 298.15K | -235.565335 |
| HF Energy | -235.551912 |
| Nuclear repulsion energy | 249.974601 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3140 |
2993 |
106.27 |
|
|
|
| 2 |
A' |
3135 |
2988 |
13.32 |
|
|
|
| 3 |
A' |
3131 |
2984 |
25.25 |
|
|
|
| 4 |
A' |
3093 |
2947 |
20.64 |
|
|
|
| 5 |
A' |
3079 |
2935 |
11.36 |
|
|
|
| 6 |
A' |
3075 |
2930 |
7.99 |
|
|
|
| 7 |
A' |
3058 |
2915 |
30.72 |
|
|
|
| 8 |
A' |
3057 |
2914 |
25.21 |
|
|
|
| 9 |
A' |
1529 |
1457 |
9.89 |
|
|
|
| 10 |
A' |
1523 |
1451 |
2.34 |
|
|
|
| 11 |
A' |
1518 |
1447 |
2.18 |
|
|
|
| 12 |
A' |
1437 |
1370 |
9.55 |
|
|
|
| 13 |
A' |
1435 |
1368 |
0.69 |
|
|
|
| 14 |
A' |
1399 |
1334 |
0.45 |
|
|
|
| 15 |
A' |
1383 |
1318 |
10.32 |
|
|
|
| 16 |
A' |
1298 |
1237 |
2.10 |
|
|
|
| 17 |
A' |
1270 |
1211 |
0.46 |
|
|
|
| 18 |
A' |
1217 |
1160 |
0.25 |
|
|
|
| 19 |
A' |
1159 |
1105 |
0.29 |
|
|
|
| 20 |
A' |
1069 |
1019 |
0.28 |
|
|
|
| 21 |
A' |
973 |
927 |
1.24 |
|
|
|
| 22 |
A' |
960 |
915 |
1.14 |
|
|
|
| 23 |
A' |
881 |
839 |
1.39 |
|
|
|
| 24 |
A' |
823 |
784 |
1.15 |
|
|
|
| 25 |
A' |
651 |
621 |
1.99 |
|
|
|
| 26 |
A' |
428 |
408 |
0.14 |
|
|
|
| 27 |
A' |
327 |
312 |
0.14 |
|
|
|
| 28 |
A' |
99 |
95 |
0.01 |
|
|
|
| 29 |
A" |
3140 |
2992 |
32.78 |
|
|
|
| 30 |
A" |
3137 |
2990 |
24.83 |
|
|
|
| 31 |
A" |
3131 |
2985 |
9.72 |
|
|
|
| 32 |
A" |
3074 |
2930 |
65.49 |
|
|
|
| 33 |
A" |
1525 |
1453 |
4.40 |
|
|
|
| 34 |
A" |
1523 |
1452 |
0.10 |
|
|
|
| 35 |
A" |
1495 |
1425 |
3.67 |
|
|
|
| 36 |
A" |
1312 |
1251 |
0.68 |
|
|
|
| 37 |
A" |
1287 |
1227 |
0.00 |
|
|
|
| 38 |
A" |
1249 |
1190 |
0.11 |
|
|
|
| 39 |
A" |
1159 |
1105 |
0.69 |
|
|
|
| 40 |
A" |
1148 |
1094 |
0.12 |
|
|
|
| 41 |
A" |
1008 |
961 |
4.00 |
|
|
|
| 42 |
A" |
932 |
888 |
0.01 |
|
|
|
| 43 |
A" |
923 |
880 |
0.00 |
|
|
|
| 44 |
A" |
817 |
778 |
0.38 |
|
|
|
| 45 |
A" |
374 |
357 |
0.01 |
|
|
|
| 46 |
A" |
261 |
248 |
0.17 |
|
|
|
| 47 |
A" |
255 |
243 |
0.00 |
|
|
|
| 48 |
A" |
236 |
225 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37066.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 35328.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.257 |
0.086 |
1.078 |
| C2 |
0.257 |
0.086 |
-1.078 |
| C3 |
-0.660 |
0.710 |
0.000 |
| C4 |
0.851 |
-0.854 |
0.000 |
| C5 |
-0.864 |
2.212 |
0.000 |
| C6 |
0.257 |
-2.254 |
0.000 |
| H7 |
0.995 |
0.809 |
1.446 |
| H8 |
-0.227 |
-0.387 |
1.941 |
| H9 |
0.995 |
0.809 |
-1.446 |
| H10 |
-0.227 |
-0.387 |
-1.941 |
| H11 |
-1.636 |
0.208 |
0.000 |
| H12 |
1.945 |
-0.922 |
0.000 |
| H13 |
0.101 |
2.735 |
0.000 |
| H14 |
-0.840 |
-2.221 |
0.000 |
| H15 |
-1.422 |
2.538 |
-0.886 |
| H16 |
-1.422 |
2.538 |
0.886 |
| H17 |
0.574 |
-2.815 |
-0.887 |
| H18 |
0.574 |
-2.815 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1566 | 1.5468 | 1.5491 | 2.6343 | 2.5766 | 1.0965 | 1.0964 | 2.7278 | 3.0941 | 2.1816 | 2.2424 | 2.8644 | 2.7726 | 3.5621 | 2.9775 | 3.5188 | 2.9250 |
C2 | 2.1566 | | 1.5468 | 1.5491 | 2.6343 | 2.5766 | 2.7278 | 3.0941 | 1.0965 | 1.0964 | 2.1816 | 2.2424 | 2.8644 | 2.7726 | 2.9775 | 3.5621 | 2.9250 | 3.5188 | C3 | 1.5468 | 1.5468 | | 2.1745 | 1.5165 | 3.1023 | 2.2004 | 2.2711 | 2.2004 | 2.2711 | 1.0970 | 3.0738 | 2.1638 | 2.9360 | 2.1698 | 2.1698 | 3.8386 | 3.8386 | C4 | 1.5491 | 1.5491 | 2.1745 | | 3.5133 | 1.5203 | 2.2087 | 2.2685 | 2.2087 | 2.2685 | 2.7037 | 1.0962 | 3.6670 | 2.1736 | 4.1781 | 4.1781 | 2.1701 | 2.1701 | C5 | 2.6343 | 2.6343 | 1.5165 | 3.5133 | | 4.6048 | 2.7418 | 3.3061 | 2.7418 | 3.3061 | 2.1481 | 4.2087 | 1.0971 | 4.4333 | 1.0968 | 1.0968 | 5.3039 | 5.3039 | C6 | 2.5766 | 2.5766 | 3.1023 | 1.5203 | 4.6048 | | 3.4665 | 2.7361 | 3.4665 | 2.7361 | 3.1053 | 2.1499 | 4.9917 | 1.0972 | 5.1541 | 5.1541 | 1.0966 | 1.0966 | H7 | 1.0965 | 2.7278 | 2.2004 | 2.2087 | 2.7418 | 3.4665 | | 1.7798 | 2.8927 | 3.7942 | 3.0617 | 2.4474 | 2.5697 | 3.8257 | 3.7779 | 3.0241 | 4.3309 | 3.6910 | H8 | 1.0964 | 3.0941 | 2.2711 | 2.2685 | 3.3061 | 2.7361 | 1.7798 | | 3.7942 | 3.8818 | 2.4710 | 2.9612 | 3.6910 | 2.7395 | 4.2399 | 3.3310 | 3.8125 | 2.7653 | H9 | 2.7278 | 1.0965 | 2.2004 | 2.2087 | 2.7418 | 3.4665 | 2.8927 | 3.7942 | | 1.7798 | 3.0617 | 2.4474 | 2.5697 | 3.8257 | 3.0241 | 3.7779 | 3.6910 | 4.3309 | H10 | 3.0941 | 1.0964 | 2.2711 | 2.2685 | 3.3061 | 2.7361 | 3.7942 | 3.8818 | 1.7798 | | 2.4710 | 2.9612 | 3.6910 | 2.7395 | 3.3310 | 4.2399 | 2.7653 | 3.8125 | H11 | 2.1816 | 2.1816 | 1.0970 | 2.7037 | 2.1481 | 3.1053 | 3.0617 | 2.4710 | 3.0617 | 2.4710 | | 3.7546 | 3.0665 | 2.5558 | 2.5020 | 2.5020 | 3.8481 | 3.8481 | H12 | 2.2424 | 2.2424 | 3.0738 | 1.0962 | 4.2087 | 2.1499 | 2.4474 | 2.9612 | 2.4474 | 2.9612 | 3.7546 | | 4.0960 | 3.0724 | 4.9083 | 4.9083 | 2.5002 | 2.5002 | H13 | 2.8644 | 2.8644 | 2.1638 | 3.6670 | 1.0971 | 4.9917 | 2.5697 | 3.6910 | 2.5697 | 3.6910 | 3.0665 | 4.0960 | | 5.0445 | 1.7728 | 1.7728 | 5.6409 | 5.6409 | H14 | 2.7726 | 2.7726 | 2.9360 | 2.1736 | 4.4333 | 1.0972 | 3.8257 | 2.7395 | 3.8257 | 2.7395 | 2.5558 | 3.0724 | 5.0445 | | 4.8753 | 4.8753 | 1.7715 | 1.7715 | H15 | 3.5621 | 2.9775 | 2.1698 | 4.1781 | 1.0968 | 5.1541 | 3.7779 | 4.2399 | 3.0241 | 3.3310 | 2.5020 | 4.9083 | 1.7728 | 4.8753 | | 1.7727 | 5.7130 | 5.9819 | H16 | 2.9775 | 3.5621 | 2.1698 | 4.1781 | 1.0968 | 5.1541 | 3.0241 | 3.3310 | 3.7779 | 4.2399 | 2.5020 | 4.9083 | 1.7728 | 4.8753 | 1.7727 | | 5.9819 | 5.7130 | H17 | 3.5188 | 2.9250 | 3.8386 | 2.1701 | 5.3039 | 1.0966 | 4.3309 | 3.8125 | 3.6910 | 2.7653 | 3.8481 | 2.5002 | 5.6409 | 1.7715 | 5.7130 | 5.9819 | | 1.7744 | H18 | 2.9250 | 3.5188 | 3.8386 | 2.1701 | 5.3039 | 1.0966 | 3.6910 | 2.7653 | 4.3309 | 3.8125 | 3.8481 | 2.5002 | 5.6409 | 1.7715 | 5.9819 | 5.7130 | 1.7744 | |
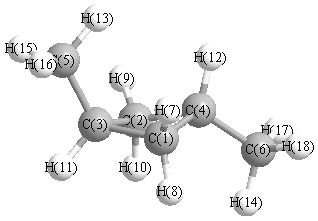 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.394 |
|
C1 |
C3 |
C5 |
118.622 |
| C1 |
C3 |
H11 |
110.051 |
|
C1 |
C4 |
C2 |
88.228 |
| C1 |
C4 |
C6 |
114.157 |
|
C1 |
C4 |
H12 |
114.844 |
| C2 |
C3 |
C5 |
118.622 |
|
C2 |
C3 |
H11 |
110.051 |
| C2 |
C4 |
C6 |
114.157 |
|
C2 |
C4 |
H12 |
114.844 |
| C3 |
C1 |
C4 |
89.237 |
|
C3 |
C1 |
H7 |
111.561 |
| C3 |
C1 |
H8 |
117.445 |
|
C3 |
C2 |
C4 |
89.237 |
| C3 |
C2 |
H9 |
111.561 |
|
C3 |
C2 |
H10 |
117.445 |
| C3 |
C5 |
H13 |
110.750 |
|
C3 |
C5 |
H15 |
111.243 |
| C3 |
C5 |
H16 |
111.243 |
|
C4 |
C1 |
H7 |
112.064 |
| C4 |
C1 |
H8 |
117.045 |
|
C4 |
C2 |
H9 |
112.064 |
| C4 |
C2 |
H10 |
117.045 |
|
C4 |
C6 |
H14 |
111.253 |
| C4 |
C6 |
H17 |
111.009 |
|
C4 |
C6 |
H18 |
111.009 |
| C5 |
C3 |
H11 |
109.507 |
|
C6 |
C4 |
H12 |
109.432 |
| H7 |
C1 |
H8 |
108.505 |
|
H9 |
C2 |
H10 |
108.505 |
| H13 |
C5 |
H15 |
107.814 |
|
H13 |
C5 |
H16 |
107.814 |
| H14 |
C6 |
H17 |
107.705 |
|
H14 |
C6 |
H18 |
107.705 |
| H15 |
C5 |
H16 |
107.821 |
|
H17 |
C6 |
H18 |
108.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.328 |
|
|
|
| 2 |
C |
-0.328 |
|
|
|
| 3 |
C |
-0.130 |
|
|
|
| 4 |
C |
-0.147 |
|
|
|
| 5 |
C |
-0.524 |
|
|
|
| 6 |
C |
-0.522 |
|
|
|
| 7 |
H |
0.162 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.158 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.167 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
| 17 |
H |
0.171 |
|
|
|
| 18 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.022 |
0.001 |
0.000 |
0.022 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.023 |
0.368 |
0.000 |
| y |
0.368 |
-39.967 |
0.000 |
| z |
0.000 |
0.000 |
-39.743 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.832 |
0.368 |
0.000 |
| y |
0.368 |
-0.584 |
0.000 |
| z |
0.000 |
0.000 |
-0.248 |
|
| Polar |
|---|
| 3z2-r2 | -0.496 |
|---|
| x2-y2 | 0.944 |
|---|
| xy | 0.368 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.473 |
-0.574 |
0.000 |
| y |
-0.574 |
10.018 |
0.000 |
| z |
0.000 |
0.000 |
8.770 |
<r2> (average value of r
2) Å
2
| <r2> |
190.740 |
| (<r2>)1/2 |
13.811 |
