Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.585928 |
| Energy at 298.15K | -235.599330 |
| HF Energy | -235.585928 |
| Nuclear repulsion energy | 250.063927 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
2993 |
104.86 |
|
|
|
| 2 |
A' |
3131 |
2988 |
12.96 |
|
|
|
| 3 |
A' |
3127 |
2984 |
24.61 |
|
|
|
| 4 |
A' |
3087 |
2946 |
21.87 |
|
|
|
| 5 |
A' |
3075 |
2935 |
12.76 |
|
|
|
| 6 |
A' |
3069 |
2929 |
6.07 |
|
|
|
| 7 |
A' |
3051 |
2911 |
30.15 |
|
|
|
| 8 |
A' |
3050 |
2910 |
27.08 |
|
|
|
| 9 |
A' |
1512 |
1443 |
8.96 |
|
|
|
| 10 |
A' |
1505 |
1436 |
2.16 |
|
|
|
| 11 |
A' |
1501 |
1433 |
2.21 |
|
|
|
| 12 |
A' |
1422 |
1357 |
8.79 |
|
|
|
| 13 |
A' |
1421 |
1356 |
1.08 |
|
|
|
| 14 |
A' |
1388 |
1324 |
0.43 |
|
|
|
| 15 |
A' |
1372 |
1310 |
10.19 |
|
|
|
| 16 |
A' |
1289 |
1231 |
1.90 |
|
|
|
| 17 |
A' |
1263 |
1205 |
0.56 |
|
|
|
| 18 |
A' |
1209 |
1154 |
0.24 |
|
|
|
| 19 |
A' |
1153 |
1100 |
0.33 |
|
|
|
| 20 |
A' |
1063 |
1014 |
0.37 |
|
|
|
| 21 |
A' |
967 |
922 |
1.26 |
|
|
|
| 22 |
A' |
956 |
912 |
1.08 |
|
|
|
| 23 |
A' |
876 |
836 |
1.37 |
|
|
|
| 24 |
A' |
819 |
782 |
1.06 |
|
|
|
| 25 |
A' |
650 |
620 |
2.01 |
|
|
|
| 26 |
A' |
427 |
408 |
0.13 |
|
|
|
| 27 |
A' |
328 |
313 |
0.13 |
|
|
|
| 28 |
A' |
99 |
95 |
0.01 |
|
|
|
| 29 |
A" |
3135 |
2992 |
32.62 |
|
|
|
| 30 |
A" |
3133 |
2990 |
24.35 |
|
|
|
| 31 |
A" |
3128 |
2985 |
9.96 |
|
|
|
| 32 |
A" |
3068 |
2927 |
66.44 |
|
|
|
| 33 |
A" |
1507 |
1439 |
3.83 |
|
|
|
| 34 |
A" |
1506 |
1437 |
0.15 |
|
|
|
| 35 |
A" |
1478 |
1410 |
3.38 |
|
|
|
| 36 |
A" |
1304 |
1244 |
0.71 |
|
|
|
| 37 |
A" |
1280 |
1221 |
0.00 |
|
|
|
| 38 |
A" |
1241 |
1184 |
0.10 |
|
|
|
| 39 |
A" |
1152 |
1100 |
0.63 |
|
|
|
| 40 |
A" |
1140 |
1088 |
0.09 |
|
|
|
| 41 |
A" |
1002 |
956 |
3.88 |
|
|
|
| 42 |
A" |
927 |
884 |
0.01 |
|
|
|
| 43 |
A" |
918 |
876 |
0.01 |
|
|
|
| 44 |
A" |
811 |
774 |
0.39 |
|
|
|
| 45 |
A" |
375 |
358 |
0.01 |
|
|
|
| 46 |
A" |
261 |
249 |
0.16 |
|
|
|
| 47 |
A" |
256 |
244 |
0.00 |
|
|
|
| 48 |
A" |
236 |
225 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36900.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 35214.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.257 |
0.086 |
1.078 |
| C2 |
0.257 |
0.086 |
-1.078 |
| C3 |
-0.660 |
0.709 |
0.000 |
| C4 |
0.851 |
-0.854 |
0.000 |
| C5 |
-0.865 |
2.211 |
0.000 |
| C6 |
0.257 |
-2.253 |
0.000 |
| H7 |
0.995 |
0.809 |
1.445 |
| H8 |
-0.225 |
-0.386 |
1.941 |
| H9 |
0.995 |
0.809 |
-1.445 |
| H10 |
-0.225 |
-0.386 |
-1.941 |
| H11 |
-1.635 |
0.206 |
0.000 |
| H12 |
1.944 |
-0.922 |
0.000 |
| H13 |
0.098 |
2.734 |
0.000 |
| H14 |
-0.838 |
-2.220 |
0.000 |
| H15 |
-1.423 |
2.536 |
-0.885 |
| H16 |
-1.423 |
2.536 |
0.885 |
| H17 |
0.573 |
-2.815 |
-0.886 |
| H18 |
0.573 |
-2.815 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1562 | 1.5465 | 1.5489 | 2.6336 | 2.5761 | 1.0957 | 1.0954 | 2.7260 | 3.0935 | 2.1809 | 2.2418 | 2.8630 | 2.7714 | 3.5607 | 2.9768 | 3.5176 | 2.9244 |
C2 | 2.1562 | | 1.5465 | 1.5489 | 2.6336 | 2.5761 | 2.7260 | 3.0935 | 1.0957 | 1.0954 | 2.1809 | 2.2418 | 2.8630 | 2.7714 | 2.9768 | 3.5607 | 2.9244 | 3.5176 | C3 | 1.5465 | 1.5465 | | 2.1740 | 1.5159 | 3.1010 | 2.1990 | 2.2706 | 2.1990 | 2.2706 | 1.0964 | 3.0732 | 2.1623 | 2.9340 | 2.1689 | 2.1689 | 3.8368 | 3.8368 | C4 | 1.5489 | 1.5489 | 2.1740 | | 3.5127 | 1.5200 | 2.2079 | 2.2678 | 2.2079 | 2.2678 | 2.7023 | 1.0955 | 3.6660 | 2.1722 | 4.1770 | 4.1770 | 2.1694 | 2.1694 | C5 | 2.6336 | 2.6336 | 1.5159 | 3.5127 | | 4.6031 | 2.7405 | 3.3048 | 2.7405 | 3.3048 | 2.1473 | 4.2083 | 1.0960 | 4.4307 | 1.0956 | 1.0956 | 5.3018 | 5.3018 | C6 | 2.5761 | 2.5761 | 3.1010 | 1.5200 | 4.6031 | | 3.4656 | 2.7360 | 3.4656 | 2.7360 | 3.1030 | 2.1490 | 4.9896 | 1.0961 | 5.1522 | 5.1522 | 1.0954 | 1.0954 | H7 | 1.0957 | 2.7260 | 2.1990 | 2.2079 | 2.7405 | 3.4656 | | 1.7780 | 2.8897 | 3.7919 | 3.0600 | 2.4470 | 2.5680 | 3.8238 | 3.7755 | 3.0230 | 4.3294 | 3.6908 | H8 | 1.0954 | 3.0935 | 2.2706 | 2.2678 | 3.3048 | 2.7360 | 1.7780 | | 3.7919 | 3.8818 | 2.4711 | 2.9595 | 3.6885 | 2.7396 | 4.2382 | 3.3302 | 3.8113 | 2.7651 | H9 | 2.7260 | 1.0957 | 2.1990 | 2.2079 | 2.7405 | 3.4656 | 2.8897 | 3.7919 | | 1.7780 | 3.0600 | 2.4470 | 2.5680 | 3.8238 | 3.0230 | 3.7755 | 3.6908 | 4.3294 | H10 | 3.0935 | 1.0954 | 2.2706 | 2.2678 | 3.3048 | 2.7360 | 3.7919 | 3.8818 | 1.7780 | | 2.4711 | 2.9595 | 3.6885 | 2.7396 | 3.3302 | 4.2382 | 2.7651 | 3.8113 | H11 | 2.1809 | 2.1809 | 1.0964 | 2.7023 | 2.1473 | 3.1030 | 3.0600 | 2.4711 | 3.0600 | 2.4711 | | 3.7527 | 3.0645 | 2.5533 | 2.5014 | 2.5014 | 3.8450 | 3.8450 | H12 | 2.2418 | 2.2418 | 3.0732 | 1.0955 | 4.2083 | 2.1490 | 2.4470 | 2.9595 | 2.4470 | 2.9595 | 3.7527 | | 4.0957 | 3.0703 | 4.9073 | 4.9073 | 2.4994 | 2.4994 | H13 | 2.8630 | 2.8630 | 2.1623 | 3.6660 | 1.0960 | 4.9896 | 2.5680 | 3.6885 | 2.5680 | 3.6885 | 3.0645 | 4.0957 | | 5.0412 | 1.7708 | 1.7708 | 5.6387 | 5.6387 | H14 | 2.7714 | 2.7714 | 2.9340 | 2.1722 | 4.4307 | 1.0961 | 3.8238 | 2.7396 | 3.8238 | 2.7396 | 2.5533 | 3.0703 | 5.0412 | | 4.8727 | 4.8727 | 1.7694 | 1.7694 | H15 | 3.5607 | 2.9768 | 2.1689 | 4.1770 | 1.0956 | 5.1522 | 3.7755 | 4.2382 | 3.0230 | 3.3302 | 2.5014 | 4.9073 | 1.7708 | 4.8727 | | 1.7704 | 5.7108 | 5.9792 | H16 | 2.9768 | 3.5607 | 2.1689 | 4.1770 | 1.0956 | 5.1522 | 3.0230 | 3.3302 | 3.7755 | 4.2382 | 2.5014 | 4.9073 | 1.7708 | 4.8727 | 1.7704 | | 5.9792 | 5.7108 | H17 | 3.5176 | 2.9244 | 3.8368 | 2.1694 | 5.3018 | 1.0954 | 4.3294 | 3.8113 | 3.6908 | 2.7651 | 3.8450 | 2.4994 | 5.6387 | 1.7694 | 5.7108 | 5.9792 | | 1.7723 | H18 | 2.9244 | 3.5176 | 3.8368 | 2.1694 | 5.3018 | 1.0954 | 3.6908 | 2.7651 | 4.3294 | 3.8113 | 3.8450 | 2.4994 | 5.6387 | 1.7694 | 5.9792 | 5.7108 | 1.7723 | |
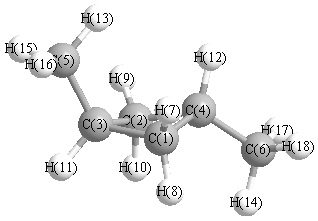 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.393 |
|
C1 |
C3 |
C5 |
118.620 |
| C1 |
C3 |
H11 |
110.046 |
|
C1 |
C4 |
C2 |
88.221 |
| C1 |
C4 |
C6 |
114.157 |
|
C1 |
C4 |
H12 |
114.852 |
| C2 |
C3 |
C5 |
118.620 |
|
C2 |
C3 |
H11 |
110.046 |
| C2 |
C4 |
C6 |
114.157 |
|
C2 |
C4 |
H12 |
114.852 |
| C3 |
C1 |
C4 |
89.229 |
|
C3 |
C1 |
H7 |
111.527 |
| C3 |
C1 |
H8 |
117.495 |
|
C3 |
C2 |
C4 |
89.229 |
| C3 |
C2 |
H9 |
111.527 |
|
C3 |
C2 |
H10 |
117.495 |
| C3 |
C5 |
H13 |
110.728 |
|
C3 |
C5 |
H15 |
111.281 |
| C3 |
C5 |
H16 |
111.281 |
|
C4 |
C1 |
H7 |
112.068 |
| C4 |
C1 |
H8 |
117.062 |
|
C4 |
C2 |
H9 |
112.068 |
| C4 |
C2 |
H10 |
117.062 |
|
C4 |
C6 |
H14 |
111.233 |
| C4 |
C6 |
H17 |
111.045 |
|
C4 |
C6 |
H18 |
111.045 |
| C5 |
C3 |
H11 |
109.518 |
|
C6 |
C4 |
H12 |
109.425 |
| H7 |
C1 |
H8 |
108.480 |
|
H9 |
C2 |
H10 |
108.480 |
| H13 |
C5 |
H15 |
107.801 |
|
H13 |
C5 |
H16 |
107.801 |
| H14 |
C6 |
H17 |
107.685 |
|
H14 |
C6 |
H18 |
107.685 |
| H15 |
C5 |
H16 |
107.791 |
|
H17 |
C6 |
H18 |
107.988 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.226 |
|
|
|
| 2 |
C |
-0.226 |
|
|
|
| 3 |
C |
-0.110 |
|
|
|
| 4 |
C |
-0.128 |
|
|
|
| 5 |
C |
-0.386 |
|
|
|
| 6 |
C |
-0.380 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
| 10 |
H |
0.116 |
|
|
|
| 11 |
H |
0.120 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
| 13 |
H |
0.120 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
| 15 |
H |
0.128 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.026 |
0.003 |
0.000 |
0.026 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.074 |
0.330 |
0.000 |
| y |
0.330 |
-39.881 |
0.000 |
| z |
0.000 |
0.000 |
-39.746 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.739 |
0.330 |
0.000 |
| y |
0.330 |
-0.471 |
0.000 |
| z |
0.000 |
0.000 |
-0.268 |
|
| Polar |
|---|
| 3z2-r2 | -0.536 |
|---|
| x2-y2 | 0.807 |
|---|
| xy | 0.330 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.602 |
-0.587 |
0.000 |
| y |
-0.587 |
10.190 |
0.000 |
| z |
0.000 |
0.000 |
8.895 |
<r2> (average value of r
2) Å
2
| <r2> |
190.618 |
| (<r2>)1/2 |
13.806 |
