Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -100.234028 |
| Energy at 298.15K | -100.233074 |
| HF Energy | -100.234028 |
| Nuclear repulsion energy | 27.555662 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.075 |
| C2 |
0.000 |
0.000 |
-0.150 |
| N3 |
0.000 |
0.000 |
1.018 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9248 | 3.0936 |
C2 | 1.9248 | | 1.1688 | N3 | 3.0936 | 1.1688 | |
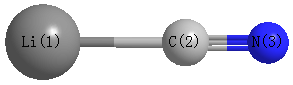 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.485 |
|
|
|
| 2 |
C |
-0.051 |
|
|
|
| 3 |
N |
-0.435 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-8.854 |
8.854 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.107 |
0.000 |
0.000 |
| y |
0.000 |
-14.107 |
0.000 |
| z |
0.000 |
0.000 |
1.642 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.874 |
0.000 |
0.000 |
| y |
0.000 |
-7.874 |
0.000 |
| z |
0.000 |
0.000 |
15.748 |
|
| Polar |
|---|
| 3z2-r2 | 31.496 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.304 |
0.000 |
0.000 |
| y |
0.000 |
2.304 |
0.000 |
| z |
0.000 |
0.000 |
4.351 |
<r2> (average value of r
2) Å
2
| <r2> |
25.848 |
| (<r2>)1/2 |
5.084 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -100.239543 |
| Energy at 298.15K | -100.238831 |
| HF Energy | -100.239543 |
| Nuclear repulsion energy | 29.197293 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.423 |
-0.619 |
0.000 |
| C2 |
-0.712 |
-0.366 |
0.000 |
| N3 |
0.000 |
0.579 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1500 | 1.8602 |
C2 | 2.1500 | | 1.1826 | N3 | 1.8602 | 1.1826 | |
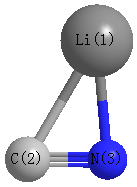 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
59.766 |
|
Li1 |
N3 |
C2 |
86.917 |
| C2 |
Li1 |
N3 |
33.317 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.406 |
|
|
|
| 2 |
C |
0.057 |
|
|
|
| 3 |
N |
-0.463 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.339 |
-2.590 |
0.000 |
6.848 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-5.876 |
-5.931 |
0.000 |
| y |
-5.931 |
-15.034 |
0.000 |
| z |
0.000 |
0.000 |
-14.520 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.901 |
-5.931 |
0.000 |
| y |
-5.931 |
-4.836 |
0.000 |
| z |
0.000 |
0.000 |
-4.065 |
|
| Polar |
|---|
| 3z2-r2 | -8.131 |
|---|
| x2-y2 | 9.158 |
|---|
| xy | -5.931 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.212 |
0.347 |
0.000 |
| y |
0.347 |
3.229 |
0.000 |
| z |
0.000 |
0.000 |
2.528 |
<r2> (average value of r
2) Å
2
| <r2> |
20.790 |
| (<r2>)1/2 |
4.560 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -100.235722 |
| Energy at 298.15K | |
| HF Energy | -100.235722 |
| Nuclear repulsion energy | 28.248948 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.892 |
| C2 |
0.000 |
0.000 |
-1.073 |
| N3 |
0.000 |
0.000 |
0.109 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9647 | 1.7826 |
C2 | 2.9647 | | 1.1821 | N3 | 1.7826 | 1.1821 | |
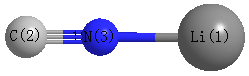 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.587 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
N |
-0.505 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
8.584 |
8.584 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.040 |
0.000 |
0.000 |
| y |
0.000 |
-14.040 |
0.000 |
| z |
0.000 |
0.000 |
-1.606 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.217 |
0.000 |
0.000 |
| y |
0.000 |
-6.217 |
0.000 |
| z |
0.000 |
0.000 |
12.434 |
|
| Polar |
|---|
| 3z2-r2 | 24.868 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.234 |
0.000 |
0.000 |
| y |
0.000 |
2.234 |
0.000 |
| z |
0.000 |
0.000 |
4.432 |
<r2> (average value of r
2) Å
2
| <r2> |
23.907 |
| (<r2>)1/2 |
4.889 |
