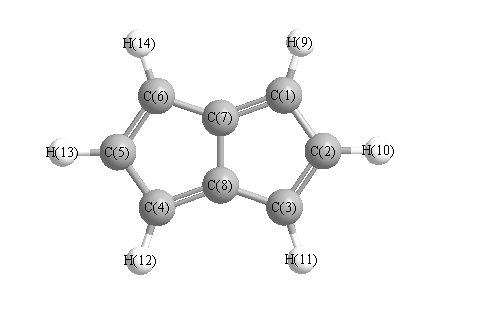
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3283 |
3129 |
0.00 |
|
|
|
| 2 |
Ag |
3261 |
3108 |
0.00 |
|
|
|
| 3 |
Ag |
3247 |
3094 |
0.00 |
|
|
|
| 4 |
Ag |
1692 |
1613 |
0.00 |
|
|
|
| 5 |
Ag |
1597 |
1522 |
0.00 |
|
|
|
| 6 |
Ag |
1414 |
1347 |
0.00 |
|
|
|
| 7 |
Ag |
1345 |
1282 |
0.00 |
|
|
|
| 8 |
Ag |
1211 |
1154 |
0.00 |
|
|
|
| 9 |
Ag |
1120 |
1068 |
0.00 |
|
|
|
| 10 |
Ag |
979 |
933 |
0.00 |
|
|
|
| 11 |
Ag |
863 |
822 |
0.00 |
|
|
|
| 12 |
Ag |
711 |
678 |
0.00 |
|
|
|
| 13 |
Ag |
662 |
631 |
0.00 |
|
|
|
| 14 |
Au |
930 |
886 |
5.95 |
|
|
|
| 15 |
Au |
884 |
842 |
15.13 |
|
|
|
| 16 |
Au |
729 |
695 |
87.82 |
|
|
|
| 17 |
Au |
513 |
489 |
7.53 |
|
|
|
| 18 |
Au |
300 |
286 |
1.39 |
|
|
|
| 19 |
Au |
155 |
148 |
5.09 |
|
|
|
| 20 |
Bg |
927 |
883 |
0.00 |
|
|
|
| 21 |
Bg |
918 |
875 |
0.00 |
|
|
|
| 22 |
Bg |
793 |
756 |
0.00 |
|
|
|
| 23 |
Bg |
668 |
637 |
0.00 |
|
|
|
| 24 |
Bg |
553 |
527 |
0.00 |
|
|
|
| 25 |
Bu |
3283 |
3129 |
24.48 |
|
|
|
| 26 |
Bu |
3261 |
3108 |
15.47 |
|
|
|
| 27 |
Bu |
3247 |
3095 |
13.67 |
|
|
|
| 28 |
Bu |
1693 |
1613 |
49.03 |
|
|
|
| 29 |
Bu |
1582 |
1507 |
15.55 |
|
|
|
| 30 |
Bu |
1360 |
1297 |
19.56 |
|
|
|
| 31 |
Bu |
1285 |
1224 |
19.72 |
|
|
|
| 32 |
Bu |
1141 |
1088 |
45.48 |
|
|
|
| 33 |
Bu |
1071 |
1020 |
6.78 |
|
|
|
| 34 |
Bu |
970 |
925 |
6.76 |
|
|
|
| 35 |
Bu |
846 |
806 |
9.34 |
|
|
|
| 36 |
Bu |
449 |
428 |
3.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24470.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 23322.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
C |
-0.225 |
|
|
|
| 3 |
C |
-0.139 |
|
|
|
| 4 |
C |
-0.209 |
|
|
|
| 5 |
C |
-0.225 |
|
|
|
| 6 |
C |
-0.139 |
|
|
|
| 7 |
C |
0.045 |
|
|
|
| 8 |
C |
0.045 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.173 |
|
|
|
| 11 |
H |
0.176 |
|
|
|
| 12 |
H |
0.179 |
|
|
|
| 13 |
H |
0.173 |
|
|
|
| 14 |
H |
0.176 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.695 |
1.221 |
0.000 |
| y |
1.221 |
-39.631 |
0.000 |
| z |
0.000 |
0.000 |
-49.798 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.019 |
1.221 |
0.000 |
| y |
1.221 |
5.116 |
0.000 |
| z |
0.000 |
0.000 |
-10.135 |
|
| Polar |
|---|
| 3z2-r2 | -20.269 |
|---|
| x2-y2 | -0.064 |
|---|
| xy | 1.221 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
17.176 |
1.108 |
0.000 |
| y |
1.108 |
11.774 |
0.000 |
| z |
0.000 |
0.000 |
4.047 |
<r2> (average value of r
2) Å
2
| <r2> |
216.872 |
| (<r2>)1/2 |
14.727 |