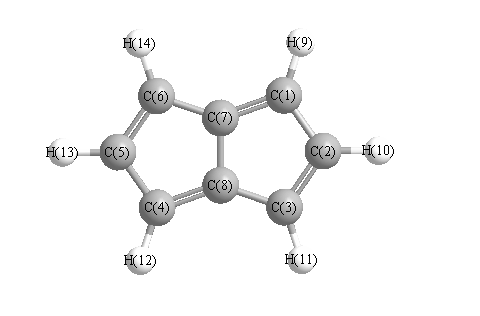
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3277 |
3127 |
0.00 |
|
|
|
| 2 |
Ag |
3255 |
3106 |
0.00 |
|
|
|
| 3 |
Ag |
3241 |
3093 |
0.00 |
|
|
|
| 4 |
Ag |
1688 |
1611 |
0.00 |
|
|
|
| 5 |
Ag |
1592 |
1519 |
0.00 |
|
|
|
| 6 |
Ag |
1409 |
1345 |
0.00 |
|
|
|
| 7 |
Ag |
1341 |
1279 |
0.00 |
|
|
|
| 8 |
Ag |
1206 |
1151 |
0.00 |
|
|
|
| 9 |
Ag |
1116 |
1065 |
0.00 |
|
|
|
| 10 |
Ag |
975 |
930 |
0.00 |
|
|
|
| 11 |
Ag |
861 |
822 |
0.00 |
|
|
|
| 12 |
Ag |
711 |
679 |
0.00 |
|
|
|
| 13 |
Ag |
661 |
631 |
0.00 |
|
|
|
| 14 |
Au |
931 |
889 |
5.24 |
|
|
|
| 15 |
Au |
885 |
845 |
14.65 |
|
|
|
| 16 |
Au |
730 |
696 |
83.18 |
|
|
|
| 17 |
Au |
513 |
490 |
7.46 |
|
|
|
| 18 |
Au |
300 |
286 |
1.27 |
|
|
|
| 19 |
Au |
155 |
148 |
4.94 |
|
|
|
| 20 |
Bg |
929 |
886 |
0.00 |
|
|
|
| 21 |
Bg |
920 |
877 |
0.00 |
|
|
|
| 22 |
Bg |
793 |
757 |
0.00 |
|
|
|
| 23 |
Bg |
668 |
638 |
0.00 |
|
|
|
| 24 |
Bg |
553 |
528 |
0.00 |
|
|
|
| 25 |
Bu |
3277 |
3127 |
23.91 |
|
|
|
| 26 |
Bu |
3256 |
3107 |
12.50 |
|
|
|
| 27 |
Bu |
3242 |
3094 |
13.40 |
|
|
|
| 28 |
Bu |
1688 |
1611 |
49.85 |
|
|
|
| 29 |
Bu |
1578 |
1506 |
15.35 |
|
|
|
| 30 |
Bu |
1355 |
1293 |
19.37 |
|
|
|
| 31 |
Bu |
1279 |
1220 |
19.41 |
|
|
|
| 32 |
Bu |
1137 |
1085 |
45.58 |
|
|
|
| 33 |
Bu |
1069 |
1020 |
6.10 |
|
|
|
| 34 |
Bu |
967 |
922 |
6.96 |
|
|
|
| 35 |
Bu |
846 |
807 |
9.28 |
|
|
|
| 36 |
Bu |
449 |
428 |
3.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24424.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 23308.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.151 |
|
|
|
| 2 |
C |
-0.175 |
|
|
|
| 3 |
C |
-0.084 |
|
|
|
| 4 |
C |
-0.151 |
|
|
|
| 5 |
C |
-0.175 |
|
|
|
| 6 |
C |
-0.084 |
|
|
|
| 7 |
C |
0.024 |
|
|
|
| 8 |
C |
0.024 |
|
|
|
| 9 |
H |
0.132 |
|
|
|
| 10 |
H |
0.125 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.132 |
|
|
|
| 13 |
H |
0.125 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.840 |
1.210 |
0.000 |
| y |
1.210 |
-39.720 |
0.000 |
| z |
0.000 |
0.000 |
-49.677 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.858 |
1.210 |
0.000 |
| y |
1.210 |
5.039 |
0.000 |
| z |
0.000 |
0.000 |
-9.897 |
|
| Polar |
|---|
| 3z2-r2 | -19.794 |
|---|
| x2-y2 | -0.121 |
|---|
| xy | 1.210 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
17.304 |
1.111 |
0.000 |
| y |
1.111 |
11.880 |
0.000 |
| z |
0.000 |
0.000 |
4.104 |
<r2> (average value of r
2) Å
2
| <r2> |
216.834 |
| (<r2>)1/2 |
14.725 |