return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Cyclobutylamine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-211.904689
Energy at 298.15K-211.916023
HF Energy-211.137136
Nuclear repulsion energy188.573160
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3558 3325 0.72      
2 A' 3250 3037 31.51      
3 A' 3215 3004 26.18      
4 A' 3171 2963 32.97      
5 A' 3149 2942 14.54      
6 A' 3075 2873 84.40      
7 A' 1693 1582 21.61      
8 A' 1563 1460 3.77      
9 A' 1535 1435 2.22      
10 A' 1433 1339 23.86      
11 A' 1335 1248 4.62      
12 A' 1279 1195 1.03      
13 A' 1197 1119 7.18      
14 A' 1125 1051 7.70      
15 A' 1003 937 9.95      
16 A' 932 871 28.92      
17 A' 911 851 41.86      
18 A' 860 804 59.23      
19 A' 680 636 1.77      
20 A' 416 389 4.34      
21 A' 213 199 2.00      
22 A" 3662 3422 0.14      
23 A" 3217 3006 11.49      
24 A" 3145 2938 57.07      
25 A" 1528 1428 0.69      
26 A" 1388 1297 0.09      
27 A" 1317 1231 0.61      
28 A" 1296 1211 0.14      
29 A" 1259 1177 0.41      
30 A" 1197 1118 0.85      
31 A" 1062 992 0.18      
32 A" 976 912 0.48      
33 A" 954 892 1.88      
34 A" 801 748 0.99      
35 A" 398 372 12.66      
36 A" 297 277 31.38      

Unscaled Zero Point Vibrational Energy (zpe) 29044.0 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 27138.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.27639 0.16222 0.13671

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.476 0.718 0.000
H2 -1.835 1.211 0.811
H3 -1.835 1.211 -0.811
C4 0.542 -0.172 -1.068
H5 -0.055 -0.314 -1.968
H6 1.556 0.110 -1.349
C7 0.542 -0.172 1.068
H8 -0.055 -0.314 1.968
H9 1.556 0.110 1.349
C10 -0.018 0.788 0.000
H11 0.384 1.808 0.000
C12 0.542 -1.284 0.000
H13 1.367 -1.992 0.000
H14 -0.401 -1.821 0.000

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 C7 H8 H9 C10 H11 C12 H13 H14
N11.01461.01462.45072.63733.37402.45072.63733.37401.45962.15492.84273.92842.7571
H21.01461.62233.33113.63524.16902.76222.61353.60622.03472.43683.54014.60163.4502
H31.01461.62232.76222.61353.60623.33113.63524.16902.03472.43683.54014.60163.4502
C42.45073.33112.76221.08891.08942.13673.09742.63641.54122.25481.54202.26642.1795
H52.63733.63522.61351.08891.77763.09743.93523.71152.25522.92652.27352.95172.5023
H63.37404.16903.60621.08941.77762.63643.71152.69762.18102.46502.18862.50493.0625
C72.45072.76223.33112.13673.09742.63641.08891.08941.54122.25481.54202.26642.1795
H82.63732.61353.63523.09743.93523.71151.08891.77762.25522.92652.27352.95172.5023
H93.37403.60624.16902.63643.71152.69761.08941.77762.18102.46502.18862.50493.0625
C101.45962.03472.03471.54122.25522.18101.54122.25522.18101.09612.14563.10592.6363
H112.15492.43682.43682.25482.92652.46502.25482.92652.46501.09613.09523.92513.7121
C122.84273.54013.54011.54202.27352.18861.54202.27352.18862.14563.09521.08771.0856
H133.92844.60164.60162.26642.95172.50492.26642.95172.50493.10593.92511.08771.7770
H142.75713.45023.45022.17952.50233.06252.17952.50233.06252.63633.71211.08561.7770

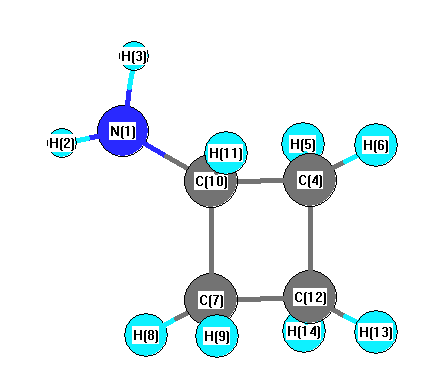
picture of Cyclobutylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C10 C4 109.479 N1 C10 C7 109.479
N1 C10 H11 114.202 H2 N1 H3 106.155
H2 N1 C10 109.324 H3 N1 C10 109.324
C4 C10 C7 87.767 C4 C10 H11 116.494
C4 C12 C7 87.714 C4 C12 H13 118.020
C4 C12 H14 110.897 H5 C4 H6 109.387
H5 C4 C10 117.024 H5 C4 C12 118.560
H6 C4 C10 110.841 H6 C4 C12 111.396
C7 C10 H11 116.494 C7 C12 H13 118.020
C7 C12 H14 110.897 H8 C7 H9 109.387
H8 C7 C10 117.024 H8 C7 C12 118.560
H9 C7 C10 110.841 H9 C7 C12 111.396
C10 C4 C12 88.198 C10 C7 C12 88.198
H13 C12 H14 109.697
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability