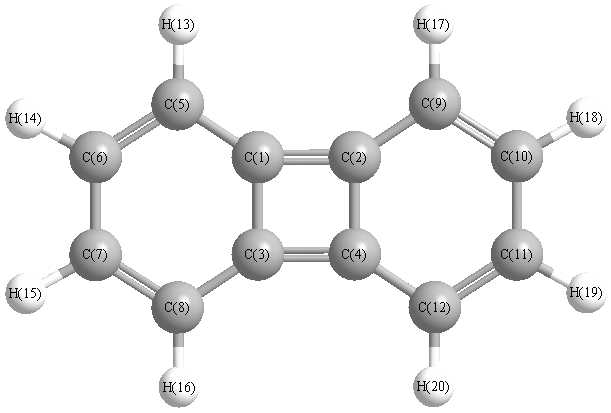
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3241 |
3089 |
0.00 |
|
|
|
| 2 |
Ag |
3225 |
3074 |
0.00 |
|
|
|
| 3 |
Ag |
1753 |
1671 |
0.00 |
|
|
|
| 4 |
Ag |
1553 |
1480 |
0.00 |
|
|
|
| 5 |
Ag |
1459 |
1390 |
0.00 |
|
|
|
| 6 |
Ag |
1206 |
1150 |
0.00 |
|
|
|
| 7 |
Ag |
1168 |
1113 |
0.00 |
|
|
|
| 8 |
Ag |
1036 |
987 |
0.00 |
|
|
|
| 9 |
Ag |
790 |
753 |
0.00 |
|
|
|
| 10 |
Ag |
405 |
386 |
0.00 |
|
|
|
| 11 |
Au |
981 |
935 |
0.00 |
|
|
|
| 12 |
Au |
904 |
862 |
0.00 |
|
|
|
| 13 |
Au |
815 |
777 |
0.00 |
|
|
|
| 14 |
Au |
588 |
561 |
0.00 |
|
|
|
| 15 |
Au |
149 |
142 |
0.00 |
|
|
|
| 16 |
B1g |
981 |
935 |
0.00 |
|
|
|
| 17 |
B1g |
890 |
848 |
0.00 |
|
|
|
| 18 |
B1g |
722 |
688 |
0.00 |
|
|
|
| 19 |
B1g |
465 |
443 |
0.00 |
|
|
|
| 20 |
B1u |
3240 |
3088 |
38.09 |
|
|
|
| 21 |
B1u |
3224 |
3073 |
17.17 |
|
|
|
| 22 |
B1u |
1677 |
1598 |
0.33 |
|
|
|
| 23 |
B1u |
1488 |
1418 |
36.93 |
|
|
|
| 24 |
B1u |
1345 |
1282 |
34.92 |
|
|
|
| 25 |
B1u |
1196 |
1140 |
8.16 |
|
|
|
| 26 |
B1u |
1067 |
1017 |
0.08 |
|
|
|
| 27 |
B1u |
1016 |
969 |
18.36 |
|
|
|
| 28 |
B1u |
626 |
597 |
4.61 |
|
|
|
| 29 |
B2g |
934 |
890 |
0.00 |
|
|
|
| 30 |
B2g |
765 |
729 |
0.00 |
|
|
|
| 31 |
B2g |
445 |
424 |
0.00 |
|
|
|
| 32 |
B2g |
325 |
310 |
0.00 |
|
|
|
| 33 |
B2u |
3235 |
3083 |
43.53 |
|
|
|
| 34 |
B2u |
3212 |
3061 |
0.51 |
|
|
|
| 35 |
B2u |
1677 |
1598 |
1.94 |
|
|
|
| 36 |
B2u |
1504 |
1433 |
6.16 |
|
|
|
| 37 |
B2u |
1310 |
1249 |
3.37 |
|
|
|
| 38 |
B2u |
1166 |
1111 |
11.52 |
|
|
|
| 39 |
B2u |
1083 |
1032 |
1.13 |
|
|
|
| 40 |
B2u |
741 |
706 |
0.48 |
|
|
|
| 41 |
B2u |
217 |
206 |
0.80 |
|
|
|
| 42 |
B3g |
3234 |
3083 |
0.00 |
|
|
|
| 43 |
B3g |
3212 |
3061 |
0.00 |
|
|
|
| 44 |
B3g |
1681 |
1602 |
0.00 |
|
|
|
| 45 |
B3g |
1504 |
1434 |
0.00 |
|
|
|
| 46 |
B3g |
1320 |
1258 |
0.00 |
|
|
|
| 47 |
B3g |
1130 |
1077 |
0.00 |
|
|
|
| 48 |
B3g |
1000 |
953 |
0.00 |
|
|
|
| 49 |
B3g |
613 |
584 |
0.00 |
|
|
|
| 50 |
B3g |
569 |
542 |
0.00 |
|
|
|
| 51 |
B3u |
929 |
885 |
6.82 |
|
|
|
| 52 |
B3u |
753 |
718 |
124.67 |
|
|
|
| 53 |
B3u |
379 |
361 |
1.87 |
|
|
|
| 54 |
B3u |
108 |
103 |
2.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35125.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 33478.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.079 |
|
|
|
| 2 |
C |
0.079 |
|
|
|
| 3 |
C |
0.079 |
|
|
|
| 4 |
C |
0.079 |
|
|
|
| 5 |
C |
-0.250 |
|
|
|
| 6 |
C |
-0.179 |
|
|
|
| 7 |
C |
-0.179 |
|
|
|
| 8 |
C |
-0.250 |
|
|
|
| 9 |
C |
-0.250 |
|
|
|
| 10 |
C |
-0.179 |
|
|
|
| 11 |
C |
-0.179 |
|
|
|
| 12 |
C |
-0.250 |
|
|
|
| 13 |
H |
0.179 |
|
|
|
| 14 |
H |
0.171 |
|
|
|
| 15 |
H |
0.171 |
|
|
|
| 16 |
H |
0.179 |
|
|
|
| 17 |
H |
0.179 |
|
|
|
| 18 |
H |
0.171 |
|
|
|
| 19 |
H |
0.171 |
|
|
|
| 20 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-73.486 |
0.000 |
0.000 |
| y |
0.000 |
-59.009 |
0.000 |
| z |
0.000 |
0.000 |
-58.201 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-14.880 |
0.000 |
0.000 |
| y |
0.000 |
6.834 |
0.000 |
| z |
0.000 |
0.000 |
8.046 |
|
| Polar |
|---|
| 3z2-r2 | 16.092 |
|---|
| x2-y2 | -14.476 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.769 |
0.000 |
0.000 |
| y |
0.000 |
18.466 |
0.000 |
| z |
0.000 |
0.000 |
30.347 |
<r2> (average value of r
2) Å
2
| <r2> |
558.785 |
| (<r2>)1/2 |
23.639 |