Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.024155 |
| Energy at 298.15K | -131.023773 |
| HF Energy | -130.679057 |
| Nuclear repulsion energy | 47.413442 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3408 |
3184 |
13.21 |
|
|
|
| 2 |
A' |
2288 |
2138 |
1.71 |
|
|
|
| 3 |
A' |
1084 |
1013 |
29.79 |
|
|
|
| 4 |
A' |
864 |
808 |
7.98 |
|
|
|
| 5 |
A' |
453 |
423 |
4.91 |
|
|
|
| 6 |
A" |
527 |
492 |
8.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4312.0 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 4029.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.277 |
-1.229 |
0.000 |
| C2 |
0.000 |
0.119 |
0.000 |
| N3 |
0.192 |
1.254 |
0.000 |
| H4 |
0.317 |
-2.121 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3760 | 2.5266 | 1.0719 |
C2 | 1.3760 | | 1.1510 | 2.2626 | N3 | 2.5266 | 1.1510 | | 3.3774 | H4 | 1.0719 | 2.2626 | 3.3774 | |
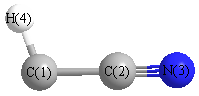 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
177.999 |
|
C2 |
C1 |
H4 |
134.747 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.018079 |
| Energy at 298.15K | |
| HF Energy | -130.678657 |
| Nuclear repulsion energy | 47.700882 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3533 |
3301 |
79.37 |
|
|
|
| 2 |
Σ |
1716 |
1604 |
33.65 |
|
|
|
| 3 |
Σ |
1210 |
1131 |
43.26 |
|
|
|
| 4 |
Π |
513 |
480 |
1.85 |
|
|
|
| 4 |
Π |
513 |
480 |
1.85 |
|
|
|
| 5 |
Π |
560i |
524i |
58.27 |
|
|
|
| 5 |
Π |
560i |
524i |
58.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3182.2 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 2973.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.211 |
| C2 |
0.000 |
0.000 |
0.099 |
| N3 |
0.000 |
0.000 |
1.278 |
| H4 |
0.000 |
0.000 |
-2.273 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3100 | 2.4887 | 1.0615 |
C2 | 1.3100 | | 1.1788 | 2.3715 | N3 | 2.4887 | 1.1788 | | 3.5503 | H4 | 1.0615 | 2.3715 | 3.5503 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.090 |
|
|
|
| 2 |
C |
0.228 |
|
|
|
| 3 |
N |
-0.394 |
|
|
|
| 4 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.691 |
3.691 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.104 |
0.000 |
0.000 |
| y |
0.000 |
-17.104 |
0.000 |
| z |
0.000 |
0.000 |
-15.564 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.770 |
0.000 |
0.000 |
| y |
0.000 |
-0.770 |
0.000 |
| z |
0.000 |
0.000 |
1.540 |
|
| Polar |
|---|
| 3z2-r2 | 3.080 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.589 |
0.000 |
0.000 |
| y |
0.000 |
1.589 |
0.000 |
| z |
0.000 |
0.000 |
6.085 |
<r2> (average value of r
2) Å
2
| <r2> |
35.813 |
| (<r2>)1/2 |
5.984 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.016649 |
| Energy at 298.15K | -131.016076 |
| HF Energy | -130.616619 |
| Nuclear repulsion energy | 46.505319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3141 |
2935 |
21.74 |
|
|
|
| 2 |
A' |
2270 |
2121 |
7.88 |
|
|
|
| 3 |
A' |
1077 |
1006 |
75.38 |
|
|
|
| 4 |
A' |
989 |
925 |
1.10 |
|
|
|
| 5 |
A' |
395 |
369 |
16.57 |
|
|
|
| 6 |
A" |
343 |
321 |
13.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4107.8 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 3838.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.096 |
-1.303 |
0.000 |
| C2 |
0.000 |
0.098 |
0.000 |
| N3 |
-0.248 |
1.259 |
0.000 |
| H4 |
1.158 |
-1.579 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4043 | 2.5851 | 1.0967 |
C2 | 1.4043 | | 1.1873 | 2.0376 | N3 | 2.5851 | 1.1873 | | 3.1669 | H4 | 1.0967 | 2.0376 | 3.1669 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.016649 |
| Energy at 298.15K | -131.016076 |
| HF Energy | -130.616619 |
| Nuclear repulsion energy | 46.505319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
