Jump to
S1C2
S2C1
S2C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.277312 |
| Energy at 298.15K | -131.276641 |
| HF Energy | -131.277312 |
| Nuclear repulsion energy | 47.275445 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3417 |
3261 |
37.50 |
|
|
|
| 2 |
A' |
1831 |
1748 |
35.40 |
|
|
|
| 3 |
A' |
1246 |
1189 |
7.67 |
|
|
|
| 4 |
A' |
488 |
466 |
28.39 |
|
|
|
| 5 |
A' |
392 |
375 |
19.29 |
|
|
|
| 6 |
A" |
449 |
429 |
1.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3911.7 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 3732.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.071 |
-1.223 |
0.000 |
| C2 |
0.000 |
0.087 |
0.000 |
| N3 |
-0.151 |
1.279 |
0.000 |
| H4 |
0.633 |
-2.134 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3113 | 2.5110 | 1.0713 |
C2 | 1.3113 | | 1.2013 | 2.3097 | N3 | 2.5110 | 1.2013 | | 3.5019 | H4 | 1.0713 | 2.3097 | 3.5019 | |
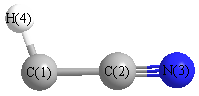 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
175.868 |
|
C2 |
C1 |
H4 |
151.414 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.276626 |
| Energy at 298.15K | |
| HF Energy | -131.276626 |
| Nuclear repulsion energy | 47.332858 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3476 |
3317 |
70.79 |
|
|
|
| 2 |
Σ |
1763 |
1683 |
40.06 |
|
|
|
| 3 |
Σ |
1288 |
1229 |
5.47 |
|
|
|
| 4 |
Π |
451 |
431 |
0.87 |
|
|
|
| 4 |
Π |
451 |
431 |
0.87 |
|
|
|
| 5 |
Π |
301i |
287i |
50.47 |
|
|
|
| 5 |
Π |
301i |
287i |
50.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3414.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 3258.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.211 |
| C2 |
0.000 |
0.000 |
0.082 |
| N3 |
0.000 |
0.000 |
1.293 |
| H4 |
0.000 |
0.000 |
-2.277 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2932 | 2.5035 | 1.0666 |
C2 | 1.2932 | | 1.2103 | 2.3598 | N3 | 2.5035 | 1.2103 | | 3.5701 | H4 | 1.0666 | 2.3598 | 3.5701 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.041 |
|
|
|
| 2 |
C |
0.276 |
|
|
|
| 3 |
N |
-0.470 |
|
|
|
| 4 |
H |
0.235 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.288 |
3.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.900 |
0.000 |
0.000 |
| y |
0.000 |
-16.900 |
0.000 |
| z |
0.000 |
0.000 |
-15.106 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.897 |
0.000 |
0.000 |
| y |
0.000 |
-0.897 |
0.000 |
| z |
0.000 |
0.000 |
1.794 |
|
| Polar |
|---|
| 3z2-r2 | 3.589 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.639 |
0.000 |
0.000 |
| y |
0.000 |
1.639 |
0.000 |
| z |
0.000 |
0.000 |
6.140 |
<r2> (average value of r
2) Å
2
| <r2> |
35.903 |
| (<r2>)1/2 |
5.992 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.243824 |
| Energy at 298.15K | -131.243263 |
| HF Energy | -131.243824 |
| Nuclear repulsion energy | 46.980735 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3099 |
2957 |
10.45 |
|
|
|
| 2 |
A' |
2132 |
2035 |
18.73 |
|
|
|
| 3 |
A' |
1107 |
1056 |
32.77 |
|
|
|
| 4 |
A' |
928 |
885 |
54.08 |
|
|
|
| 5 |
A' |
445 |
425 |
23.90 |
|
|
|
| 6 |
A" |
321 |
306 |
10.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4015.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 3832.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.102 |
-1.279 |
0.000 |
| C2 |
0.000 |
0.091 |
0.000 |
| N3 |
-0.253 |
1.246 |
0.000 |
| H4 |
1.156 |
-1.599 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3740 | 2.5500 | 1.1009 |
C2 | 1.3740 | | 1.1823 | 2.0477 | N3 | 2.5500 | 1.1823 | | 3.1748 | H4 | 1.1009 | 2.0477 | 3.1748 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.081 |
|
|
|
| 2 |
C |
0.311 |
|
|
|
| 3 |
N |
-0.407 |
|
|
|
| 4 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.793 |
-1.381 |
0.000 |
2.263 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.558 |
-2.283 |
0.000 |
| y |
-2.283 |
-20.722 |
0.000 |
| z |
0.000 |
0.000 |
-15.560 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.583 |
-2.283 |
0.000 |
| y |
-2.283 |
-4.663 |
0.000 |
| z |
0.000 |
0.000 |
3.080 |
|
| Polar |
|---|
| 3z2-r2 | 6.161 |
|---|
| x2-y2 | 4.164 |
|---|
| xy | -2.283 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.433 |
-0.858 |
0.000 |
| y |
-0.858 |
6.065 |
0.000 |
| z |
0.000 |
0.000 |
2.195 |
<r2> (average value of r
2) Å
2
| <r2> |
36.139 |
| (<r2>)1/2 |
6.012 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.243824 |
| Energy at 298.15K | -131.243263 |
| HF Energy | -131.243824 |
| Nuclear repulsion energy | 46.980735 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
