Jump to
S1C2
S2C1
S2C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.292972 |
| Energy at 298.15K | -131.292400 |
| HF Energy | -131.170205 |
| Nuclear repulsion energy | 47.221587 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3400 |
3400 |
26.60 |
|
|
|
| 2 |
A' |
1905 |
1905 |
19.06 |
|
|
|
| 3 |
A' |
1189 |
1189 |
13.33 |
|
|
|
| 4 |
A' |
602 |
602 |
34.93 |
|
|
|
| 5 |
A' |
409 |
409 |
7.05 |
|
|
|
| 6 |
A" |
459 |
459 |
3.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3982.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3982.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.002 |
-1.235 |
0.000 |
| C2 |
0.000 |
0.092 |
0.000 |
| N3 |
-0.090 |
1.282 |
0.000 |
| H4 |
0.616 |
-2.113 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3276 | 2.5189 | 1.0709 |
C2 | 1.3276 | | 1.1930 | 2.2899 | N3 | 2.5189 | 1.1930 | | 3.4677 | H4 | 1.0709 | 2.2899 | 3.4677 | |
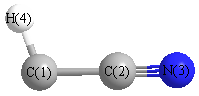 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
175.777 |
|
C2 |
C1 |
H4 |
145.165 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.291316 |
| Energy at 298.15K | |
| HF Energy | -131.169310 |
| Nuclear repulsion energy | 47.348414 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3488 |
3488 |
73.73 |
|
|
|
| 2 |
Σ |
1728 |
1728 |
29.97 |
|
|
|
| 3 |
Σ |
1277 |
1277 |
9.98 |
|
|
|
| 4 |
Π |
462 |
462 |
1.01 |
|
|
|
| 4 |
Π |
462 |
462 |
1.01 |
|
|
|
| 5 |
Π |
386i |
386i |
52.57 |
|
|
|
| 5 |
Π |
386i |
386i |
52.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3323.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3323.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.212 |
| C2 |
0.000 |
0.000 |
0.084 |
| N3 |
0.000 |
0.000 |
1.292 |
| H4 |
0.000 |
0.000 |
-2.276 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2960 | 2.5037 | 1.0637 |
C2 | 1.2960 | | 1.2077 | 2.3597 | N3 | 2.5037 | 1.2077 | | 3.5674 | H4 | 1.0637 | 2.3597 | 3.5674 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULL/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.054 |
|
|
|
| 2 |
C |
0.268 |
|
|
|
| 3 |
N |
-0.435 |
|
|
|
| 4 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
35.973 |
| (<r2>)1/2 |
5.998 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.271218 |
| Energy at 298.15K | -131.270653 |
| HF Energy | -131.132743 |
| Nuclear repulsion energy | 46.758418 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3111 |
3111 |
14.50 |
|
|
|
| 2 |
A' |
2099 |
2099 |
21.67 |
|
|
|
| 3 |
A' |
1086 |
1086 |
41.88 |
|
|
|
| 4 |
A' |
956 |
956 |
38.37 |
|
|
|
| 5 |
A' |
428 |
428 |
20.65 |
|
|
|
| 6 |
A" |
328 |
328 |
10.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4003.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4003.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.129 |
-1.284 |
0.000 |
| C2 |
0.000 |
0.092 |
0.000 |
| N3 |
-0.281 |
1.247 |
0.000 |
| H4 |
1.188 |
-1.574 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3817 | 2.5640 | 1.0979 |
C2 | 1.3817 | | 1.1889 | 2.0458 | N3 | 2.5640 | 1.1889 | | 3.1802 | H4 | 1.0979 | 2.0458 | 3.1802 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -131.271218 |
| Energy at 298.15K | -131.270653 |
| HF Energy | -131.132743 |
| Nuclear repulsion energy | 46.758418 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
