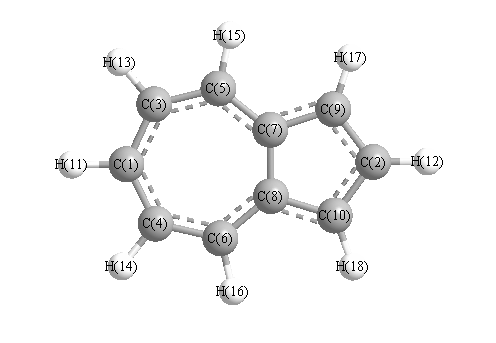
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3272 |
3119 |
12.01 |
|
|
|
| 2 |
A1 |
3245 |
3093 |
6.94 |
|
|
|
| 3 |
A1 |
3216 |
3065 |
22.98 |
|
|
|
| 4 |
A1 |
3188 |
3038 |
7.53 |
|
|
|
| 5 |
A1 |
3178 |
3029 |
0.08 |
|
|
|
| 6 |
A1 |
1683 |
1604 |
61.20 |
|
|
|
| 7 |
A1 |
1614 |
1539 |
8.02 |
|
|
|
| 8 |
A1 |
1519 |
1448 |
33.91 |
|
|
|
| 9 |
A1 |
1454 |
1386 |
76.71 |
|
|
|
| 10 |
A1 |
1328 |
1265 |
0.02 |
|
|
|
| 11 |
A1 |
1259 |
1200 |
0.31 |
|
|
|
| 12 |
A1 |
1099 |
1047 |
7.08 |
|
|
|
| 13 |
A1 |
981 |
935 |
1.42 |
|
|
|
| 14 |
A1 |
915 |
872 |
2.25 |
|
|
|
| 15 |
A1 |
836 |
796 |
5.12 |
|
|
|
| 16 |
A1 |
693 |
660 |
0.75 |
|
|
|
| 17 |
A1 |
412 |
392 |
1.28 |
|
|
|
| 18 |
A2 |
1002 |
955 |
0.00 |
|
|
|
| 19 |
A2 |
885 |
844 |
0.00 |
|
|
|
| 20 |
A2 |
799 |
762 |
0.00 |
|
|
|
| 21 |
A2 |
730 |
696 |
0.00 |
|
|
|
| 22 |
A2 |
432 |
412 |
0.00 |
|
|
|
| 23 |
A2 |
164 |
156 |
0.00 |
|
|
|
| 24 |
B1 |
1014 |
966 |
0.06 |
|
|
|
| 25 |
B1 |
985 |
939 |
6.57 |
|
|
|
| 26 |
B1 |
943 |
899 |
0.10 |
|
|
|
| 27 |
B1 |
789 |
752 |
108.61 |
|
|
|
| 28 |
B1 |
748 |
713 |
3.66 |
|
|
|
| 29 |
B1 |
615 |
586 |
1.82 |
|
|
|
| 30 |
B1 |
575 |
548 |
6.89 |
|
|
|
| 31 |
B1 |
320 |
305 |
8.00 |
|
|
|
| 32 |
B1 |
170 |
162 |
1.48 |
|
|
|
| 33 |
B2 |
3263 |
3110 |
11.36 |
|
|
|
| 34 |
B2 |
3208 |
3057 |
30.73 |
|
|
|
| 35 |
B2 |
3180 |
3031 |
13.44 |
|
|
|
| 36 |
B2 |
1683 |
1604 |
3.51 |
|
|
|
| 37 |
B2 |
1579 |
1505 |
9.51 |
|
|
|
| 38 |
B2 |
1510 |
1440 |
6.96 |
|
|
|
| 39 |
B2 |
1451 |
1383 |
1.00 |
|
|
|
| 40 |
B2 |
1352 |
1289 |
0.68 |
|
|
|
| 41 |
B2 |
1335 |
1272 |
3.82 |
|
|
|
| 42 |
B2 |
1257 |
1198 |
7.39 |
|
|
|
| 43 |
B2 |
1204 |
1148 |
0.34 |
|
|
|
| 44 |
B2 |
1078 |
1028 |
0.04 |
|
|
|
| 45 |
B2 |
1049 |
1000 |
13.80 |
|
|
|
| 46 |
B2 |
753 |
717 |
0.39 |
|
|
|
| 47 |
B2 |
499 |
476 |
1.76 |
|
|
|
| 48 |
B2 |
334 |
319 |
0.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32398.1 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 30878.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.142 |
|
|
|
| 2 |
C |
-0.184 |
|
|
|
| 3 |
C |
-0.187 |
|
|
|
| 4 |
C |
-0.187 |
|
|
|
| 5 |
C |
-0.190 |
|
|
|
| 6 |
C |
-0.190 |
|
|
|
| 7 |
C |
0.082 |
|
|
|
| 8 |
C |
0.082 |
|
|
|
| 9 |
C |
-0.225 |
|
|
|
| 10 |
C |
-0.225 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
H |
0.171 |
|
|
|
| 15 |
H |
0.178 |
|
|
|
| 16 |
H |
0.178 |
|
|
|
| 17 |
H |
0.163 |
|
|
|
| 18 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.113 |
1.113 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.762 |
0.000 |
0.000 |
| y |
0.000 |
-49.599 |
0.000 |
| z |
0.000 |
0.000 |
-50.412 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.757 |
0.000 |
0.000 |
| y |
0.000 |
6.988 |
0.000 |
| z |
0.000 |
0.000 |
5.769 |
|
| Polar |
|---|
| 3z2-r2 | 11.538 |
|---|
| x2-y2 | -13.163 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.917 |
0.000 |
0.000 |
| y |
0.000 |
17.135 |
0.000 |
| z |
0.000 |
0.000 |
25.817 |
<r2> (average value of r
2) Å
2
| <r2> |
364.324 |
| (<r2>)1/2 |
19.087 |