Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G**
Geometric Data calculated at B2PLYP/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -344.960447 |
| Energy at 298.15K | -344.977606 |
| HF Energy | -344.570709 |
| Nuclear repulsion energy | 424.535660 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3116 |
3116 |
0.00 |
|
|
|
| 2 |
A1 |
3087 |
3087 |
0.00 |
|
|
|
| 3 |
A1 |
1534 |
1534 |
0.00 |
|
|
|
| 4 |
A1 |
1385 |
1385 |
0.00 |
|
|
|
| 5 |
A1 |
1284 |
1284 |
0.00 |
|
|
|
| 6 |
A1 |
1035 |
1035 |
0.00 |
|
|
|
| 7 |
A1 |
984 |
984 |
0.00 |
|
|
|
| 8 |
A1 |
821 |
821 |
0.00 |
|
|
|
| 9 |
A1 |
612 |
612 |
0.00 |
|
|
|
| 10 |
A1 |
73 |
73 |
0.00 |
|
|
|
| 11 |
A2 |
3136 |
3136 |
0.00 |
|
|
|
| 12 |
A2 |
3074 |
3074 |
110.70 |
|
|
|
| 13 |
A2 |
1525 |
1525 |
2.36 |
|
|
|
| 14 |
A2 |
1406 |
1406 |
8.28 |
|
|
|
| 15 |
A2 |
1210 |
1210 |
0.00 |
|
|
|
| 16 |
A2 |
1004 |
1004 |
21.57 |
|
|
|
| 17 |
A2 |
820 |
820 |
0.00 |
|
|
|
| 18 |
A2 |
774 |
774 |
62.60 |
|
|
|
| 19 |
E |
3143 |
3143 |
88.93 |
|
|
|
| 19 |
E |
3143 |
3143 |
88.99 |
|
|
|
| 20 |
E |
3119 |
3119 |
0.00 |
|
|
|
| 20 |
E |
3119 |
3119 |
0.00 |
|
|
|
| 21 |
E |
3080 |
3080 |
103.93 |
|
|
|
| 21 |
E |
3080 |
3080 |
103.85 |
|
|
|
| 22 |
E |
3072 |
3072 |
0.00 |
|
|
|
| 22 |
E |
3072 |
3072 |
0.00 |
|
|
|
| 23 |
E |
1524 |
1524 |
6.43 |
|
|
|
| 23 |
E |
1524 |
1524 |
6.44 |
|
|
|
| 24 |
E |
1512 |
1512 |
0.00 |
|
|
|
| 24 |
E |
1512 |
1512 |
0.00 |
|
|
|
| 25 |
E |
1372 |
1372 |
8.26 |
|
|
|
| 25 |
E |
1372 |
1372 |
8.26 |
|
|
|
| 26 |
E |
1364 |
1364 |
0.00 |
|
|
|
| 26 |
E |
1364 |
1364 |
0.00 |
|
|
|
| 27 |
E |
1351 |
1351 |
0.04 |
|
|
|
| 27 |
E |
1351 |
1351 |
0.04 |
|
|
|
| 28 |
E |
1345 |
1345 |
0.00 |
|
|
|
| 28 |
E |
1345 |
1345 |
0.00 |
|
|
|
| 29 |
E |
1223 |
1223 |
0.00 |
|
|
|
| 29 |
E |
1223 |
1223 |
0.00 |
|
|
|
| 30 |
E |
1100 |
1100 |
39.30 |
|
|
|
| 30 |
E |
1100 |
1100 |
39.30 |
|
|
|
| 31 |
E |
1056 |
1056 |
0.00 |
|
|
|
| 31 |
E |
1056 |
1056 |
0.00 |
|
|
|
| 32 |
E |
913 |
913 |
4.34 |
|
|
|
| 32 |
E |
913 |
913 |
4.34 |
|
|
|
| 33 |
E |
842 |
842 |
4.00 |
|
|
|
| 33 |
E |
842 |
842 |
4.00 |
|
|
|
| 34 |
E |
590 |
590 |
0.00 |
|
|
|
| 34 |
E |
590 |
590 |
0.00 |
|
|
|
| 35 |
E |
428 |
428 |
0.00 |
|
|
|
| 35 |
E |
428 |
428 |
0.00 |
|
|
|
| 36 |
E |
335 |
335 |
0.00 |
|
|
|
| 36 |
E |
335 |
335 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40810.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 40810.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.289 |
| N2 |
0.000 |
0.000 |
-1.289 |
| C3 |
-0.001 |
1.381 |
0.780 |
| C4 |
1.196 |
-0.690 |
0.780 |
| C5 |
-1.195 |
-0.691 |
0.780 |
| C6 |
0.001 |
1.381 |
-0.780 |
| C7 |
-1.196 |
-0.690 |
-0.780 |
| C8 |
1.195 |
-0.691 |
-0.780 |
| H9 |
0.880 |
1.890 |
1.182 |
| H10 |
-0.884 |
1.887 |
1.180 |
| H11 |
1.197 |
-1.706 |
1.182 |
| H12 |
2.076 |
-0.178 |
1.180 |
| H13 |
-2.076 |
-0.183 |
1.182 |
| H14 |
-1.192 |
-1.709 |
1.180 |
| H15 |
-0.880 |
1.890 |
-1.182 |
| H16 |
0.884 |
1.887 |
-1.180 |
| H17 |
-1.197 |
-1.706 |
-1.182 |
| H18 |
-2.076 |
-0.178 |
-1.180 |
| H19 |
2.076 |
-0.183 |
-1.182 |
| H20 |
1.192 |
-1.709 |
-1.180 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5787 | 1.4718 | 1.4718 | 1.4718 | 2.4879 | 2.4879 | 2.4879 | 2.0870 | 2.0867 | 2.0870 | 2.0867 | 2.0870 | 2.0867 | 3.2329 | 3.2308 | 3.2329 | 3.2308 | 3.2329 | 3.2308 |
N2 | 2.5787 | | 2.4879 | 2.4879 | 2.4879 | 1.4718 | 1.4718 | 1.4718 | 3.2329 | 3.2308 | 3.2329 | 3.2308 | 3.2329 | 3.2308 | 2.0870 | 2.0867 | 2.0870 | 2.0867 | 2.0870 | 2.0867 | C3 | 1.4718 | 2.4879 | | 2.3917 | 2.3917 | 1.5603 | 2.8548 | 2.8565 | 1.0934 | 1.0934 | 3.3358 | 2.6276 | 2.6295 | 3.3357 | 2.2092 | 2.2091 | 3.8485 | 3.2522 | 3.2574 | 3.8487 | C4 | 1.4718 | 2.4879 | 2.3917 | | 2.3917 | 2.8548 | 2.8565 | 1.5603 | 2.6295 | 3.3357 | 1.0934 | 1.0934 | 3.3358 | 2.6276 | 3.8485 | 3.2522 | 3.2574 | 3.8487 | 2.2092 | 2.2091 | C5 | 1.4718 | 2.4879 | 2.3917 | 2.3917 | | 2.8565 | 1.5603 | 2.8548 | 3.3358 | 2.6276 | 2.6295 | 3.3357 | 1.0934 | 1.0934 | 3.2574 | 3.8487 | 2.2092 | 2.2091 | 3.8485 | 3.2522 | C6 | 2.4879 | 1.4718 | 1.5603 | 2.8548 | 2.8565 | | 2.3917 | 2.3917 | 2.2092 | 2.2091 | 3.8485 | 3.2522 | 3.2574 | 3.8487 | 1.0934 | 1.0934 | 3.3358 | 2.6276 | 2.6295 | 3.3357 | C7 | 2.4879 | 1.4718 | 2.8548 | 2.8565 | 1.5603 | 2.3917 | | 2.3917 | 3.8485 | 3.2522 | 3.2574 | 3.8487 | 2.2092 | 2.2091 | 2.6295 | 3.3357 | 1.0934 | 1.0934 | 3.3358 | 2.6276 | C8 | 2.4879 | 1.4718 | 2.8565 | 1.5603 | 2.8548 | 2.3917 | 2.3917 | | 3.2574 | 3.8487 | 2.2092 | 2.2091 | 3.8485 | 3.2522 | 3.3358 | 2.6276 | 2.6295 | 3.3357 | 1.0934 | 1.0934 | H9 | 2.0870 | 3.2329 | 1.0934 | 2.6295 | 3.3358 | 2.2092 | 3.8485 | 3.2574 | | 1.7636 | 3.6100 | 2.3889 | 3.6100 | 4.1524 | 2.9466 | 2.3616 | 4.7782 | 4.3114 | 3.3640 | 4.3158 | H10 | 2.0867 | 3.2308 | 1.0934 | 3.3357 | 2.6276 | 2.2091 | 3.2522 | 3.8487 | 1.7636 | | 4.1524 | 3.6093 | 2.3889 | 3.6093 | 2.3616 | 2.9483 | 4.3114 | 3.3543 | 4.3158 | 4.7759 | H11 | 2.0870 | 3.2329 | 3.3358 | 1.0934 | 2.6295 | 3.8485 | 3.2574 | 2.2092 | 3.6100 | 4.1524 | | 1.7636 | 3.6100 | 2.3889 | 4.7782 | 4.3114 | 3.3640 | 4.3158 | 2.9466 | 2.3616 | H12 | 2.0867 | 3.2308 | 2.6276 | 1.0934 | 3.3357 | 3.2522 | 3.8487 | 2.2091 | 2.3889 | 3.6093 | 1.7636 | | 4.1524 | 3.6093 | 4.3114 | 3.3543 | 4.3158 | 4.7759 | 2.3616 | 2.9483 | H13 | 2.0870 | 3.2329 | 2.6295 | 3.3358 | 1.0934 | 3.2574 | 2.2092 | 3.8485 | 3.6100 | 2.3889 | 3.6100 | 4.1524 | | 1.7636 | 3.3640 | 4.3158 | 2.9466 | 2.3616 | 4.7782 | 4.3114 | H14 | 2.0867 | 3.2308 | 3.3357 | 2.6276 | 1.0934 | 3.8487 | 2.2091 | 3.2522 | 4.1524 | 3.6093 | 2.3889 | 3.6093 | 1.7636 | | 4.3158 | 4.7759 | 2.3616 | 2.9483 | 4.3114 | 3.3543 | H15 | 3.2329 | 2.0870 | 2.2092 | 3.8485 | 3.2574 | 1.0934 | 2.6295 | 3.3358 | 2.9466 | 2.3616 | 4.7782 | 4.3114 | 3.3640 | 4.3158 | | 1.7636 | 3.6100 | 2.3889 | 3.6100 | 4.1524 | H16 | 3.2308 | 2.0867 | 2.2091 | 3.2522 | 3.8487 | 1.0934 | 3.3357 | 2.6276 | 2.3616 | 2.9483 | 4.3114 | 3.3543 | 4.3158 | 4.7759 | 1.7636 | | 4.1524 | 3.6093 | 2.3889 | 3.6093 | H17 | 3.2329 | 2.0870 | 3.8485 | 3.2574 | 2.2092 | 3.3358 | 1.0934 | 2.6295 | 4.7782 | 4.3114 | 3.3640 | 4.3158 | 2.9466 | 2.3616 | 3.6100 | 4.1524 | | 1.7636 | 3.6100 | 2.3889 | H18 | 3.2308 | 2.0867 | 3.2522 | 3.8487 | 2.2091 | 2.6276 | 1.0934 | 3.3357 | 4.3114 | 3.3543 | 4.3158 | 4.7759 | 2.3616 | 2.9483 | 2.3889 | 3.6093 | 1.7636 | | 4.1524 | 3.6093 | H19 | 3.2329 | 2.0870 | 3.2574 | 2.2092 | 3.8485 | 2.6295 | 3.3358 | 1.0934 | 3.3640 | 4.3158 | 2.9466 | 2.3616 | 4.7782 | 4.3114 | 3.6100 | 2.3889 | 3.6100 | 4.1524 | | 1.7636 | H20 | 3.2308 | 2.0867 | 3.8487 | 2.2091 | 3.2522 | 3.3357 | 2.6276 | 1.0934 | 4.3158 | 4.7759 | 2.3616 | 2.9483 | 4.3114 | 3.3543 | 4.1524 | 3.6093 | 2.3889 | 3.6093 | 1.7636 | |
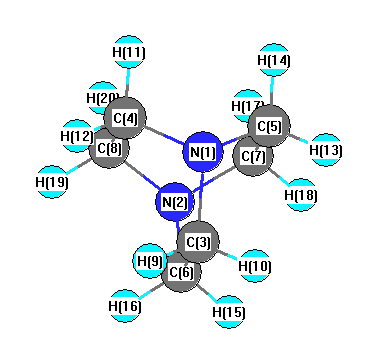 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.243 |
|
N1 |
C3 |
H9 |
107.988 |
| N1 |
C3 |
H10 |
107.965 |
|
N1 |
C4 |
C8 |
110.243 |
| N1 |
C4 |
H11 |
107.989 |
|
N1 |
C4 |
H12 |
107.965 |
| N1 |
C5 |
C7 |
110.243 |
|
N1 |
C5 |
H13 |
107.989 |
| N1 |
C5 |
H14 |
107.965 |
|
N2 |
C6 |
C3 |
110.243 |
| N2 |
C6 |
H15 |
107.988 |
|
N2 |
C6 |
H16 |
107.965 |
| N2 |
C7 |
C5 |
110.243 |
|
N2 |
C7 |
H17 |
107.989 |
| N2 |
C7 |
H18 |
107.965 |
|
N2 |
C8 |
C4 |
110.243 |
| N2 |
C8 |
H19 |
107.989 |
|
N2 |
C8 |
H20 |
107.965 |
| C3 |
N1 |
C4 |
108.689 |
|
C3 |
N1 |
C5 |
108.689 |
| C3 |
C6 |
H15 |
111.500 |
|
C3 |
C6 |
H16 |
111.493 |
| C4 |
N1 |
C5 |
108.689 |
|
C4 |
C8 |
H19 |
111.500 |
| C4 |
C8 |
H20 |
111.493 |
|
C5 |
C6 |
H15 |
101.595 |
| C5 |
C6 |
H16 |
150.881 |
|
C6 |
N2 |
C7 |
108.689 |
| C6 |
N2 |
C8 |
108.689 |
|
C6 |
C3 |
H9 |
111.500 |
| C6 |
C3 |
H10 |
111.493 |
|
C7 |
N2 |
C8 |
108.689 |
| C7 |
C5 |
H13 |
111.500 |
|
C7 |
C5 |
H14 |
111.493 |
| C8 |
C4 |
H11 |
111.500 |
|
C8 |
C4 |
H12 |
111.493 |
| H9 |
C3 |
H10 |
107.499 |
|
H11 |
C4 |
H12 |
107.499 |
| H13 |
C5 |
H14 |
107.499 |
|
H15 |
C6 |
H16 |
107.499 |
| H17 |
C7 |
H18 |
107.499 |
|
H19 |
C8 |
H20 |
107.499 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.422 |
|
|
|
| 2 |
N |
-0.422 |
|
|
|
| 3 |
C |
-0.071 |
|
|
|
| 4 |
C |
-0.071 |
|
|
|
| 5 |
C |
-0.071 |
|
|
|
| 6 |
C |
-0.071 |
|
|
|
| 7 |
C |
-0.071 |
|
|
|
| 8 |
C |
-0.071 |
|
|
|
| 9 |
H |
0.106 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
| 11 |
H |
0.106 |
|
|
|
| 12 |
H |
0.106 |
|
|
|
| 13 |
H |
0.106 |
|
|
|
| 14 |
H |
0.106 |
|
|
|
| 15 |
H |
0.106 |
|
|
|
| 16 |
H |
0.106 |
|
|
|
| 17 |
H |
0.106 |
|
|
|
| 18 |
H |
0.106 |
|
|
|
| 19 |
H |
0.106 |
|
|
|
| 20 |
H |
0.106 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.904 |
0.000 |
0.000 |
| y |
0.000 |
-46.904 |
0.000 |
| z |
0.000 |
0.000 |
-57.830 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.463 |
0.000 |
0.000 |
| y |
0.000 |
5.463 |
0.000 |
| z |
0.000 |
0.000 |
-10.927 |
|
| Polar |
|---|
| 3z2-r2 | -21.853 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.872 |
0.000 |
0.000 |
| y |
0.000 |
10.872 |
0.000 |
| z |
0.000 |
0.000 |
9.322 |
<r2> (average value of r
2) Å
2
| <r2> |
214.249 |
| (<r2>)1/2 |
14.637 |
