Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -344.969884 |
| Energy at 298.15K | -344.987066 |
| HF Energy | -344.969884 |
| Nuclear repulsion energy | 426.003647 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3100 |
2958 |
0.00 |
|
|
|
| 2 |
A1 |
3069 |
2928 |
0.00 |
|
|
|
| 3 |
A1 |
1507 |
1438 |
0.00 |
|
|
|
| 4 |
A1 |
1374 |
1311 |
0.00 |
|
|
|
| 5 |
A1 |
1269 |
1211 |
0.00 |
|
|
|
| 6 |
A1 |
1028 |
981 |
0.00 |
|
|
|
| 7 |
A1 |
993 |
947 |
0.00 |
|
|
|
| 8 |
A1 |
837 |
799 |
0.00 |
|
|
|
| 9 |
A1 |
613 |
585 |
0.00 |
|
|
|
| 10 |
A1 |
80 |
77 |
0.00 |
|
|
|
| 11 |
A2 |
3119 |
2977 |
0.00 |
|
|
|
| 12 |
A2 |
3056 |
2916 |
112.73 |
|
|
|
| 13 |
A2 |
1498 |
1429 |
5.36 |
|
|
|
| 14 |
A2 |
1390 |
1327 |
5.46 |
|
|
|
| 15 |
A2 |
1203 |
1148 |
0.00 |
|
|
|
| 16 |
A2 |
1009 |
963 |
19.92 |
|
|
|
| 17 |
A2 |
811 |
774 |
0.01 |
|
|
|
| 18 |
A2 |
790 |
754 |
65.97 |
|
|
|
| 19 |
E |
3125 |
2983 |
83.04 |
|
|
|
| 19 |
E |
3125 |
2983 |
83.10 |
|
|
|
| 20 |
E |
3102 |
2960 |
0.01 |
|
|
|
| 20 |
E |
3102 |
2960 |
0.01 |
|
|
|
| 21 |
E |
3062 |
2922 |
107.52 |
|
|
|
| 21 |
E |
3062 |
2922 |
107.53 |
|
|
|
| 22 |
E |
3055 |
2915 |
0.00 |
|
|
|
| 22 |
E |
3055 |
2915 |
0.00 |
|
|
|
| 23 |
E |
1497 |
1428 |
9.87 |
|
|
|
| 23 |
E |
1497 |
1428 |
9.88 |
|
|
|
| 24 |
E |
1484 |
1416 |
0.00 |
|
|
|
| 24 |
E |
1484 |
1416 |
0.00 |
|
|
|
| 25 |
E |
1370 |
1307 |
10.88 |
|
|
|
| 25 |
E |
1370 |
1307 |
10.89 |
|
|
|
| 26 |
E |
1360 |
1298 |
0.00 |
|
|
|
| 26 |
E |
1360 |
1298 |
0.00 |
|
|
|
| 27 |
E |
1338 |
1277 |
0.58 |
|
|
|
| 27 |
E |
1338 |
1277 |
0.58 |
|
|
|
| 28 |
E |
1332 |
1271 |
0.00 |
|
|
|
| 28 |
E |
1332 |
1271 |
0.00 |
|
|
|
| 29 |
E |
1215 |
1160 |
0.00 |
|
|
|
| 29 |
E |
1215 |
1160 |
0.00 |
|
|
|
| 30 |
E |
1117 |
1066 |
39.32 |
|
|
|
| 30 |
E |
1117 |
1066 |
39.34 |
|
|
|
| 31 |
E |
1067 |
1018 |
0.00 |
|
|
|
| 31 |
E |
1067 |
1018 |
0.00 |
|
|
|
| 32 |
E |
921 |
879 |
3.68 |
|
|
|
| 32 |
E |
921 |
879 |
3.68 |
|
|
|
| 33 |
E |
833 |
795 |
4.15 |
|
|
|
| 33 |
E |
833 |
795 |
4.15 |
|
|
|
| 34 |
E |
592 |
565 |
0.00 |
|
|
|
| 34 |
E |
592 |
565 |
0.00 |
|
|
|
| 35 |
E |
427 |
407 |
0.00 |
|
|
|
| 35 |
E |
427 |
407 |
0.00 |
|
|
|
| 36 |
E |
339 |
323 |
0.00 |
|
|
|
| 36 |
E |
339 |
323 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40606.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 38750.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.281 |
| N2 |
0.000 |
0.000 |
-1.281 |
| C3 |
-0.000 |
1.374 |
0.778 |
| C4 |
1.190 |
-0.687 |
0.778 |
| C5 |
-1.190 |
-0.687 |
0.778 |
| C6 |
0.000 |
1.374 |
-0.778 |
| C7 |
-1.190 |
-0.687 |
-0.778 |
| C8 |
1.190 |
-0.687 |
-0.778 |
| H9 |
0.882 |
1.885 |
1.182 |
| H10 |
-0.884 |
1.884 |
1.181 |
| H11 |
1.191 |
-1.706 |
1.182 |
| H12 |
2.073 |
-0.176 |
1.181 |
| H13 |
-2.073 |
-0.179 |
1.182 |
| H14 |
-1.189 |
-1.707 |
1.181 |
| H15 |
-0.882 |
1.885 |
-1.182 |
| H16 |
0.884 |
1.884 |
-1.181 |
| H17 |
-1.191 |
-1.706 |
-1.182 |
| H18 |
-2.073 |
-0.176 |
-1.181 |
| H19 |
2.073 |
-0.179 |
-1.182 |
| H20 |
1.189 |
-1.707 |
-1.181 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5622 | 1.4633 | 1.4633 | 1.4633 | 2.4756 | 2.4756 | 2.4756 | 2.0833 | 2.0830 | 2.0833 | 2.0830 | 2.0833 | 2.0830 | 3.2241 | 3.2232 | 3.2241 | 3.2232 | 3.2241 | 3.2232 |
N2 | 2.5622 | | 2.4756 | 2.4756 | 2.4756 | 1.4633 | 1.4633 | 1.4633 | 3.2241 | 3.2232 | 3.2241 | 3.2232 | 3.2241 | 3.2232 | 2.0833 | 2.0830 | 2.0833 | 2.0830 | 2.0833 | 2.0830 | C3 | 1.4633 | 2.4756 | | 2.3800 | 2.3800 | 1.5563 | 2.8434 | 2.8440 | 1.0963 | 1.0963 | 3.3273 | 2.6202 | 2.6213 | 3.3271 | 2.2087 | 2.2087 | 3.8402 | 3.2462 | 3.2483 | 3.8401 | C4 | 1.4633 | 2.4756 | 2.3800 | | 2.3800 | 2.8434 | 2.8440 | 1.5563 | 2.6213 | 3.3271 | 1.0963 | 1.0963 | 3.3273 | 2.6202 | 3.8402 | 3.2462 | 3.2483 | 3.8401 | 2.2087 | 2.2087 | C5 | 1.4633 | 2.4756 | 2.3800 | 2.3800 | | 2.8440 | 1.5563 | 2.8434 | 3.3273 | 2.6202 | 2.6213 | 3.3271 | 1.0963 | 1.0963 | 3.2483 | 3.8401 | 2.2087 | 2.2087 | 3.8402 | 3.2462 | C6 | 2.4756 | 1.4633 | 1.5563 | 2.8434 | 2.8440 | | 2.3800 | 2.3800 | 2.2087 | 2.2087 | 3.8402 | 3.2462 | 3.2483 | 3.8401 | 1.0963 | 1.0963 | 3.3273 | 2.6202 | 2.6213 | 3.3271 | C7 | 2.4756 | 1.4633 | 2.8434 | 2.8440 | 1.5563 | 2.3800 | | 2.3800 | 3.8402 | 3.2462 | 3.2483 | 3.8401 | 2.2087 | 2.2087 | 2.6213 | 3.3271 | 1.0963 | 1.0963 | 3.3273 | 2.6202 | C8 | 2.4756 | 1.4633 | 2.8440 | 1.5563 | 2.8434 | 2.3800 | 2.3800 | | 3.2483 | 3.8401 | 2.2087 | 2.2087 | 3.8402 | 3.2462 | 3.3273 | 2.6202 | 2.6213 | 3.3271 | 1.0963 | 1.0963 | H9 | 2.0833 | 3.2241 | 1.0963 | 2.6213 | 3.3273 | 2.2087 | 3.8402 | 3.2483 | | 1.7655 | 3.6043 | 2.3808 | 3.6043 | 4.1463 | 2.9487 | 2.3622 | 4.7726 | 4.3082 | 3.3559 | 4.3101 | H10 | 2.0830 | 3.2232 | 1.0963 | 3.3271 | 2.6202 | 2.2087 | 3.2462 | 3.8401 | 1.7655 | | 4.1463 | 3.6037 | 2.3808 | 3.6037 | 2.3622 | 2.9495 | 4.3082 | 3.3518 | 4.3101 | 4.7715 | H11 | 2.0833 | 3.2241 | 3.3273 | 1.0963 | 2.6213 | 3.8402 | 3.2483 | 2.2087 | 3.6043 | 4.1463 | | 1.7655 | 3.6043 | 2.3808 | 4.7726 | 4.3082 | 3.3559 | 4.3101 | 2.9487 | 2.3622 | H12 | 2.0830 | 3.2232 | 2.6202 | 1.0963 | 3.3271 | 3.2462 | 3.8401 | 2.2087 | 2.3808 | 3.6037 | 1.7655 | | 4.1463 | 3.6037 | 4.3082 | 3.3518 | 4.3101 | 4.7715 | 2.3622 | 2.9495 | H13 | 2.0833 | 3.2241 | 2.6213 | 3.3273 | 1.0963 | 3.2483 | 2.2087 | 3.8402 | 3.6043 | 2.3808 | 3.6043 | 4.1463 | | 1.7655 | 3.3559 | 4.3101 | 2.9487 | 2.3622 | 4.7726 | 4.3082 | H14 | 2.0830 | 3.2232 | 3.3271 | 2.6202 | 1.0963 | 3.8401 | 2.2087 | 3.2462 | 4.1463 | 3.6037 | 2.3808 | 3.6037 | 1.7655 | | 4.3101 | 4.7715 | 2.3622 | 2.9495 | 4.3082 | 3.3518 | H15 | 3.2241 | 2.0833 | 2.2087 | 3.8402 | 3.2483 | 1.0963 | 2.6213 | 3.3273 | 2.9487 | 2.3622 | 4.7726 | 4.3082 | 3.3559 | 4.3101 | | 1.7655 | 3.6043 | 2.3808 | 3.6043 | 4.1463 | H16 | 3.2232 | 2.0830 | 2.2087 | 3.2462 | 3.8401 | 1.0963 | 3.3271 | 2.6202 | 2.3622 | 2.9495 | 4.3082 | 3.3518 | 4.3101 | 4.7715 | 1.7655 | | 4.1463 | 3.6037 | 2.3808 | 3.6037 | H17 | 3.2241 | 2.0833 | 3.8402 | 3.2483 | 2.2087 | 3.3273 | 1.0963 | 2.6213 | 4.7726 | 4.3082 | 3.3559 | 4.3101 | 2.9487 | 2.3622 | 3.6043 | 4.1463 | | 1.7655 | 3.6043 | 2.3808 | H18 | 3.2232 | 2.0830 | 3.2462 | 3.8401 | 2.2087 | 2.6202 | 1.0963 | 3.3271 | 4.3082 | 3.3518 | 4.3101 | 4.7715 | 2.3622 | 2.9495 | 2.3808 | 3.6037 | 1.7655 | | 4.1463 | 3.6037 | H19 | 3.2241 | 2.0833 | 3.2483 | 2.2087 | 3.8402 | 2.6213 | 3.3273 | 1.0963 | 3.3559 | 4.3101 | 2.9487 | 2.3622 | 4.7726 | 4.3082 | 3.6043 | 2.3808 | 3.6043 | 4.1463 | | 1.7655 | H20 | 3.2232 | 2.0830 | 3.8401 | 2.2087 | 3.2462 | 3.3271 | 2.6202 | 1.0963 | 4.3101 | 4.7715 | 2.3622 | 2.9495 | 4.3082 | 3.3518 | 4.1463 | 3.6037 | 2.3808 | 3.6037 | 1.7655 | |
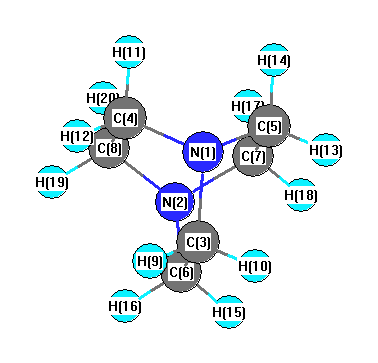 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.103 |
|
N1 |
C3 |
H9 |
108.109 |
| N1 |
C3 |
H10 |
108.084 |
|
N1 |
C4 |
C8 |
110.103 |
| N1 |
C4 |
H11 |
108.109 |
|
N1 |
C4 |
H12 |
108.084 |
| N1 |
C5 |
C7 |
110.103 |
|
N1 |
C5 |
H13 |
108.109 |
| N1 |
C5 |
H14 |
108.084 |
|
N2 |
C6 |
C3 |
110.103 |
| N2 |
C6 |
H15 |
108.109 |
|
N2 |
C6 |
H16 |
108.084 |
| N2 |
C7 |
C5 |
110.103 |
|
N2 |
C7 |
H17 |
108.109 |
| N2 |
C7 |
H18 |
108.084 |
|
N2 |
C8 |
C4 |
110.103 |
| N2 |
C8 |
H19 |
108.109 |
|
N2 |
C8 |
H20 |
108.084 |
| C3 |
N1 |
C4 |
108.832 |
|
C3 |
N1 |
C5 |
108.832 |
| C3 |
C6 |
H15 |
111.568 |
|
C3 |
C6 |
H16 |
111.562 |
| C4 |
N1 |
C5 |
108.832 |
|
C4 |
C8 |
H19 |
111.568 |
| C4 |
C8 |
H20 |
111.562 |
|
C5 |
C6 |
H15 |
101.671 |
| C5 |
C6 |
H16 |
151.041 |
|
C6 |
N2 |
C7 |
108.832 |
| C6 |
N2 |
C8 |
108.832 |
|
C6 |
C3 |
H9 |
111.568 |
| C6 |
C3 |
H10 |
111.562 |
|
C7 |
N2 |
C8 |
108.832 |
| C7 |
C5 |
H13 |
111.568 |
|
C7 |
C5 |
H14 |
111.562 |
| C8 |
C4 |
H11 |
111.568 |
|
C8 |
C4 |
H12 |
111.562 |
| H9 |
C3 |
H10 |
107.266 |
|
H11 |
C4 |
H12 |
107.266 |
| H13 |
C5 |
H14 |
107.266 |
|
H15 |
C6 |
H16 |
107.266 |
| H17 |
C7 |
H18 |
107.266 |
|
H19 |
C8 |
H20 |
107.266 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.409 |
|
|
|
| 2 |
N |
-0.409 |
|
|
|
| 3 |
C |
-0.120 |
|
|
|
| 4 |
C |
-0.120 |
|
|
|
| 5 |
C |
-0.120 |
|
|
|
| 6 |
C |
-0.120 |
|
|
|
| 7 |
C |
-0.120 |
|
|
|
| 8 |
C |
-0.120 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.128 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.128 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.128 |
|
|
|
| 18 |
H |
0.128 |
|
|
|
| 19 |
H |
0.128 |
|
|
|
| 20 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.365 |
0.000 |
0.000 |
| y |
0.000 |
-46.365 |
0.000 |
| z |
0.000 |
0.000 |
-57.168 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.402 |
0.000 |
0.000 |
| y |
0.000 |
5.402 |
0.000 |
| z |
0.000 |
0.000 |
-10.803 |
|
| Polar |
|---|
| 3z2-r2 | -21.607 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.978 |
0.000 |
0.000 |
| y |
0.000 |
10.978 |
0.000 |
| z |
0.000 |
0.000 |
9.436 |
<r2> (average value of r
2) Å
2
| <r2> |
212.655 |
| (<r2>)1/2 |
14.583 |
