Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
Geometric Data calculated at B2PLYP=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -344.972028 |
| Energy at 298.15K | -344.989194 |
| HF Energy | -344.570761 |
| Nuclear repulsion energy | 424.693710 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3117 |
3117 |
0.00 |
|
|
|
| 2 |
A1 |
3088 |
3088 |
0.00 |
|
|
|
| 3 |
A1 |
1535 |
1535 |
0.00 |
|
|
|
| 4 |
A1 |
1386 |
1386 |
0.00 |
|
|
|
| 5 |
A1 |
1285 |
1285 |
0.00 |
|
|
|
| 6 |
A1 |
1036 |
1036 |
0.00 |
|
|
|
| 7 |
A1 |
984 |
984 |
0.00 |
|
|
|
| 8 |
A1 |
821 |
821 |
0.00 |
|
|
|
| 9 |
A1 |
612 |
612 |
0.00 |
|
|
|
| 10 |
A1 |
73 |
73 |
0.00 |
|
|
|
| 11 |
A2 |
3137 |
3137 |
0.00 |
|
|
|
| 12 |
A2 |
3075 |
3075 |
110.46 |
|
|
|
| 13 |
A2 |
1526 |
1526 |
2.42 |
|
|
|
| 14 |
A2 |
1407 |
1407 |
8.18 |
|
|
|
| 15 |
A2 |
1212 |
1212 |
0.00 |
|
|
|
| 16 |
A2 |
1005 |
1005 |
21.56 |
|
|
|
| 17 |
A2 |
821 |
821 |
0.00 |
|
|
|
| 18 |
A2 |
775 |
775 |
62.68 |
|
|
|
| 19 |
E |
3144 |
3144 |
88.69 |
|
|
|
| 19 |
E |
3144 |
3144 |
88.74 |
|
|
|
| 20 |
E |
3120 |
3120 |
0.00 |
|
|
|
| 20 |
E |
3120 |
3120 |
0.00 |
|
|
|
| 21 |
E |
3081 |
3081 |
103.88 |
|
|
|
| 21 |
E |
3081 |
3081 |
103.80 |
|
|
|
| 22 |
E |
3073 |
3073 |
0.00 |
|
|
|
| 22 |
E |
3073 |
3073 |
0.00 |
|
|
|
| 23 |
E |
1525 |
1525 |
6.51 |
|
|
|
| 23 |
E |
1525 |
1525 |
6.52 |
|
|
|
| 24 |
E |
1513 |
1513 |
0.00 |
|
|
|
| 24 |
E |
1513 |
1513 |
0.00 |
|
|
|
| 25 |
E |
1373 |
1373 |
8.30 |
|
|
|
| 25 |
E |
1373 |
1373 |
8.29 |
|
|
|
| 26 |
E |
1365 |
1365 |
0.00 |
|
|
|
| 26 |
E |
1365 |
1365 |
0.00 |
|
|
|
| 27 |
E |
1352 |
1352 |
0.04 |
|
|
|
| 27 |
E |
1352 |
1352 |
0.04 |
|
|
|
| 28 |
E |
1346 |
1346 |
0.00 |
|
|
|
| 28 |
E |
1346 |
1346 |
0.00 |
|
|
|
| 29 |
E |
1224 |
1224 |
0.00 |
|
|
|
| 29 |
E |
1224 |
1224 |
0.00 |
|
|
|
| 30 |
E |
1101 |
1101 |
39.37 |
|
|
|
| 30 |
E |
1101 |
1101 |
39.37 |
|
|
|
| 31 |
E |
1057 |
1057 |
0.00 |
|
|
|
| 31 |
E |
1057 |
1057 |
0.00 |
|
|
|
| 32 |
E |
914 |
914 |
4.32 |
|
|
|
| 32 |
E |
914 |
914 |
4.32 |
|
|
|
| 33 |
E |
842 |
842 |
4.00 |
|
|
|
| 33 |
E |
842 |
842 |
4.00 |
|
|
|
| 34 |
E |
591 |
591 |
0.00 |
|
|
|
| 34 |
E |
591 |
591 |
0.00 |
|
|
|
| 35 |
E |
429 |
429 |
0.00 |
|
|
|
| 35 |
E |
429 |
429 |
0.00 |
|
|
|
| 36 |
E |
336 |
336 |
0.00 |
|
|
|
| 36 |
E |
336 |
336 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40830.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 40830.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-31G**
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.289 |
| N2 |
0.000 |
0.000 |
-1.289 |
| C3 |
-0.001 |
1.380 |
0.780 |
| C4 |
1.196 |
-0.690 |
0.780 |
| C5 |
-1.195 |
-0.691 |
0.780 |
| C6 |
0.001 |
1.380 |
-0.780 |
| C7 |
-1.196 |
-0.690 |
-0.780 |
| C8 |
1.195 |
-0.691 |
-0.780 |
| H9 |
0.880 |
1.889 |
1.181 |
| H10 |
-0.883 |
1.887 |
1.179 |
| H11 |
1.196 |
-1.706 |
1.181 |
| H12 |
2.076 |
-0.178 |
1.179 |
| H13 |
-2.076 |
-0.183 |
1.181 |
| H14 |
-1.192 |
-1.708 |
1.179 |
| H15 |
-0.880 |
1.889 |
-1.181 |
| H16 |
0.883 |
1.887 |
-1.179 |
| H17 |
-1.196 |
-1.706 |
-1.181 |
| H18 |
-2.076 |
-0.178 |
-1.179 |
| H19 |
2.076 |
-0.183 |
-1.181 |
| H20 |
1.192 |
-1.708 |
-1.179 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5773 | 1.4712 | 1.4712 | 1.4712 | 2.4868 | 2.4868 | 2.4868 | 2.0863 | 2.0861 | 2.0863 | 2.0861 | 2.0863 | 2.0861 | 3.2314 | 3.2297 | 3.2314 | 3.2297 | 3.2314 | 3.2297 |
N2 | 2.5773 | | 2.4868 | 2.4868 | 2.4868 | 1.4712 | 1.4712 | 1.4712 | 3.2314 | 3.2297 | 3.2314 | 3.2297 | 3.2314 | 3.2297 | 2.0863 | 2.0861 | 2.0863 | 2.0861 | 2.0863 | 2.0861 | C3 | 1.4712 | 2.4868 | | 2.3909 | 2.3909 | 1.5597 | 2.8540 | 2.8553 | 1.0931 | 1.0931 | 3.3347 | 2.6269 | 2.6284 | 3.3346 | 2.2084 | 2.2084 | 3.8473 | 3.2516 | 3.2558 | 3.8474 | C4 | 1.4712 | 2.4868 | 2.3909 | | 2.3909 | 2.8540 | 2.8553 | 1.5597 | 2.6284 | 3.3346 | 1.0931 | 1.0931 | 3.3347 | 2.6269 | 3.8473 | 3.2516 | 3.2558 | 3.8474 | 2.2084 | 2.2084 | C5 | 1.4712 | 2.4868 | 2.3909 | 2.3909 | | 2.8553 | 1.5597 | 2.8540 | 3.3347 | 2.6269 | 2.6284 | 3.3346 | 1.0931 | 1.0931 | 3.2558 | 3.8474 | 2.2084 | 2.2084 | 3.8473 | 3.2516 | C6 | 2.4868 | 1.4712 | 1.5597 | 2.8540 | 2.8553 | | 2.3909 | 2.3909 | 2.2084 | 2.2084 | 3.8473 | 3.2516 | 3.2558 | 3.8474 | 1.0931 | 1.0931 | 3.3347 | 2.6269 | 2.6284 | 3.3346 | C7 | 2.4868 | 1.4712 | 2.8540 | 2.8553 | 1.5597 | 2.3909 | | 2.3909 | 3.8473 | 3.2516 | 3.2558 | 3.8474 | 2.2084 | 2.2084 | 2.6284 | 3.3346 | 1.0931 | 1.0931 | 3.3347 | 2.6269 | C8 | 2.4868 | 1.4712 | 2.8553 | 1.5597 | 2.8540 | 2.3909 | 2.3909 | | 3.2558 | 3.8474 | 2.2084 | 2.2084 | 3.8473 | 3.2516 | 3.3347 | 2.6269 | 2.6284 | 3.3346 | 1.0931 | 1.0931 | H9 | 2.0863 | 3.2314 | 1.0931 | 2.6284 | 3.3347 | 2.2084 | 3.8473 | 3.2558 | | 1.7631 | 3.6089 | 2.3881 | 3.6089 | 4.1511 | 2.9458 | 2.3607 | 4.7764 | 4.3104 | 3.3619 | 4.3139 | H10 | 2.0861 | 3.2297 | 1.0931 | 3.3346 | 2.6269 | 2.2084 | 3.2516 | 3.8474 | 1.7631 | | 4.1511 | 3.6082 | 2.3881 | 3.6082 | 2.3607 | 2.9471 | 4.3104 | 3.3540 | 4.3139 | 4.7745 | H11 | 2.0863 | 3.2314 | 3.3347 | 1.0931 | 2.6284 | 3.8473 | 3.2558 | 2.2084 | 3.6089 | 4.1511 | | 1.7631 | 3.6089 | 2.3881 | 4.7764 | 4.3104 | 3.3619 | 4.3139 | 2.9458 | 2.3607 | H12 | 2.0861 | 3.2297 | 2.6269 | 1.0931 | 3.3346 | 3.2516 | 3.8474 | 2.2084 | 2.3881 | 3.6082 | 1.7631 | | 4.1511 | 3.6082 | 4.3104 | 3.3540 | 4.3139 | 4.7745 | 2.3607 | 2.9471 | H13 | 2.0863 | 3.2314 | 2.6284 | 3.3347 | 1.0931 | 3.2558 | 2.2084 | 3.8473 | 3.6089 | 2.3881 | 3.6089 | 4.1511 | | 1.7631 | 3.3619 | 4.3139 | 2.9458 | 2.3607 | 4.7764 | 4.3104 | H14 | 2.0861 | 3.2297 | 3.3346 | 2.6269 | 1.0931 | 3.8474 | 2.2084 | 3.2516 | 4.1511 | 3.6082 | 2.3881 | 3.6082 | 1.7631 | | 4.3139 | 4.7745 | 2.3607 | 2.9471 | 4.3104 | 3.3540 | H15 | 3.2314 | 2.0863 | 2.2084 | 3.8473 | 3.2558 | 1.0931 | 2.6284 | 3.3347 | 2.9458 | 2.3607 | 4.7764 | 4.3104 | 3.3619 | 4.3139 | | 1.7631 | 3.6089 | 2.3881 | 3.6089 | 4.1511 | H16 | 3.2297 | 2.0861 | 2.2084 | 3.2516 | 3.8474 | 1.0931 | 3.3346 | 2.6269 | 2.3607 | 2.9471 | 4.3104 | 3.3540 | 4.3139 | 4.7745 | 1.7631 | | 4.1511 | 3.6082 | 2.3881 | 3.6082 | H17 | 3.2314 | 2.0863 | 3.8473 | 3.2558 | 2.2084 | 3.3347 | 1.0931 | 2.6284 | 4.7764 | 4.3104 | 3.3619 | 4.3139 | 2.9458 | 2.3607 | 3.6089 | 4.1511 | | 1.7631 | 3.6089 | 2.3881 | H18 | 3.2297 | 2.0861 | 3.2516 | 3.8474 | 2.2084 | 2.6269 | 1.0931 | 3.3346 | 4.3104 | 3.3540 | 4.3139 | 4.7745 | 2.3607 | 2.9471 | 2.3881 | 3.6082 | 1.7631 | | 4.1511 | 3.6082 | H19 | 3.2314 | 2.0863 | 3.2558 | 2.2084 | 3.8473 | 2.6284 | 3.3347 | 1.0931 | 3.3619 | 4.3139 | 2.9458 | 2.3607 | 4.7764 | 4.3104 | 3.6089 | 2.3881 | 3.6089 | 4.1511 | | 1.7631 | H20 | 3.2297 | 2.0861 | 3.8474 | 2.2084 | 3.2516 | 3.3346 | 2.6269 | 1.0931 | 4.3139 | 4.7745 | 2.3607 | 2.9471 | 4.3104 | 3.3540 | 4.1511 | 3.6082 | 2.3881 | 3.6082 | 1.7631 | |
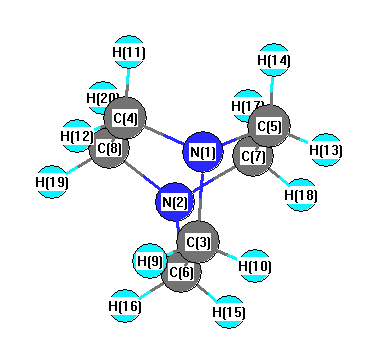 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.234 |
|
N1 |
C3 |
H9 |
107.995 |
| N1 |
C3 |
H10 |
107.974 |
|
N1 |
C4 |
C8 |
110.234 |
| N1 |
C4 |
H11 |
107.995 |
|
N1 |
C4 |
H12 |
107.974 |
| N1 |
C5 |
C7 |
110.234 |
|
N1 |
C5 |
H13 |
107.995 |
| N1 |
C5 |
H14 |
107.974 |
|
N2 |
C6 |
C3 |
110.234 |
| N2 |
C6 |
H15 |
107.995 |
|
N2 |
C6 |
H16 |
107.974 |
| N2 |
C7 |
C5 |
110.234 |
|
N2 |
C7 |
H17 |
107.995 |
| N2 |
C7 |
H18 |
107.974 |
|
N2 |
C8 |
C4 |
110.234 |
| N2 |
C8 |
H19 |
107.995 |
|
N2 |
C8 |
H20 |
107.974 |
| C3 |
N1 |
C4 |
108.697 |
|
C3 |
N1 |
C5 |
108.697 |
| C3 |
C6 |
H15 |
111.497 |
|
C3 |
C6 |
H16 |
111.491 |
| C4 |
N1 |
C5 |
108.697 |
|
C4 |
C8 |
H19 |
111.497 |
| C4 |
C8 |
H20 |
111.491 |
|
C5 |
C6 |
H15 |
101.574 |
| C5 |
C6 |
H16 |
150.902 |
|
C6 |
N2 |
C7 |
108.697 |
| C6 |
N2 |
C8 |
108.697 |
|
C6 |
C3 |
H9 |
111.497 |
| C6 |
C3 |
H10 |
111.491 |
|
C7 |
N2 |
C8 |
108.697 |
| C7 |
C5 |
H13 |
111.497 |
|
C7 |
C5 |
H14 |
111.491 |
| C8 |
C4 |
H11 |
111.497 |
|
C8 |
C4 |
H12 |
111.491 |
| H9 |
C3 |
H10 |
107.499 |
|
H11 |
C4 |
H12 |
107.499 |
| H13 |
C5 |
H14 |
107.499 |
|
H15 |
C6 |
H16 |
107.499 |
| H17 |
C7 |
H18 |
107.499 |
|
H19 |
C8 |
H20 |
107.499 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
