Jump to
S1C2
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -345.373072 |
| Energy at 298.15K | -345.390075 |
| HF Energy | -345.373072 |
| Nuclear repulsion energy | 423.648062 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3073 |
2946 |
0.00 |
|
|
|
| 2 |
A1 |
3046 |
2920 |
0.00 |
|
|
|
| 3 |
A1 |
1518 |
1455 |
0.00 |
|
|
|
| 4 |
A1 |
1366 |
1309 |
0.00 |
|
|
|
| 5 |
A1 |
1270 |
1217 |
0.00 |
|
|
|
| 6 |
A1 |
1020 |
978 |
0.00 |
|
|
|
| 7 |
A1 |
965 |
925 |
0.00 |
|
|
|
| 8 |
A1 |
807 |
774 |
0.00 |
|
|
|
| 9 |
A1 |
595 |
570 |
0.00 |
|
|
|
| 10 |
A1 |
83 |
80 |
0.00 |
|
|
|
| 11 |
A2 |
3093 |
2965 |
0.00 |
|
|
|
| 12 |
A2 |
3032 |
2906 |
133.27 |
|
|
|
| 13 |
A2 |
1510 |
1447 |
1.96 |
|
|
|
| 14 |
A2 |
1388 |
1331 |
9.92 |
|
|
|
| 15 |
A2 |
1194 |
1144 |
0.00 |
|
|
|
| 16 |
A2 |
987 |
947 |
20.50 |
|
|
|
| 17 |
A2 |
802 |
769 |
0.01 |
|
|
|
| 18 |
A2 |
763 |
731 |
62.78 |
|
|
|
| 19 |
E |
3099 |
2971 |
104.04 |
|
|
|
| 19 |
E |
3099 |
2971 |
104.02 |
|
|
|
| 20 |
E |
3075 |
2948 |
0.00 |
|
|
|
| 20 |
E |
3075 |
2948 |
0.00 |
|
|
|
| 21 |
E |
3039 |
2914 |
124.00 |
|
|
|
| 21 |
E |
3039 |
2914 |
123.87 |
|
|
|
| 22 |
E |
3031 |
2906 |
0.00 |
|
|
|
| 22 |
E |
3031 |
2906 |
0.00 |
|
|
|
| 23 |
E |
1509 |
1447 |
5.89 |
|
|
|
| 23 |
E |
1509 |
1447 |
5.92 |
|
|
|
| 24 |
E |
1497 |
1435 |
0.00 |
|
|
|
| 24 |
E |
1497 |
1435 |
0.00 |
|
|
|
| 25 |
E |
1355 |
1299 |
7.94 |
|
|
|
| 25 |
E |
1355 |
1299 |
7.94 |
|
|
|
| 26 |
E |
1348 |
1292 |
0.00 |
|
|
|
| 26 |
E |
1348 |
1292 |
0.00 |
|
|
|
| 27 |
E |
1333 |
1277 |
0.22 |
|
|
|
| 27 |
E |
1333 |
1277 |
0.22 |
|
|
|
| 28 |
E |
1330 |
1275 |
0.00 |
|
|
|
| 28 |
E |
1330 |
1275 |
0.00 |
|
|
|
| 29 |
E |
1206 |
1156 |
0.00 |
|
|
|
| 29 |
E |
1206 |
1156 |
0.00 |
|
|
|
| 30 |
E |
1077 |
1033 |
39.25 |
|
|
|
| 30 |
E |
1077 |
1033 |
39.26 |
|
|
|
| 31 |
E |
1039 |
996 |
0.01 |
|
|
|
| 31 |
E |
1039 |
996 |
0.01 |
|
|
|
| 32 |
E |
898 |
860 |
3.83 |
|
|
|
| 32 |
E |
898 |
860 |
3.83 |
|
|
|
| 33 |
E |
822 |
788 |
3.92 |
|
|
|
| 33 |
E |
822 |
788 |
3.92 |
|
|
|
| 34 |
E |
574 |
550 |
0.00 |
|
|
|
| 34 |
E |
574 |
550 |
0.00 |
|
|
|
| 35 |
E |
406 |
389 |
0.00 |
|
|
|
| 35 |
E |
406 |
389 |
0.00 |
|
|
|
| 36 |
E |
322 |
309 |
0.00 |
|
|
|
| 36 |
E |
322 |
309 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40212.4 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 38551.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.292 |
| N2 |
0.000 |
0.000 |
-1.292 |
| C3 |
-0.001 |
1.384 |
0.782 |
| C4 |
1.199 |
-0.691 |
0.782 |
| C5 |
-1.198 |
-0.693 |
0.782 |
| C6 |
0.001 |
1.384 |
-0.782 |
| C7 |
-1.199 |
-0.691 |
-0.782 |
| C8 |
1.198 |
-0.693 |
-0.782 |
| H9 |
0.882 |
1.892 |
1.186 |
| H10 |
-0.888 |
1.888 |
1.183 |
| H11 |
1.197 |
-1.710 |
1.186 |
| H12 |
2.079 |
-0.175 |
1.183 |
| H13 |
-2.079 |
-0.182 |
1.186 |
| H14 |
-1.191 |
-1.713 |
1.183 |
| H15 |
-0.882 |
1.892 |
-1.186 |
| H16 |
0.888 |
1.888 |
-1.183 |
| H17 |
-1.197 |
-1.710 |
-1.186 |
| H18 |
-2.079 |
-0.175 |
-1.183 |
| H19 |
2.079 |
-0.182 |
-1.186 |
| H20 |
1.191 |
-1.713 |
-1.183 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5846 | 1.4748 | 1.4748 | 1.4748 | 2.4934 | 2.4934 | 2.4934 | 2.0899 | 2.0895 | 2.0899 | 2.0895 | 2.0899 | 2.0895 | 3.2400 | 3.2374 | 3.2400 | 3.2374 | 3.2400 | 3.2374 |
N2 | 2.5846 | | 2.4934 | 2.4934 | 2.4934 | 1.4748 | 1.4748 | 1.4748 | 3.2400 | 3.2374 | 3.2400 | 3.2374 | 3.2400 | 3.2374 | 2.0899 | 2.0895 | 2.0899 | 2.0895 | 2.0899 | 2.0895 | C3 | 1.4748 | 2.4934 | | 2.3967 | 2.3967 | 1.5638 | 2.8607 | 2.8628 | 1.0961 | 1.0961 | 3.3420 | 2.6305 | 2.6330 | 3.3417 | 2.2150 | 2.2149 | 3.8564 | 3.2572 | 3.2638 | 3.8566 | C4 | 1.4748 | 2.4934 | 2.3967 | | 2.3967 | 2.8607 | 2.8628 | 1.5638 | 2.6330 | 3.3417 | 1.0961 | 1.0961 | 3.3420 | 2.6305 | 3.8564 | 3.2572 | 3.2638 | 3.8566 | 2.2150 | 2.2149 | C5 | 1.4748 | 2.4934 | 2.3967 | 2.3967 | | 2.8628 | 1.5638 | 2.8607 | 3.3420 | 2.6305 | 2.6330 | 3.3417 | 1.0961 | 1.0961 | 3.2638 | 3.8566 | 2.2150 | 2.2149 | 3.8564 | 3.2572 | C6 | 2.4934 | 1.4748 | 1.5638 | 2.8607 | 2.8628 | | 2.3967 | 2.3967 | 2.2150 | 2.2149 | 3.8564 | 3.2572 | 3.2638 | 3.8566 | 1.0961 | 1.0961 | 3.3420 | 2.6305 | 2.6330 | 3.3417 | C7 | 2.4934 | 1.4748 | 2.8607 | 2.8628 | 1.5638 | 2.3967 | | 2.3967 | 3.8564 | 3.2572 | 3.2638 | 3.8566 | 2.2150 | 2.2149 | 2.6330 | 3.3417 | 1.0961 | 1.0961 | 3.3420 | 2.6305 | C8 | 2.4934 | 1.4748 | 2.8628 | 1.5638 | 2.8607 | 2.3967 | 2.3967 | | 3.2638 | 3.8566 | 2.2150 | 2.2149 | 3.8564 | 3.2572 | 3.3420 | 2.6305 | 2.6330 | 3.3417 | 1.0961 | 1.0961 | H9 | 2.0899 | 3.2400 | 1.0961 | 2.6330 | 3.3420 | 2.2150 | 3.8564 | 3.2638 | | 1.7699 | 3.6151 | 2.3886 | 3.6151 | 4.1585 | 2.9559 | 2.3688 | 4.7873 | 4.3189 | 3.3703 | 4.3245 | H10 | 2.0895 | 3.2374 | 1.0961 | 3.3417 | 2.6305 | 2.2149 | 3.2572 | 3.8566 | 1.7699 | | 4.1585 | 3.6141 | 2.3886 | 3.6141 | 2.3688 | 2.9581 | 4.3189 | 3.3578 | 4.3245 | 4.7844 | H11 | 2.0899 | 3.2400 | 3.3420 | 1.0961 | 2.6330 | 3.8564 | 3.2638 | 2.2150 | 3.6151 | 4.1585 | | 1.7699 | 3.6151 | 2.3886 | 4.7873 | 4.3189 | 3.3703 | 4.3245 | 2.9559 | 2.3688 | H12 | 2.0895 | 3.2374 | 2.6305 | 1.0961 | 3.3417 | 3.2572 | 3.8566 | 2.2149 | 2.3886 | 3.6141 | 1.7699 | | 4.1585 | 3.6141 | 4.3189 | 3.3578 | 4.3245 | 4.7844 | 2.3688 | 2.9581 | H13 | 2.0899 | 3.2400 | 2.6330 | 3.3420 | 1.0961 | 3.2638 | 2.2150 | 3.8564 | 3.6151 | 2.3886 | 3.6151 | 4.1585 | | 1.7699 | 3.3703 | 4.3245 | 2.9559 | 2.3688 | 4.7873 | 4.3189 | H14 | 2.0895 | 3.2374 | 3.3417 | 2.6305 | 1.0961 | 3.8566 | 2.2149 | 3.2572 | 4.1585 | 3.6141 | 2.3886 | 3.6141 | 1.7699 | | 4.3245 | 4.7844 | 2.3688 | 2.9581 | 4.3189 | 3.3578 | H15 | 3.2400 | 2.0899 | 2.2150 | 3.8564 | 3.2638 | 1.0961 | 2.6330 | 3.3420 | 2.9559 | 2.3688 | 4.7873 | 4.3189 | 3.3703 | 4.3245 | | 1.7699 | 3.6151 | 2.3886 | 3.6151 | 4.1585 | H16 | 3.2374 | 2.0895 | 2.2149 | 3.2572 | 3.8566 | 1.0961 | 3.3417 | 2.6305 | 2.3688 | 2.9581 | 4.3189 | 3.3578 | 4.3245 | 4.7844 | 1.7699 | | 4.1585 | 3.6141 | 2.3886 | 3.6141 | H17 | 3.2400 | 2.0899 | 3.8564 | 3.2638 | 2.2150 | 3.3420 | 1.0961 | 2.6330 | 4.7873 | 4.3189 | 3.3703 | 4.3245 | 2.9559 | 2.3688 | 3.6151 | 4.1585 | | 1.7699 | 3.6151 | 2.3886 | H18 | 3.2374 | 2.0895 | 3.2572 | 3.8566 | 2.2149 | 2.6305 | 1.0961 | 3.3417 | 4.3189 | 3.3578 | 4.3245 | 4.7844 | 2.3688 | 2.9581 | 2.3886 | 3.6141 | 1.7699 | | 4.1585 | 3.6141 | H19 | 3.2400 | 2.0899 | 3.2638 | 2.2150 | 3.8564 | 2.6330 | 3.3420 | 1.0961 | 3.3703 | 4.3245 | 2.9559 | 2.3688 | 4.7873 | 4.3189 | 3.6151 | 2.3886 | 3.6151 | 4.1585 | | 1.7699 | H20 | 3.2374 | 2.0895 | 3.8566 | 2.2149 | 3.2572 | 3.3417 | 2.6305 | 1.0961 | 4.3245 | 4.7844 | 2.3688 | 2.9581 | 4.3189 | 3.3578 | 4.1585 | 3.6141 | 2.3886 | 3.6141 | 1.7699 | |
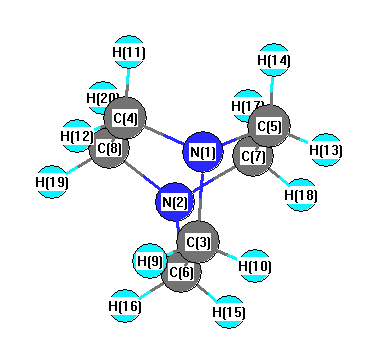 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.246 |
|
N1 |
C3 |
H9 |
107.852 |
| N1 |
C3 |
H10 |
107.818 |
|
N1 |
C4 |
C8 |
110.246 |
| N1 |
C4 |
H11 |
107.852 |
|
N1 |
C4 |
H12 |
107.818 |
| N1 |
C5 |
C7 |
110.246 |
|
N1 |
C5 |
H13 |
107.852 |
| N1 |
C5 |
H14 |
107.818 |
|
N2 |
C6 |
C3 |
110.246 |
| N2 |
C6 |
H15 |
107.852 |
|
N2 |
C6 |
H16 |
107.818 |
| N2 |
C7 |
C5 |
110.246 |
|
N2 |
C7 |
H17 |
107.852 |
| N2 |
C7 |
H18 |
107.818 |
|
N2 |
C8 |
C4 |
110.246 |
| N2 |
C8 |
H19 |
107.852 |
|
N2 |
C8 |
H20 |
107.818 |
| C3 |
N1 |
C4 |
108.686 |
|
C3 |
N1 |
C5 |
108.686 |
| C3 |
C6 |
H15 |
111.547 |
|
C3 |
C6 |
H16 |
111.541 |
| C4 |
N1 |
C5 |
108.686 |
|
C4 |
C8 |
H19 |
111.547 |
| C4 |
C8 |
H20 |
111.541 |
|
C5 |
C6 |
H15 |
101.537 |
| C5 |
C6 |
H16 |
150.754 |
|
C6 |
N2 |
C7 |
108.686 |
| C6 |
N2 |
C8 |
108.686 |
|
C6 |
C3 |
H9 |
111.547 |
| C6 |
C3 |
H10 |
111.541 |
|
C7 |
N2 |
C8 |
108.686 |
| C7 |
C5 |
H13 |
111.547 |
|
C7 |
C5 |
H14 |
111.541 |
| C8 |
C4 |
H11 |
111.547 |
|
C8 |
C4 |
H12 |
111.541 |
| H9 |
C3 |
H10 |
107.676 |
|
H11 |
C4 |
H12 |
107.677 |
| H13 |
C5 |
H14 |
107.677 |
|
H15 |
C6 |
H16 |
107.676 |
| H17 |
C7 |
H18 |
107.677 |
|
H19 |
C8 |
H20 |
107.677 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.411 |
|
|
|
| 2 |
N |
-0.411 |
|
|
|
| 3 |
C |
-0.107 |
|
|
|
| 4 |
C |
-0.107 |
|
|
|
| 5 |
C |
-0.107 |
|
|
|
| 6 |
C |
-0.107 |
|
|
|
| 7 |
C |
-0.107 |
|
|
|
| 8 |
C |
-0.107 |
|
|
|
| 9 |
H |
0.122 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.122 |
|
|
|
| 16 |
H |
0.122 |
|
|
|
| 17 |
H |
0.122 |
|
|
|
| 18 |
H |
0.122 |
|
|
|
| 19 |
H |
0.122 |
|
|
|
| 20 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.175 |
0.000 |
0.000 |
| y |
0.000 |
11.174 |
0.000 |
| z |
0.000 |
0.000 |
9.619 |
<r2> (average value of r
2) Å
2
| <r2> |
214.646 |
| (<r2>)1/2 |
14.651 |
