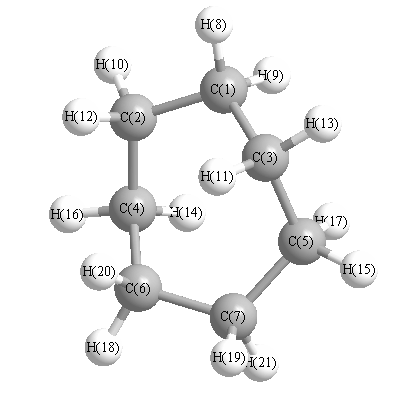
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3102 |
2956 |
5.05 |
|
|
|
| 2 |
A |
3095 |
2950 |
25.85 |
|
|
|
| 3 |
A |
3088 |
2943 |
74.87 |
|
|
|
| 4 |
A |
3061 |
2917 |
24.54 |
|
|
|
| 5 |
A |
3052 |
2909 |
2.38 |
|
|
|
| 6 |
A |
3049 |
2906 |
6.03 |
|
|
|
| 7 |
A |
3043 |
2900 |
18.44 |
|
|
|
| 8 |
A |
1537 |
1465 |
0.34 |
|
|
|
| 9 |
A |
1520 |
1449 |
6.10 |
|
|
|
| 10 |
A |
1511 |
1440 |
0.01 |
|
|
|
| 11 |
A |
1508 |
1437 |
0.30 |
|
|
|
| 12 |
A |
1423 |
1356 |
0.23 |
|
|
|
| 13 |
A |
1417 |
1350 |
1.13 |
|
|
|
| 14 |
A |
1395 |
1329 |
2.00 |
|
|
|
| 15 |
A |
1333 |
1271 |
0.00 |
|
|
|
| 16 |
A |
1323 |
1261 |
1.52 |
|
|
|
| 17 |
A |
1249 |
1191 |
0.48 |
|
|
|
| 18 |
A |
1235 |
1177 |
0.12 |
|
|
|
| 19 |
A |
1148 |
1094 |
0.08 |
|
|
|
| 20 |
A |
1066 |
1016 |
0.51 |
|
|
|
| 21 |
A |
1045 |
996 |
0.00 |
|
|
|
| 22 |
A |
973 |
927 |
1.82 |
|
|
|
| 23 |
A |
863 |
823 |
0.03 |
|
|
|
| 24 |
A |
853 |
813 |
2.15 |
|
|
|
| 25 |
A |
755 |
720 |
0.03 |
|
|
|
| 26 |
A |
520 |
496 |
0.30 |
|
|
|
| 27 |
A |
399 |
380 |
0.00 |
|
|
|
| 28 |
A |
308 |
294 |
0.01 |
|
|
|
| 29 |
A |
173 |
165 |
0.01 |
|
|
|
| 30 |
B |
3111 |
2965 |
111.07 |
|
|
|
| 31 |
B |
3100 |
2955 |
88.92 |
|
|
|
| 32 |
B |
3094 |
2949 |
57.35 |
|
|
|
| 33 |
B |
3085 |
2940 |
9.46 |
|
|
|
| 34 |
B |
3053 |
2910 |
42.65 |
|
|
|
| 35 |
B |
3048 |
2905 |
22.64 |
|
|
|
| 36 |
B |
3042 |
2900 |
23.07 |
|
|
|
| 37 |
B |
1529 |
1458 |
19.99 |
|
|
|
| 38 |
B |
1513 |
1442 |
8.24 |
|
|
|
| 39 |
B |
1506 |
1436 |
0.75 |
|
|
|
| 40 |
B |
1417 |
1350 |
1.61 |
|
|
|
| 41 |
B |
1408 |
1342 |
0.17 |
|
|
|
| 42 |
B |
1401 |
1336 |
1.79 |
|
|
|
| 43 |
B |
1367 |
1303 |
0.13 |
|
|
|
| 44 |
B |
1326 |
1264 |
1.40 |
|
|
|
| 45 |
B |
1247 |
1188 |
0.60 |
|
|
|
| 46 |
B |
1220 |
1163 |
1.21 |
|
|
|
| 47 |
B |
1165 |
1110 |
0.37 |
|
|
|
| 48 |
B |
1046 |
997 |
0.38 |
|
|
|
| 49 |
B |
1036 |
988 |
0.63 |
|
|
|
| 50 |
B |
977 |
931 |
2.44 |
|
|
|
| 51 |
B |
881 |
840 |
0.34 |
|
|
|
| 52 |
B |
829 |
790 |
3.16 |
|
|
|
| 53 |
B |
742 |
708 |
1.63 |
|
|
|
| 54 |
B |
492 |
469 |
0.36 |
|
|
|
| 55 |
B |
335 |
319 |
0.07 |
|
|
|
| 56 |
B |
276 |
263 |
0.12 |
|
|
|
| 57 |
B |
117 |
111 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 44201.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 42128.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.322 |
|
|
|
| 2 |
C |
-0.326 |
|
|
|
| 3 |
C |
-0.326 |
|
|
|
| 4 |
C |
-0.320 |
|
|
|
| 5 |
C |
-0.320 |
|
|
|
| 6 |
C |
-0.320 |
|
|
|
| 7 |
C |
-0.320 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.160 |
|
|
|
| 11 |
H |
0.160 |
|
|
|
| 12 |
H |
0.164 |
|
|
|
| 13 |
H |
0.164 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.163 |
|
|
|
| 16 |
H |
0.160 |
|
|
|
| 17 |
H |
0.160 |
|
|
|
| 18 |
H |
0.157 |
|
|
|
| 19 |
H |
0.157 |
|
|
|
| 20 |
H |
0.161 |
|
|
|
| 21 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.001 |
0.001 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.908 |
-0.103 |
0.000 |
| y |
-0.103 |
-45.958 |
0.000 |
| z |
0.000 |
0.000 |
-46.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.224 |
-0.103 |
0.000 |
| y |
-0.103 |
0.149 |
0.000 |
| z |
0.000 |
0.000 |
-0.373 |
|
| Polar |
|---|
| 3z2-r2 | -0.746 |
|---|
| x2-y2 | 0.050 |
|---|
| xy | -0.103 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.568 |
0.352 |
0.000 |
| y |
0.352 |
10.840 |
0.000 |
| z |
0.000 |
0.000 |
11.147 |
<r2> (average value of r
2) Å
2
| <r2> |
227.499 |
| (<r2>)1/2 |
15.083 |