Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -957.757169 |
| Energy at 298.15K | |
| HF Energy | -957.358260 |
| Nuclear repulsion energy | 126.221684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3384 |
3162 |
0.05 |
|
|
|
| 2 |
A1 |
793 |
741 |
11.03 |
|
|
|
| 3 |
A1 |
327 |
306 |
0.48 |
|
|
|
| 4 |
B1 |
443i |
414i |
54.49 |
|
|
|
| 5 |
B2 |
1321 |
1235 |
63.65 |
|
|
|
| 6 |
B2 |
983 |
919 |
145.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3182.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 2973.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.667 |
| H2 |
0.000 |
0.000 |
1.739 |
| Cl3 |
0.000 |
1.471 |
-0.169 |
| Cl4 |
0.000 |
-1.471 |
-0.169 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0721 | 1.6918 | 1.6918 |
H2 | 1.0721 | | 2.4091 | 2.4091 | Cl3 | 1.6918 | 2.4091 | | 2.9419 | Cl4 | 1.6918 | 2.4091 | 2.9419 | |
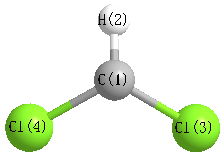 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.606 |
|
Cl3 |
C1 |
Cl4 |
120.788 |
| Cl4 |
C1 |
H2 |
119.606 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -957.758356 |
| Energy at 298.15K | -957.759251 |
| HF Energy | -957.359687 |
| Nuclear repulsion energy | 126.026183 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3333 |
3114 |
2.49 |
|
|
|
| 2 |
A' |
801 |
748 |
17.75 |
|
|
|
| 3 |
A' |
559 |
522 |
22.36 |
|
|
|
| 4 |
A' |
321 |
300 |
0.48 |
|
|
|
| 5 |
A" |
1325 |
1238 |
53.86 |
|
|
|
| 6 |
A" |
949 |
886 |
170.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3643.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 3404.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
0.692 |
0.000 |
| H2 |
-0.494 |
1.642 |
0.000 |
| Cl3 |
0.012 |
-0.170 |
1.467 |
| Cl4 |
0.012 |
-0.170 |
-1.467 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0763 | 1.7020 | 1.7020 |
H2 | 1.0763 | | 2.3861 | 2.3861 | Cl3 | 1.7020 | 2.3861 | | 2.9348 | Cl4 | 1.7020 | 2.3861 | 2.9348 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
116.558 |
|
Cl3 |
C1 |
Cl4 |
119.115 |
| Cl4 |
C1 |
H2 |
116.558 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
