Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -958.698260 |
| Energy at 298.15K | |
| HF Energy | -958.698260 |
| Nuclear repulsion energy | 125.859331 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3304 |
3153 |
0.18 |
|
|
|
| 2 |
A1 |
773 |
737 |
10.64 |
|
|
|
| 3 |
A1 |
319 |
304 |
0.43 |
|
|
|
| 4 |
B1 |
325i |
310i |
57.11 |
|
|
|
| 5 |
B2 |
1264 |
1206 |
54.33 |
|
|
|
| 6 |
B2 |
950 |
907 |
156.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3142.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 2998.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.668 |
| H2 |
0.000 |
0.000 |
1.747 |
| Cl3 |
0.000 |
1.475 |
-0.169 |
| Cl4 |
0.000 |
-1.475 |
-0.169 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0793 | 1.6962 | 1.6962 |
H2 | 1.0793 | | 2.4185 | 2.4185 | Cl3 | 1.6962 | 2.4185 | | 2.9504 | Cl4 | 1.6962 | 2.4185 | 2.9504 | |
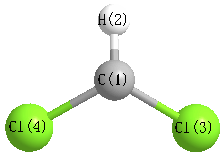 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.575 |
|
Cl3 |
C1 |
Cl4 |
120.849 |
| Cl4 |
C1 |
H2 |
119.575 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.356 |
|
|
|
| 2 |
H |
0.217 |
|
|
|
| 3 |
Cl |
0.070 |
|
|
|
| 4 |
Cl |
0.070 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.009 |
1.009 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.693 |
0.000 |
0.000 |
| y |
0.000 |
-30.909 |
0.000 |
| z |
0.000 |
0.000 |
-28.645 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.916 |
0.000 |
0.000 |
| y |
0.000 |
-0.740 |
0.000 |
| z |
0.000 |
0.000 |
2.656 |
|
| Polar |
|---|
| 3z2-r2 | 5.313 |
|---|
| x2-y2 | -0.784 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.260 |
0.000 |
0.000 |
| y |
0.000 |
6.462 |
0.000 |
| z |
0.000 |
0.000 |
3.644 |
<r2> (average value of r
2) Å
2
| <r2> |
99.692 |
| (<r2>)1/2 |
9.985 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -958.698759 |
| Energy at 298.15K | -958.699530 |
| HF Energy | -958.698759 |
| Nuclear repulsion energy | 125.738197 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3271 |
3121 |
1.36 |
|
|
|
| 2 |
A' |
777 |
742 |
15.12 |
|
|
|
| 3 |
A' |
437 |
417 |
33.38 |
|
|
|
| 4 |
A' |
311 |
297 |
1.50 |
|
|
|
| 5 |
A" |
1267 |
1210 |
46.85 |
|
|
|
| 6 |
A" |
926 |
884 |
177.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3494.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 3334.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.686 |
0.000 |
| H2 |
-0.435 |
1.672 |
0.000 |
| Cl3 |
0.011 |
-0.170 |
1.472 |
| Cl4 |
0.011 |
-0.170 |
-1.472 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0819 | 1.7034 | 1.7034 |
H2 | 1.0819 | | 2.4003 | 2.4003 | Cl3 | 1.7034 | 2.4003 | | 2.9447 | Cl4 | 1.7034 | 2.4003 | 2.9447 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
117.277 |
|
Cl3 |
C1 |
Cl4 |
119.614 |
| Cl4 |
C1 |
H2 |
117.277 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.339 |
|
|
|
| 2 |
H |
0.214 |
|
|
|
| 3 |
Cl |
0.062 |
|
|
|
| 4 |
Cl |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.482 |
0.932 |
0.000 |
1.049 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.460 |
-0.842 |
0.000 |
| y |
-0.842 |
-29.028 |
0.000 |
| z |
0.000 |
0.000 |
-31.039 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.426 |
-0.842 |
0.000 |
| y |
-0.842 |
2.222 |
0.000 |
| z |
0.000 |
0.000 |
-0.796 |
|
| Polar |
|---|
| 3z2-r2 | -1.591 |
|---|
| x2-y2 | -2.432 |
|---|
| xy | -0.842 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.342 |
-0.183 |
0.000 |
| y |
-0.183 |
3.641 |
0.000 |
| z |
0.000 |
0.000 |
6.522 |
<r2> (average value of r
2) Å
2
| <r2> |
99.564 |
| (<r2>)1/2 |
9.978 |
