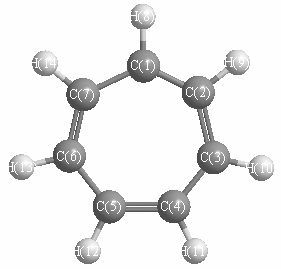
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3221 |
3074 |
3.66 |
|
|
|
| 2 |
A1 |
3210 |
3063 |
41.44 |
|
|
|
| 3 |
A1 |
3177 |
3032 |
4.44 |
|
|
|
| 4 |
A1 |
3171 |
3026 |
6.97 |
|
|
|
| 5 |
A1 |
1663 |
1587 |
3.00 |
|
|
|
| 6 |
A1 |
1564 |
1492 |
10.59 |
|
|
|
| 7 |
A1 |
1482 |
1414 |
4.69 |
|
|
|
| 8 |
A1 |
1288 |
1229 |
0.89 |
|
|
|
| 9 |
A1 |
1199 |
1144 |
2.22 |
|
|
|
| 10 |
A1 |
976 |
932 |
1.93 |
|
|
|
| 11 |
A1 |
915 |
873 |
0.83 |
|
|
|
| 12 |
A1 |
861 |
821 |
0.23 |
|
|
|
| 13 |
A1 |
444 |
423 |
0.27 |
|
|
|
| 14 |
A2 |
997 |
951 |
0.00 |
|
|
|
| 15 |
A2 |
847 |
808 |
0.00 |
|
|
|
| 16 |
A2 |
759 |
724 |
0.00 |
|
|
|
| 17 |
A2 |
567 |
541 |
0.00 |
|
|
|
| 18 |
A2 |
154 |
147 |
0.00 |
|
|
|
| 19 |
B1 |
1011 |
965 |
0.00 |
|
|
|
| 20 |
B1 |
986 |
941 |
0.41 |
|
|
|
| 21 |
B1 |
772 |
736 |
0.89 |
|
|
|
| 22 |
B1 |
654 |
624 |
60.18 |
|
|
|
| 23 |
B1 |
514 |
491 |
37.75 |
|
|
|
| 24 |
B1 |
288 |
275 |
0.00 |
|
|
|
| 25 |
B2 |
3213 |
3066 |
31.35 |
|
|
|
| 26 |
B2 |
3196 |
3050 |
19.27 |
|
|
|
| 27 |
B2 |
3172 |
3027 |
6.66 |
|
|
|
| 28 |
B2 |
1668 |
1592 |
9.17 |
|
|
|
| 29 |
B2 |
1492 |
1423 |
3.22 |
|
|
|
| 30 |
B2 |
1413 |
1348 |
0.01 |
|
|
|
| 31 |
B2 |
1307 |
1247 |
0.00 |
|
|
|
| 32 |
B2 |
1251 |
1194 |
1.41 |
|
|
|
| 33 |
B2 |
989 |
944 |
3.29 |
|
|
|
| 34 |
B2 |
906 |
865 |
1.43 |
|
|
|
| 35 |
B2 |
416 |
397 |
0.30 |
|
|
|
| 36 |
B2 |
56i |
53i |
8.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24841.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 23706.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.081 |
|
|
|
| 2 |
C |
-0.158 |
|
|
|
| 3 |
C |
-0.158 |
|
|
|
| 4 |
C |
-0.096 |
|
|
|
| 5 |
C |
-0.096 |
|
|
|
| 6 |
C |
-0.131 |
|
|
|
| 7 |
C |
-0.131 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.039 |
0.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.292 |
0.000 |
0.000 |
| y |
0.000 |
-36.195 |
0.000 |
| z |
0.000 |
0.000 |
-36.199 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.095 |
0.000 |
0.000 |
| y |
0.000 |
4.550 |
0.000 |
| z |
0.000 |
0.000 |
4.545 |
|
| Polar |
|---|
| 3z2-r2 | 9.090 |
|---|
| x2-y2 | -9.097 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.698 |
0.000 |
0.000 |
| y |
0.000 |
13.322 |
0.000 |
| z |
0.000 |
0.000 |
13.335 |
<r2> (average value of r
2) Å
2
| <r2> |
185.435 |
| (<r2>)1/2 |
13.617 |