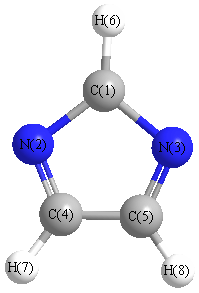
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3280 |
3127 |
2.96 |
|
|
|
| 2 |
A1 |
3272 |
3118 |
13.78 |
|
|
|
| 3 |
A1 |
1493 |
1423 |
25.52 |
|
|
|
| 4 |
A1 |
1336 |
1274 |
54.83 |
|
|
|
| 5 |
A1 |
1193 |
1137 |
27.12 |
|
|
|
| 6 |
A1 |
1014 |
966 |
2.66 |
|
|
|
| 7 |
A1 |
924 |
881 |
3.83 |
|
|
|
| 8 |
A2 |
886 |
844 |
0.00 |
|
|
|
| 9 |
A2 |
567 |
541 |
0.00 |
|
|
|
| 10 |
B1 |
876 |
835 |
0.35 |
|
|
|
| 11 |
B1 |
763 |
727 |
20.81 |
|
|
|
| 12 |
B1 |
519 |
494 |
30.04 |
|
|
|
| 13 |
B2 |
3258 |
3105 |
1.52 |
|
|
|
| 14 |
B2 |
1566 |
1493 |
19.31 |
|
|
|
| 15 |
B2 |
1309 |
1248 |
0.02 |
|
|
|
| 16 |
B2 |
1207 |
1150 |
5.83 |
|
|
|
| 17 |
B2 |
1046 |
997 |
71.76 |
|
|
|
| 18 |
B2 |
813 |
775 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12659.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12066.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.182 |
|
|
|
| 2 |
N |
-0.404 |
|
|
|
| 3 |
N |
-0.404 |
|
|
|
| 4 |
C |
0.009 |
|
|
|
| 5 |
C |
0.009 |
|
|
|
| 6 |
H |
0.206 |
|
|
|
| 7 |
H |
0.201 |
|
|
|
| 8 |
H |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.680 |
1.680 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.898 |
0.000 |
0.000 |
| y |
0.000 |
-33.900 |
0.000 |
| z |
0.000 |
0.000 |
-21.957 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.970 |
0.000 |
0.000 |
| y |
0.000 |
-8.472 |
0.000 |
| z |
0.000 |
0.000 |
9.442 |
|
| Polar |
|---|
| 3z2-r2 | 18.884 |
|---|
| x2-y2 | 5.001 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.584 |
0.000 |
0.000 |
| y |
0.000 |
5.857 |
0.000 |
| z |
0.000 |
0.000 |
7.172 |
<r2> (average value of r
2) Å
2
| <r2> |
75.893 |
| (<r2>)1/2 |
8.712 |