return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-307.429432
Energy at 298.15K-307.434552
HF Energy-306.344274
Nuclear repulsion energy312.529965
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3325 3107 10.95      
2 A1 3282 3067 15.34      
3 A1 3267 3053 8.19      
4 A1 1727 1613 1.37      
5 A1 1572 1469 1.38      
6 A1 1544 1443 2.02      
7 A1 1474 1377 9.27      
8 A1 1212 1133 0.10      
9 A1 1146 1070 0.24      
10 A1 1093 1022 0.32      
11 A1 1015 948 2.57      
12 A1 828 774 0.37      
13 A1 551 515 0.06      
14 A2 893 834 0.00      
15 A2 868 811 0.00      
16 A2 799 747 0.00      
17 A2 509 475 0.00      
18 A2 338 316 0.00      
19 A2 281 262 0.00      
20 B1 854 798 3.69      
21 B1 739 690 71.83      
22 B1 682 637 31.55      
23 B1 330 308 0.25      
24 B1 212 198 9.13      
25 B2 3296 3080 1.83      
26 B2 3274 3060 26.32      
27 B2 3256 3042 0.08      
28 B2 1678 1568 0.53      
29 B2 1496 1398 3.53      
30 B2 1332 1245 4.17      
31 B2 1279 1195 13.35      
32 B2 1131 1056 1.10      
33 B2 1060 991 2.08      
34 B2 874 817 0.37      
35 B2 657 614 1.32      
36 B2 423 395 3.22      

Unscaled Zero Point Vibrational Energy (zpe) 24147.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 22563.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.16082 0.07344 0.05041

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.444 -0.622
C2 0.000 0.688 -1.841
C3 0.000 -0.688 -1.841
C4 0.000 -1.444 -0.622
C5 0.000 -0.678 2.041
C6 0.000 0.678 2.041
C7 0.000 0.716 0.528
C8 0.000 -0.716 0.528
H9 0.000 2.525 -0.651
H10 0.000 1.222 -2.781
H11 0.000 -1.222 -2.781
H12 0.000 -2.525 -0.651
H13 0.000 -1.443 2.802
H14 0.000 1.443 2.802

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.43502.45662.88803.40442.77021.36052.44641.08172.17103.43133.96944.47833.4233
C21.43501.37662.45664.11553.88212.36922.75392.18921.08102.12933.42715.10914.7042
C32.45661.37661.43503.88214.11552.75392.36923.42712.12931.08102.18924.70425.1091
C42.88802.45661.43502.77023.40442.44641.36053.96943.43132.17101.08173.42334.4783
C53.40444.11553.88212.77021.35582.05701.51354.18385.18284.85253.26441.07932.2535
C62.77023.88214.11553.40441.35581.51352.05703.26444.85255.18284.18382.25351.0793
C71.36052.36922.75392.44642.05701.51351.43152.15943.34743.83473.44863.13572.3877
C82.44642.75392.36921.36051.51352.05701.43153.44863.83473.34742.15942.38773.1357
H91.08172.18923.42713.96944.18383.26442.15943.44862.49744.31105.05075.26013.6181
H102.17101.08102.12933.43135.18284.85253.34743.83472.49742.44484.31106.18675.5874
H113.43132.12931.08102.17104.85255.18283.83473.34744.31102.44482.49745.58746.1867
H123.96943.42712.18921.08173.26444.18383.44862.15945.05074.31102.49743.61815.2601
H134.47835.10914.70423.42331.07932.25353.13572.38775.26016.18675.58743.61812.8864
H143.42334.70425.10914.47832.25351.07932.38773.13573.61815.58746.18675.26012.8864

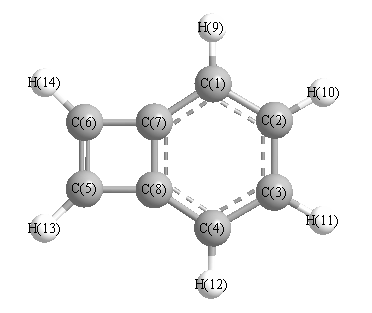
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.777 C1 C2 H10 118.615
C1 C7 C6 149.068 C1 C7 N8 122.363
C2 C1 C7 115.859 C2 C1 H9 120.232
C2 C3 C4 121.777 C2 C3 H11 119.608
C3 C2 H10 119.608 C3 C4 N8 115.859
C3 C4 H12 120.232 C4 C3 H11 118.615
C4 N8 C5 149.068 C4 N8 C7 122.363
C5 C6 C7 91.432 C5 C6 H14 135.157
C5 N8 C7 88.568 C6 C5 N8 91.432
C6 C5 H13 135.157 C6 C7 N8 88.568
C7 C1 H9 123.908 C7 C6 H14 133.411
N8 C4 H12 123.908 N8 C5 H13 133.411
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability