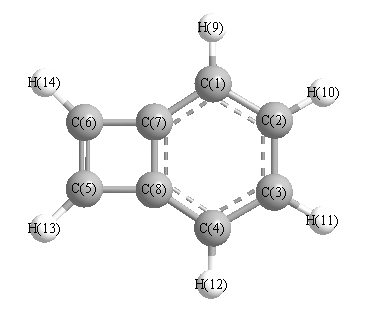
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3287 |
3133 |
11.70 |
|
|
|
| 2 |
A1 |
3239 |
3087 |
13.54 |
|
|
|
| 3 |
A1 |
3224 |
3073 |
11.51 |
|
|
|
| 4 |
A1 |
1744 |
1663 |
3.07 |
|
|
|
| 5 |
A1 |
1615 |
1539 |
0.68 |
|
|
|
| 6 |
A1 |
1541 |
1469 |
2.22 |
|
|
|
| 7 |
A1 |
1467 |
1398 |
11.11 |
|
|
|
| 8 |
A1 |
1200 |
1144 |
0.01 |
|
|
|
| 9 |
A1 |
1129 |
1076 |
0.06 |
|
|
|
| 10 |
A1 |
1080 |
1030 |
0.80 |
|
|
|
| 11 |
A1 |
1003 |
956 |
4.06 |
|
|
|
| 12 |
A1 |
834 |
795 |
0.39 |
|
|
|
| 13 |
A1 |
552 |
526 |
0.13 |
|
|
|
| 14 |
A2 |
968 |
923 |
0.00 |
|
|
|
| 15 |
A2 |
917 |
874 |
0.00 |
|
|
|
| 16 |
A2 |
882 |
840 |
0.00 |
|
|
|
| 17 |
A2 |
782 |
746 |
0.00 |
|
|
|
| 18 |
A2 |
584 |
557 |
0.00 |
|
|
|
| 19 |
A2 |
297 |
283 |
0.00 |
|
|
|
| 20 |
B1 |
915 |
872 |
8.34 |
|
|
|
| 21 |
B1 |
756 |
720 |
105.71 |
|
|
|
| 22 |
B1 |
728 |
694 |
0.60 |
|
|
|
| 23 |
B1 |
363 |
346 |
1.80 |
|
|
|
| 24 |
B1 |
238 |
226 |
10.14 |
|
|
|
| 25 |
B2 |
3256 |
3104 |
1.72 |
|
|
|
| 26 |
B2 |
3233 |
3081 |
29.34 |
|
|
|
| 27 |
B2 |
3211 |
3060 |
0.07 |
|
|
|
| 28 |
B2 |
1693 |
1614 |
0.47 |
|
|
|
| 29 |
B2 |
1482 |
1413 |
3.85 |
|
|
|
| 30 |
B2 |
1321 |
1259 |
4.72 |
|
|
|
| 31 |
B2 |
1266 |
1207 |
12.17 |
|
|
|
| 32 |
B2 |
1114 |
1062 |
1.76 |
|
|
|
| 33 |
B2 |
1060 |
1010 |
1.10 |
|
|
|
| 34 |
B2 |
881 |
839 |
0.54 |
|
|
|
| 35 |
B2 |
655 |
625 |
0.79 |
|
|
|
| 36 |
B2 |
427 |
407 |
4.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24472.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 23324.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.265 |
|
|
|
| 2 |
C |
-0.176 |
|
|
|
| 3 |
C |
-0.176 |
|
|
|
| 4 |
C |
-0.265 |
|
|
|
| 5 |
C |
-0.182 |
|
|
|
| 6 |
C |
-0.182 |
|
|
|
| 7 |
C |
0.103 |
|
|
|
| 8 |
C |
0.103 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.172 |
|
|
|
| 13 |
H |
0.183 |
|
|
|
| 14 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.694 |
0.694 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.776 |
0.000 |
0.000 |
| y |
0.000 |
-40.022 |
0.000 |
| z |
0.000 |
0.000 |
-39.233 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.148 |
0.000 |
0.000 |
| y |
0.000 |
4.482 |
0.000 |
| z |
0.000 |
0.000 |
5.666 |
|
| Polar |
|---|
| 3z2-r2 | 11.332 |
|---|
| x2-y2 | -9.754 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.053 |
0.000 |
0.000 |
| y |
0.000 |
12.859 |
0.000 |
| z |
0.000 |
0.000 |
15.570 |
<r2> (average value of r
2) Å
2
| <r2> |
219.472 |
| (<r2>)1/2 |
14.815 |