Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.586896 |
| Energy at 298.15K | -235.599107 |
| HF Energy | -235.586896 |
| Nuclear repulsion energy | 226.732438 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3016 |
49.05 |
|
|
|
| 2 |
A |
3146 |
3002 |
30.46 |
|
|
|
| 3 |
A |
3144 |
3001 |
25.84 |
|
|
|
| 4 |
A |
3142 |
2998 |
4.78 |
|
|
|
| 5 |
A |
3138 |
2994 |
44.82 |
|
|
|
| 6 |
A |
3110 |
2968 |
22.00 |
|
|
|
| 7 |
A |
3103 |
2961 |
20.14 |
|
|
|
| 8 |
A |
3080 |
2939 |
7.38 |
|
|
|
| 9 |
A |
3058 |
2918 |
40.72 |
|
|
|
| 10 |
A |
3055 |
2916 |
22.04 |
|
|
|
| 11 |
A |
3048 |
2908 |
35.98 |
|
|
|
| 12 |
A |
3029 |
2890 |
25.53 |
|
|
|
| 13 |
A |
1779 |
1698 |
0.44 |
|
|
|
| 14 |
A |
1518 |
1449 |
5.26 |
|
|
|
| 15 |
A |
1509 |
1440 |
6.27 |
|
|
|
| 16 |
A |
1502 |
1434 |
2.35 |
|
|
|
| 17 |
A |
1501 |
1433 |
7.79 |
|
|
|
| 18 |
A |
1488 |
1420 |
5.91 |
|
|
|
| 19 |
A |
1485 |
1417 |
2.36 |
|
|
|
| 20 |
A |
1423 |
1358 |
1.61 |
|
|
|
| 21 |
A |
1422 |
1357 |
2.55 |
|
|
|
| 22 |
A |
1398 |
1335 |
0.88 |
|
|
|
| 23 |
A |
1348 |
1287 |
0.94 |
|
|
|
| 24 |
A |
1341 |
1279 |
0.43 |
|
|
|
| 25 |
A |
1323 |
1262 |
0.03 |
|
|
|
| 26 |
A |
1291 |
1232 |
5.94 |
|
|
|
| 27 |
A |
1257 |
1199 |
0.51 |
|
|
|
| 28 |
A |
1188 |
1133 |
0.04 |
|
|
|
| 29 |
A |
1124 |
1073 |
1.37 |
|
|
|
| 30 |
A |
1113 |
1062 |
2.47 |
|
|
|
| 31 |
A |
1072 |
1023 |
1.88 |
|
|
|
| 32 |
A |
1068 |
1019 |
0.22 |
|
|
|
| 33 |
A |
1048 |
1000 |
7.84 |
|
|
|
| 34 |
A |
1012 |
966 |
28.29 |
|
|
|
| 35 |
A |
935 |
892 |
6.52 |
|
|
|
| 36 |
A |
912 |
870 |
2.53 |
|
|
|
| 37 |
A |
878 |
838 |
0.41 |
|
|
|
| 38 |
A |
783 |
747 |
0.66 |
|
|
|
| 39 |
A |
747 |
713 |
3.10 |
|
|
|
| 40 |
A |
533 |
509 |
0.16 |
|
|
|
| 41 |
A |
391 |
373 |
1.21 |
|
|
|
| 42 |
A |
312 |
298 |
0.78 |
|
|
|
| 43 |
A |
286 |
273 |
1.46 |
|
|
|
| 44 |
A |
249 |
238 |
0.05 |
|
|
|
| 45 |
A |
211 |
201 |
0.79 |
|
|
|
| 46 |
A |
143 |
136 |
0.29 |
|
|
|
| 47 |
A |
99 |
94 |
0.04 |
|
|
|
| 48 |
A |
76 |
73 |
0.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36487.7 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 34820.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.176 |
0.215 |
-0.158 |
| C2 |
1.800 |
-0.366 |
-0.218 |
| C3 |
0.738 |
0.105 |
0.437 |
| C4 |
-0.642 |
-0.469 |
0.372 |
| C5 |
-1.678 |
0.519 |
-0.172 |
| C6 |
-3.089 |
-0.055 |
-0.190 |
| H7 |
3.212 |
1.094 |
0.491 |
| H8 |
3.901 |
-0.516 |
0.221 |
| H9 |
3.527 |
0.513 |
-1.154 |
| H10 |
1.674 |
-1.247 |
-0.849 |
| H11 |
0.862 |
0.989 |
1.066 |
| H12 |
-0.958 |
-0.785 |
1.378 |
| H13 |
-0.638 |
-1.375 |
-0.248 |
| H14 |
-1.383 |
0.822 |
-1.184 |
| H15 |
-1.659 |
1.433 |
0.437 |
| H16 |
-3.812 |
0.667 |
-0.582 |
| H17 |
-3.143 |
-0.952 |
-0.816 |
| H18 |
-3.414 |
-0.337 |
0.817 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4943 | 2.5115 | 3.9151 | 4.8641 | 6.2710 | 1.0939 | 1.0969 | 1.0969 | 2.2073 | 2.7304 | 4.5219 | 4.1330 | 4.7121 | 5.0213 | 7.0152 | 6.4595 | 6.6850 |
C2 | 1.4943 | | 1.3332 | 2.5149 | 3.5898 | 4.8993 | 2.1510 | 2.1516 | 2.1516 | 1.0916 | 2.0895 | 3.2140 | 2.6391 | 3.5321 | 3.9532 | 5.7178 | 5.0138 | 5.3164 | C3 | 2.5115 | 1.3332 | | 1.4967 | 2.5264 | 3.8817 | 2.6646 | 3.2302 | 3.2359 | 2.0871 | 1.0926 | 2.1343 | 2.1339 | 2.7639 | 2.7403 | 4.6966 | 4.2134 | 4.1935 | C4 | 3.9151 | 2.5149 | 1.4967 | | 1.5314 | 2.5444 | 4.1609 | 4.5461 | 4.5470 | 2.7312 | 2.2071 | 1.1006 | 1.0978 | 2.1531 | 2.1574 | 3.4995 | 2.8105 | 2.8106 | C5 | 4.8641 | 3.5898 | 2.5264 | 1.5314 | | 1.5229 | 4.9685 | 5.6883 | 5.2973 | 3.8489 | 2.8650 | 2.1495 | 2.1624 | 1.0969 | 1.0981 | 2.1774 | 2.1736 | 2.1735 | C6 | 6.2710 | 4.8993 | 3.8817 | 2.5444 | 1.5229 | | 6.4410 | 7.0174 | 6.7099 | 4.9536 | 4.2754 | 2.7447 | 2.7848 | 2.1605 | 2.1567 | 1.0942 | 1.0956 | 1.0956 | H7 | 1.0939 | 2.1510 | 2.6646 | 4.1609 | 4.9685 | 6.4410 | | 1.7721 | 1.7722 | 3.1057 | 2.4217 | 4.6589 | 4.6331 | 4.8978 | 4.8827 | 7.1179 | 6.8031 | 6.7869 | H8 | 1.0969 | 2.1516 | 3.2302 | 4.5461 | 5.6883 | 7.0174 | 1.7721 | | 1.7577 | 2.5775 | 3.4951 | 5.0019 | 4.6435 | 5.6290 | 5.8954 | 7.8443 | 7.1336 | 7.3418 | H9 | 1.0969 | 2.1516 | 3.2359 | 4.5470 | 5.2973 | 6.7099 | 1.7722 | 1.7577 | | 2.5745 | 3.5011 | 5.3111 | 4.6619 | 4.9197 | 5.5015 | 7.3625 | 6.8375 | 7.2656 | H10 | 2.2073 | 1.0916 | 2.0871 | 2.7312 | 3.8489 | 4.9536 | 3.1057 | 2.5775 | 2.5745 | | 3.0545 | 3.4781 | 2.3918 | 3.7064 | 4.4655 | 5.8159 | 4.8257 | 5.4305 | H11 | 2.7304 | 2.0895 | 1.0926 | 2.2071 | 2.8650 | 4.2754 | 2.4217 | 3.4951 | 3.5011 | 3.0545 | | 2.5603 | 3.0930 | 3.1827 | 2.6358 | 4.9664 | 4.8325 | 4.4841 | H12 | 4.5219 | 3.2140 | 2.1343 | 1.1006 | 2.1495 | 2.7447 | 4.6589 | 5.0019 | 5.3111 | 3.4781 | 2.5603 | | 1.7589 | 3.0537 | 2.5090 | 3.7544 | 3.1011 | 2.5590 | H13 | 4.1330 | 2.6391 | 2.1339 | 1.0978 | 2.1624 | 2.7848 | 4.6331 | 4.6435 | 4.6619 | 2.3918 | 3.0930 | 1.7589 | | 2.5020 | 3.0652 | 3.7889 | 2.6034 | 3.1496 | H14 | 4.7121 | 3.5321 | 2.7639 | 2.1531 | 1.0969 | 2.1605 | 4.8978 | 5.6290 | 4.9197 | 3.7064 | 3.1827 | 3.0537 | 2.5020 | | 1.7532 | 2.5070 | 2.5265 | 3.0782 | H15 | 5.0213 | 3.9532 | 2.7403 | 2.1574 | 1.0981 | 2.1567 | 4.8827 | 5.8954 | 5.5015 | 4.4655 | 2.6358 | 2.5090 | 3.0652 | 1.7532 | | 2.5019 | 3.0759 | 2.5219 | H16 | 7.0152 | 5.7178 | 4.6966 | 3.4995 | 2.1774 | 1.0942 | 7.1179 | 7.8443 | 7.3625 | 5.8159 | 4.9664 | 3.7544 | 3.7889 | 2.5070 | 2.5019 | | 1.7676 | 1.7680 | H17 | 6.4595 | 5.0138 | 4.2134 | 2.8105 | 2.1736 | 1.0956 | 6.8031 | 7.1336 | 6.8375 | 4.8257 | 4.8325 | 3.1011 | 2.6034 | 2.5265 | 3.0759 | 1.7676 | | 1.7663 | H18 | 6.6850 | 5.3164 | 4.1935 | 2.8106 | 2.1735 | 1.0956 | 6.7869 | 7.3418 | 7.2656 | 5.4305 | 4.4841 | 2.5590 | 3.1496 | 3.0782 | 2.5219 | 1.7680 | 1.7663 | |
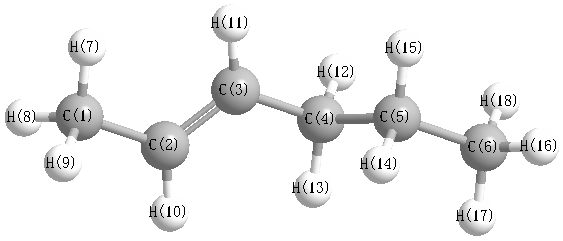 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.209 |
|
C1 |
C2 |
H10 |
116.336 |
| C2 |
C1 |
H7 |
111.480 |
|
C2 |
C1 |
H8 |
111.342 |
| C2 |
C1 |
H9 |
111.347 |
|
C2 |
C3 |
C4 |
125.321 |
| C2 |
C3 |
H11 |
118.600 |
|
C3 |
C2 |
H10 |
118.454 |
| C3 |
C4 |
C5 |
113.086 |
|
C3 |
C4 |
H12 |
109.579 |
| C3 |
C4 |
H13 |
109.711 |
|
C4 |
C3 |
H11 |
116.074 |
| C4 |
C5 |
C6 |
112.830 |
|
C4 |
C5 |
H14 |
108.882 |
| C4 |
C5 |
H15 |
109.149 |
|
C5 |
C4 |
H12 |
108.395 |
| C5 |
C4 |
H13 |
109.557 |
|
C5 |
C6 |
H16 |
111.560 |
| C5 |
C6 |
H17 |
111.170 |
|
C5 |
C6 |
H18 |
111.162 |
| C6 |
C5 |
H14 |
110.048 |
|
C6 |
C5 |
H15 |
109.684 |
| H7 |
C1 |
H8 |
107.971 |
|
H7 |
C1 |
H9 |
107.986 |
| H8 |
C1 |
H9 |
106.495 |
|
H12 |
C4 |
H13 |
106.277 |
| H14 |
C5 |
H15 |
106.015 |
|
H16 |
C6 |
H17 |
107.647 |
| H16 |
C6 |
H18 |
107.677 |
|
H17 |
C6 |
H18 |
107.428 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.426 |
|
|
|
| 2 |
C |
-0.099 |
|
|
|
| 3 |
C |
-0.082 |
|
|
|
| 4 |
C |
-0.268 |
|
|
|
| 5 |
C |
-0.236 |
|
|
|
| 6 |
C |
-0.391 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
| 9 |
H |
0.139 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
| 11 |
H |
0.102 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.122 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.128 |
|
|
|
| 18 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.032 |
0.037 |
-0.049 |
0.069 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.249 |
0.003 |
-0.647 |
| y |
0.003 |
-39.283 |
1.185 |
| z |
-0.647 |
1.185 |
-39.505 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.145 |
0.003 |
-0.647 |
| y |
0.003 |
-0.406 |
1.185 |
| z |
-0.647 |
1.185 |
-0.739 |
|
| Polar |
|---|
| 3z2-r2 | -1.479 |
|---|
| x2-y2 | 1.034 |
|---|
| xy | 0.003 |
|---|
| xz | -0.647 |
|---|
| yz | 1.185 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.309 |
-0.217 |
-0.944 |
| y |
-0.217 |
8.289 |
0.704 |
| z |
-0.944 |
0.704 |
8.058 |
<r2> (average value of r
2) Å
2
| <r2> |
294.380 |
| (<r2>)1/2 |
17.158 |
