return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCHCHCH3 (2-Butenal)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-230.554781
Energy at 298.15K-230.560618
HF Energy-229.811249
Nuclear repulsion energy153.002713
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3270 3056 3.78      
2 A' 3241 3028 18.33      
3 A' 3218 3007 6.08      
4 A' 3125 2920 12.54      
5 A' 2990 2794 102.08      
6 A' 1780 1663 170.86      
7 A' 1735 1621 4.73      
8 A' 1547 1446 12.07      
9 A' 1467 1371 1.26      
10 A' 1456 1360 1.42      
11 A' 1358 1269 3.68      
12 A' 1310 1224 0.61      
13 A' 1202 1123 47.06      
14 A' 1133 1059 16.82      
15 A' 970 907 17.86      
16 A' 547 511 8.02      
17 A' 467 436 2.23      
18 A' 209 195 6.53      
19 A" 3207 2997 11.71      
20 A" 1538 1437 6.95      
21 A" 1097 1025 1.17      
22 A" 1039 971 11.26      
23 A" 1013 947 22.39      
24 A" 784 733 0.02      
25 A" 290 271 7.03      
26 A" 197 184 0.51      
27 A" 126 117 1.90      

Unscaled Zero Point Vibrational Energy (zpe) 20157.8 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18835.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
1.09687 0.07270 0.06905

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.382 -0.479 0.000
C2 0.901 -0.659 0.000
C3 0.000 0.336 0.000
C4 -1.437 0.045 0.000
O5 -2.315 0.901 0.000
H6 -1.691 -1.032 0.000
H7 0.295 1.378 0.000
H8 0.533 -1.681 0.000
H9 2.651 0.574 0.000
H10 2.826 -0.950 0.876
H11 2.826 -0.950 -0.876

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.49142.51733.85394.89474.11042.79332.20501.08701.08971.0897
C21.49141.34262.44123.57392.61922.12511.08642.14092.13522.1352
C32.51731.34261.46592.38242.17581.08252.08672.66193.22673.2267
C43.85392.44121.46591.22631.10652.18552.61864.12214.46434.4643
O54.89473.57392.38241.22632.03092.65293.84374.97675.53385.5338
H64.11042.61922.17581.10652.03093.12332.31714.63024.60264.6026
H72.79332.12511.08252.18552.65293.12333.06832.48973.54923.5492
H82.20501.08642.08672.61863.84372.31713.06833.09402.56142.5614
H91.08702.14092.66194.12214.97674.63022.48973.09401.76681.7668
H101.08972.13523.22674.46435.53384.60263.54922.56141.76681.7519
H111.08972.13523.22674.46435.53384.60263.54922.56141.76681.7519

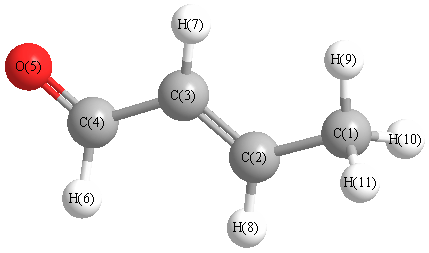
picture of 2-Butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.228 C1 C2 H8 116.729
C2 C1 H9 111.295 C2 C1 H10 110.673
C2 C1 H11 110.673 C2 C3 C4 120.672
C2 C3 H7 122.032 C3 C2 H8 118.042
C3 C4 O5 124.240 C3 C4 H6 114.795
C4 C3 H7 117.296 O5 C4 H6 120.965
H9 C1 H10 108.529 H9 C1 H11 108.529
H10 C1 H11 107.005
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability