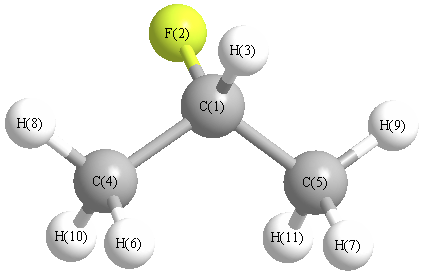
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3167 |
3022 |
24.45 |
|
|
|
| 2 |
A' |
3162 |
3017 |
44.56 |
|
|
|
| 3 |
A' |
3073 |
2933 |
7.53 |
|
|
|
| 4 |
A' |
3054 |
2914 |
39.55 |
|
|
|
| 5 |
A' |
1522 |
1453 |
5.19 |
|
|
|
| 6 |
A' |
1501 |
1432 |
5.63 |
|
|
|
| 7 |
A' |
1427 |
1362 |
22.26 |
|
|
|
| 8 |
A' |
1392 |
1329 |
13.84 |
|
|
|
| 9 |
A' |
1208 |
1153 |
15.40 |
|
|
|
| 10 |
A' |
1181 |
1127 |
53.54 |
|
|
|
| 11 |
A' |
981 |
936 |
31.11 |
|
|
|
| 12 |
A' |
844 |
806 |
6.03 |
|
|
|
| 13 |
A' |
477 |
455 |
2.60 |
|
|
|
| 14 |
A' |
350 |
334 |
1.04 |
|
|
|
| 15 |
A' |
263 |
251 |
0.08 |
|
|
|
| 16 |
A" |
3165 |
3021 |
14.98 |
|
|
|
| 17 |
A" |
3155 |
3011 |
0.02 |
|
|
|
| 18 |
A" |
3070 |
2929 |
13.71 |
|
|
|
| 19 |
A" |
1495 |
1427 |
0.00 |
|
|
|
| 20 |
A" |
1489 |
1421 |
0.04 |
|
|
|
| 21 |
A" |
1426 |
1360 |
34.02 |
|
|
|
| 22 |
A" |
1379 |
1316 |
1.84 |
|
|
|
| 23 |
A" |
1177 |
1124 |
12.60 |
|
|
|
| 24 |
A" |
949 |
906 |
0.00 |
|
|
|
| 25 |
A" |
931 |
889 |
0.99 |
|
|
|
| 26 |
A" |
406 |
388 |
6.21 |
|
|
|
| 27 |
A" |
222 |
212 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21232.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 20262.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.212 |
|
|
|
| 2 |
F |
-0.330 |
|
|
|
| 3 |
H |
0.110 |
|
|
|
| 4 |
C |
-0.409 |
|
|
|
| 5 |
C |
-0.409 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.313 |
-1.074 |
0.000 |
1.697 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.452 |
1.346 |
0.000 |
| y |
1.346 |
-25.933 |
0.000 |
| z |
0.000 |
0.000 |
-24.312 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.330 |
1.346 |
0.000 |
| y |
1.346 |
-1.051 |
0.000 |
| z |
0.000 |
0.000 |
1.381 |
|
| Polar |
|---|
| 3z2-r2 | 2.762 |
|---|
| x2-y2 | 0.481 |
|---|
| xy | 1.346 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.844 |
-0.019 |
0.000 |
| y |
-0.019 |
4.891 |
0.000 |
| z |
0.000 |
0.000 |
5.393 |
<r2> (average value of r
2) Å
2
| <r2> |
85.137 |
| (<r2>)1/2 |
9.227 |