Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -376.590463 |
| Energy at 298.15K | |
| HF Energy | -375.789226 |
| Nuclear repulsion energy | 190.789880 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3240 |
3028 |
27.64 |
36.00 |
0.72 |
0.84 |
| 2 |
A |
3197 |
2988 |
31.81 |
81.49 |
0.31 |
0.48 |
| 3 |
A |
3163 |
2955 |
16.75 |
88.78 |
0.09 |
0.17 |
| 4 |
A |
1568 |
1465 |
3.17 |
8.77 |
0.74 |
0.85 |
| 5 |
A |
1519 |
1419 |
16.28 |
2.50 |
0.62 |
0.77 |
| 6 |
A |
1466 |
1370 |
27.81 |
4.60 |
0.74 |
0.85 |
| 7 |
A |
1391 |
1300 |
28.21 |
3.06 |
0.74 |
0.85 |
| 8 |
A |
1299 |
1214 |
9.72 |
8.98 |
0.73 |
0.84 |
| 9 |
A |
1206 |
1127 |
90.10 |
5.22 |
0.74 |
0.85 |
| 10 |
A |
1184 |
1106 |
19.73 |
2.20 |
0.48 |
0.64 |
| 11 |
A |
1161 |
1084 |
176.57 |
1.54 |
0.41 |
0.58 |
| 12 |
A |
1132 |
1058 |
31.38 |
6.14 |
0.47 |
0.64 |
| 13 |
A |
946 |
884 |
32.96 |
5.18 |
0.32 |
0.49 |
| 14 |
A |
579 |
541 |
4.55 |
1.76 |
0.39 |
0.56 |
| 15 |
A |
490 |
457 |
18.94 |
0.73 |
0.72 |
0.84 |
| 16 |
A |
431 |
403 |
6.06 |
1.41 |
0.53 |
0.70 |
| 17 |
A |
242 |
227 |
8.44 |
0.04 |
0.53 |
0.70 |
| 18 |
A |
129 |
120 |
7.34 |
0.01 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12171.0 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 11372.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.755 |
-0.613 |
-0.273 |
| C2 |
0.466 |
0.022 |
0.334 |
| F3 |
-1.871 |
0.096 |
0.143 |
| F4 |
1.550 |
-0.739 |
-0.014 |
| F5 |
0.636 |
1.272 |
-0.182 |
| H6 |
-0.674 |
-0.570 |
-1.357 |
| H7 |
-0.837 |
-1.647 |
0.055 |
| H8 |
0.417 |
0.099 |
1.418 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5045 | 1.3860 | 2.3233 | 2.3451 | 1.0880 | 1.0879 | 2.1782 |
C2 | 1.5045 | | 2.3463 | 1.3695 | 1.3635 | 2.1239 | 2.1358 | 1.0883 | F3 | 1.3860 | 2.3463 | | 3.5252 | 2.7889 | 2.0313 | 2.0285 | 2.6202 | F4 | 2.3233 | 1.3695 | 3.5252 | | 2.2160 | 2.6041 | 2.5549 | 2.0093 | F5 | 2.3451 | 1.3635 | 2.7889 | 2.2160 | | 2.5479 | 3.2788 | 1.9966 | H6 | 1.0880 | 2.1239 | 2.0313 | 2.6041 | 2.5479 | | 1.7836 | 3.0568 | H7 | 1.0879 | 2.1358 | 2.0285 | 2.5549 | 3.2788 | 1.7836 | | 2.5462 | H8 | 2.1782 | 1.0883 | 2.6202 | 2.0093 | 1.9966 | 3.0568 | 2.5462 | |
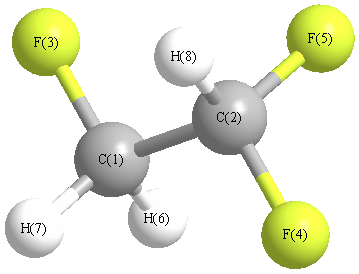 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.789 |
|
C1 |
C2 |
F5 |
109.609 |
| C1 |
C2 |
H8 |
113.320 |
|
C2 |
C1 |
F3 |
108.463 |
| C2 |
C1 |
H6 |
108.958 |
|
C2 |
C1 |
H7 |
109.913 |
| F3 |
C1 |
H6 |
109.798 |
|
F3 |
C1 |
H7 |
109.573 |
| F4 |
C2 |
F5 |
108.352 |
|
F4 |
C2 |
H8 |
109.140 |
| F5 |
C2 |
H8 |
108.521 |
|
H6 |
C1 |
H7 |
110.109 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -376.587963 |
| Energy at 298.15K | |
| HF Energy | -375.786197 |
| Nuclear repulsion energy | 193.843507 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3168 |
2961 |
63.20 |
94.64 |
0.09 |
0.17 |
| 2 |
A' |
3149 |
2943 |
8.19 |
63.37 |
0.29 |
0.44 |
| 3 |
A' |
1565 |
1462 |
4.30 |
7.58 |
0.74 |
0.85 |
| 4 |
A' |
1495 |
1397 |
28.04 |
2.95 |
0.54 |
0.70 |
| 5 |
A' |
1473 |
1376 |
28.82 |
3.40 |
0.72 |
0.84 |
| 6 |
A' |
1219 |
1139 |
99.75 |
5.91 |
0.44 |
0.61 |
| 7 |
A' |
1137 |
1063 |
12.55 |
2.95 |
0.75 |
0.86 |
| 8 |
A' |
919 |
859 |
36.98 |
7.06 |
0.16 |
0.27 |
| 9 |
A' |
775 |
724 |
49.23 |
2.69 |
0.53 |
0.70 |
| 10 |
A' |
512 |
478 |
12.21 |
1.49 |
0.75 |
0.86 |
| 11 |
A' |
236 |
221 |
1.87 |
0.10 |
0.39 |
0.56 |
| 12 |
A" |
3222 |
3011 |
23.18 |
48.83 |
0.75 |
0.86 |
| 13 |
A" |
1461 |
1365 |
37.78 |
1.87 |
0.75 |
0.86 |
| 14 |
A" |
1315 |
1229 |
14.64 |
15.46 |
0.75 |
0.86 |
| 15 |
A" |
1191 |
1113 |
87.60 |
0.98 |
0.75 |
0.86 |
| 16 |
A" |
1002 |
937 |
59.26 |
3.77 |
0.75 |
0.86 |
| 17 |
A" |
381 |
356 |
0.12 |
0.15 |
0.75 |
0.86 |
| 18 |
A" |
120 |
112 |
1.99 |
0.03 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12170.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 11372.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.752 |
-0.820 |
0.000 |
| C2 |
0.361 |
0.632 |
0.000 |
| F3 |
-0.392 |
-1.595 |
0.000 |
| F4 |
-0.392 |
0.904 |
1.105 |
| F5 |
-0.392 |
0.904 |
-1.105 |
| H6 |
1.339 |
-1.040 |
-0.891 |
| H7 |
1.339 |
-1.040 |
0.891 |
| H8 |
1.231 |
1.290 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5034 | 1.3823 | 2.3462 | 2.3462 | 1.0890 | 1.0890 | 2.1642 |
C2 | 1.5034 | | 2.3512 | 1.3651 | 1.3651 | 2.1312 | 2.1312 | 1.0908 | F3 | 1.3823 | 2.3512 | | 2.7332 | 2.7332 | 2.0246 | 2.0246 | 3.3111 | F4 | 2.3462 | 1.3651 | 2.7332 | | 2.2102 | 3.2801 | 2.6117 | 2.0011 | F5 | 2.3462 | 1.3651 | 2.7332 | 2.2102 | | 2.6117 | 3.2801 | 2.0011 | H6 | 1.0890 | 2.1312 | 2.0246 | 3.2801 | 2.6117 | | 1.7817 | 2.4969 | H7 | 1.0890 | 2.1312 | 2.0246 | 2.6117 | 3.2801 | 1.7817 | | 2.4969 | H8 | 2.1642 | 1.0908 | 3.3111 | 2.0011 | 2.0011 | 2.4969 | 2.4969 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.657 |
|
C1 |
C2 |
F5 |
109.657 |
| C1 |
C2 |
H8 |
112.090 |
|
C2 |
C1 |
F3 |
109.057 |
| C2 |
C1 |
H6 |
109.551 |
|
C2 |
C1 |
H7 |
109.551 |
| F3 |
C1 |
H6 |
109.442 |
|
F3 |
C1 |
H7 |
109.442 |
| F4 |
C2 |
F5 |
108.097 |
|
F4 |
C2 |
H8 |
108.624 |
| F5 |
C2 |
H8 |
108.624 |
|
H6 |
C1 |
H7 |
109.781 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
