Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -377.185353 |
| Energy at 298.15K | |
| HF Energy | -377.185353 |
| Nuclear repulsion energy | 191.253963 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3002 |
32.51 |
40.49 |
0.71 |
0.83 |
| 2 |
A |
3103 |
2961 |
35.26 |
89.88 |
0.34 |
0.50 |
| 3 |
A |
3081 |
2940 |
18.05 |
105.90 |
0.09 |
0.16 |
| 4 |
A |
1514 |
1445 |
2.72 |
9.26 |
0.73 |
0.84 |
| 5 |
A |
1479 |
1412 |
17.64 |
3.26 |
0.68 |
0.81 |
| 6 |
A |
1433 |
1367 |
29.42 |
5.02 |
0.74 |
0.85 |
| 7 |
A |
1358 |
1296 |
24.38 |
3.31 |
0.75 |
0.86 |
| 8 |
A |
1276 |
1218 |
12.96 |
9.21 |
0.73 |
0.84 |
| 9 |
A |
1193 |
1139 |
106.26 |
4.82 |
0.69 |
0.82 |
| 10 |
A |
1162 |
1109 |
198.88 |
1.56 |
0.67 |
0.80 |
| 11 |
A |
1155 |
1102 |
16.97 |
2.58 |
0.60 |
0.75 |
| 12 |
A |
1136 |
1084 |
17.61 |
5.55 |
0.44 |
0.61 |
| 13 |
A |
928 |
885 |
26.82 |
5.11 |
0.36 |
0.53 |
| 14 |
A |
580 |
554 |
3.86 |
1.71 |
0.42 |
0.60 |
| 15 |
A |
481 |
459 |
16.80 |
0.74 |
0.73 |
0.84 |
| 16 |
A |
429 |
409 |
5.61 |
1.46 |
0.52 |
0.69 |
| 17 |
A |
238 |
227 |
7.47 |
0.05 |
0.54 |
0.70 |
| 18 |
A |
121 |
115 |
6.62 |
0.01 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 11905.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 11361.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.767 |
-0.599 |
-0.272 |
| C2 |
0.470 |
0.022 |
0.330 |
| F3 |
-1.871 |
0.105 |
0.144 |
| F4 |
1.536 |
-0.749 |
-0.013 |
| F5 |
0.658 |
1.262 |
-0.182 |
| H6 |
-0.688 |
-0.561 |
-1.364 |
| H7 |
-0.848 |
-1.642 |
0.053 |
| H8 |
0.417 |
0.101 |
1.422 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5091 | 1.3737 | 2.3219 | 2.3459 | 1.0953 | 1.0949 | 2.1816 |
C2 | 1.5091 | | 2.3493 | 1.3596 | 1.3543 | 2.1327 | 2.1404 | 1.0962 | F3 | 1.3737 | 2.3493 | | 3.5153 | 2.7999 | 2.0283 | 2.0259 | 2.6203 | F4 | 2.3219 | 1.3596 | 3.5153 | | 2.2012 | 2.6090 | 2.5460 | 2.0077 | F5 | 2.3459 | 1.3543 | 2.7999 | 2.2012 | | 2.5560 | 3.2795 | 1.9946 | H6 | 1.0953 | 2.1327 | 2.0283 | 2.6090 | 2.5560 | | 1.7894 | 3.0688 | H7 | 1.0949 | 2.1404 | 2.0259 | 2.5460 | 3.2795 | 1.7894 | | 2.5509 | H8 | 2.1816 | 1.0962 | 2.6203 | 2.0077 | 1.9946 | 3.0688 | 2.5509 | |
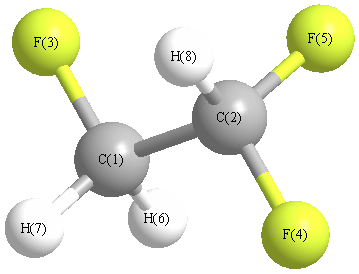 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.959 |
|
C1 |
C2 |
F5 |
109.907 |
| C1 |
C2 |
H8 |
112.769 |
|
C2 |
C1 |
F3 |
109.067 |
| C2 |
C1 |
H6 |
108.909 |
|
C2 |
C1 |
H7 |
109.533 |
| F3 |
C1 |
H6 |
109.957 |
|
F3 |
C1 |
H7 |
109.788 |
| F4 |
C2 |
F5 |
108.403 |
|
F4 |
C2 |
H8 |
109.206 |
| F5 |
C2 |
H8 |
108.509 |
|
H6 |
C1 |
H7 |
109.569 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.035 |
|
|
|
| 2 |
C |
0.445 |
|
|
|
| 3 |
F |
-0.305 |
|
|
|
| 4 |
F |
-0.290 |
|
|
|
| 5 |
F |
-0.281 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
| 8 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.179 |
-1.334 |
0.347 |
1.389 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.111 |
1.360 |
1.171 |
| y |
1.360 |
-27.028 |
0.471 |
| z |
1.171 |
0.471 |
-24.421 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.386 |
1.360 |
1.171 |
| y |
1.360 |
0.738 |
0.471 |
| z |
1.171 |
0.471 |
4.648 |
|
| Polar |
|---|
| 3z2-r2 | 9.296 |
|---|
| x2-y2 | -4.082 |
|---|
| xy | 1.360 |
|---|
| xz | 1.171 |
|---|
| yz | 0.471 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.495 |
-0.004 |
0.019 |
| y |
-0.004 |
3.530 |
0.034 |
| z |
0.019 |
0.034 |
3.477 |
<r2> (average value of r
2) Å
2
| <r2> |
110.156 |
| (<r2>)1/2 |
10.496 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -377.182934 |
| Energy at 298.15K | |
| HF Energy | -377.182934 |
| Nuclear repulsion energy | 194.093861 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3080 |
2939 |
67.45 |
123.85 |
0.03 |
0.06 |
| 2 |
A' |
3065 |
2925 |
9.44 |
53.58 |
0.64 |
0.78 |
| 3 |
A' |
1510 |
1441 |
5.20 |
8.10 |
0.73 |
0.84 |
| 4 |
A' |
1458 |
1392 |
30.28 |
2.99 |
0.57 |
0.73 |
| 5 |
A' |
1434 |
1369 |
26.14 |
4.35 |
0.72 |
0.84 |
| 6 |
A' |
1213 |
1158 |
117.74 |
5.23 |
0.40 |
0.57 |
| 7 |
A' |
1141 |
1089 |
17.94 |
3.09 |
0.74 |
0.85 |
| 8 |
A' |
897 |
856 |
30.49 |
6.90 |
0.21 |
0.34 |
| 9 |
A' |
775 |
739 |
46.25 |
2.69 |
0.53 |
0.70 |
| 10 |
A' |
515 |
491 |
11.56 |
1.48 |
0.75 |
0.86 |
| 11 |
A' |
229 |
218 |
1.66 |
0.10 |
0.38 |
0.55 |
| 12 |
A" |
3129 |
2986 |
26.48 |
55.64 |
0.75 |
0.86 |
| 13 |
A" |
1426 |
1361 |
39.34 |
1.80 |
0.75 |
0.86 |
| 14 |
A" |
1294 |
1235 |
17.10 |
16.38 |
0.75 |
0.86 |
| 15 |
A" |
1180 |
1126 |
99.01 |
1.09 |
0.75 |
0.86 |
| 16 |
A" |
980 |
936 |
48.93 |
3.75 |
0.75 |
0.86 |
| 17 |
A" |
372 |
355 |
0.17 |
0.14 |
0.75 |
0.86 |
| 18 |
A" |
110 |
105 |
1.83 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11903.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 11359.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.739 |
-0.828 |
0.000 |
| C2 |
0.355 |
0.630 |
0.000 |
| F3 |
-0.388 |
-1.607 |
0.000 |
| F4 |
-0.388 |
0.915 |
1.097 |
| F5 |
-0.388 |
0.915 |
-1.097 |
| H6 |
1.336 |
-1.044 |
-0.894 |
| H7 |
1.336 |
-1.044 |
0.894 |
| H8 |
1.244 |
1.276 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5080 | 1.3703 | 2.3478 | 2.3478 | 1.0961 | 1.0961 | 2.1644 |
C2 | 1.5080 | | 2.3580 | 1.3558 | 1.3558 | 2.1360 | 2.1360 | 1.0987 | F3 | 1.3703 | 2.3580 | | 2.7505 | 2.7505 | 2.0221 | 2.0221 | 3.3138 | F4 | 2.3478 | 1.3558 | 2.7505 | | 2.1948 | 3.2822 | 2.6173 | 1.9999 | F5 | 2.3478 | 1.3558 | 2.7505 | 2.1948 | | 2.6173 | 3.2822 | 1.9999 | H6 | 1.0961 | 2.1360 | 2.0221 | 3.2822 | 2.6173 | | 1.7871 | 2.4881 | H7 | 1.0961 | 2.1360 | 2.0221 | 2.6173 | 3.2822 | 1.7871 | | 2.4881 | H8 | 2.1644 | 1.0987 | 3.3138 | 1.9999 | 1.9999 | 2.4881 | 2.4881 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.022 |
|
C1 |
C2 |
F5 |
110.022 |
| C1 |
C2 |
H8 |
111.293 |
|
C2 |
C1 |
F3 |
109.926 |
| C2 |
C1 |
H6 |
109.201 |
|
C2 |
C1 |
H7 |
109.201 |
| F3 |
C1 |
H6 |
109.639 |
|
F3 |
C1 |
H7 |
109.639 |
| F4 |
C2 |
F5 |
108.071 |
|
F4 |
C2 |
H8 |
108.678 |
| F5 |
C2 |
H8 |
108.678 |
|
H6 |
C1 |
H7 |
109.217 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.048 |
|
|
|
| 2 |
C |
0.455 |
|
|
|
| 3 |
F |
-0.299 |
|
|
|
| 4 |
F |
-0.284 |
|
|
|
| 5 |
F |
-0.284 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.836 |
0.089 |
0.000 |
2.837 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.654 |
-0.301 |
0.000 |
| y |
-0.301 |
-28.045 |
0.000 |
| z |
0.000 |
0.000 |
-27.742 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.239 |
-0.301 |
0.000 |
| y |
-0.301 |
-1.847 |
0.000 |
| z |
0.000 |
0.000 |
-1.392 |
|
| Polar |
|---|
| 3z2-r2 | -2.783 |
|---|
| x2-y2 | 3.391 |
|---|
| xy | -0.301 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.523 |
0.016 |
0.000 |
| y |
0.016 |
3.519 |
0.000 |
| z |
0.000 |
0.000 |
3.520 |
<r2> (average value of r
2) Å
2
| <r2> |
101.862 |
| (<r2>)1/2 |
10.093 |
