return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11 (2-Methylbut-2-yl radical)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-196.455687
Energy at 298.15K-196.466681
HF Energy-195.724390
Nuclear repulsion energy182.713221
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3235 3022 27.22      
2 A 3228 3016 22.03      
3 A 3223 3012 38.09      
4 A 3216 3005 19.04      
5 A 3174 2966 22.36      
6 A 3165 2958 24.73      
7 A 3143 2937 21.79      
8 A 3133 2928 21.36      
9 A 3069 2867 37.36      
10 A 3065 2864 26.72      
11 A 3041 2842 33.78      
12 A 1573 1469 1.27      
13 A 1564 1461 4.83      
14 A 1556 1454 2.30      
15 A 1550 1448 14.92      
16 A 1544 1443 1.18      
17 A 1537 1436 2.53      
18 A 1529 1429 0.09      
19 A 1470 1374 3.68      
20 A 1465 1369 1.10      
21 A 1456 1360 3.65      
22 A 1427 1333 0.02      
23 A 1356 1267 1.01      
24 A 1324 1237 5.04      
25 A 1292 1208 5.76      
26 A 1157 1082 1.09      
27 A 1111 1038 0.21      
28 A 1081 1010 3.78      
29 A 1027 960 0.54      
30 A 1018 952 1.23      
31 A 976 912 0.46      
32 A 969 906 0.35      
33 A 792 740 0.98      
34 A 768 717 1.38      
35 A 465 434 0.62      
36 A 388 363 0.08      
37 A 316 295 1.78      
38 A 293 274 0.02      
39 A 222 207 0.77      
40 A 152 142 0.00      
41 A 142 132 0.05      
42 A 69 65 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 33140.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 30966.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.24943 0.11563 0.08552

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.586 1.475 0.067
H2 0.276 1.998 -0.341
H3 -1.480 1.885 -0.400
H4 -0.636 1.726 1.134
C5 2.049 -0.042 -0.163
H6 2.183 0.908 0.348
H7 2.905 -0.669 0.078
H8 2.060 0.147 -1.235
C9 0.749 -0.726 0.250
H10 0.720 -1.731 -0.177
H11 0.760 -0.874 1.340
C12 -0.503 -0.002 -0.140
C13 -1.779 -0.777 -0.099
H14 -1.652 -1.777 -0.513
H15 -2.140 -0.900 0.930
H16 -2.570 -0.275 -0.654

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15 H16
C11.08821.08881.09743.04922.84014.09683.23432.58043.46982.99101.49372.55403.47062.96682.7417
H21.08821.76051.75592.70862.30163.76822.72172.82713.75853.36252.15613.46224.24233.98203.6558
H31.08881.76051.75794.02763.86365.09694.03083.49394.23803.95652.14072.69623.66773.15692.4328
H41.09741.75591.75793.46753.03914.40413.92172.95213.93832.95832.15173.01564.00203.03373.3077
C53.04922.70864.02763.46751.08751.08821.08841.52582.14882.14802.55273.89874.10234.41384.6508
H62.84012.30163.86363.03911.08751.75561.76082.17613.06262.48682.87794.32884.75994.72214.9993
H74.09683.76825.09694.40411.08821.75561.76192.16362.44282.49743.47994.68894.72695.12195.5379
H83.23432.72174.03083.92171.08841.76081.76192.16492.53793.06022.79154.10944.24314.84034.6855
C92.58042.82713.49392.95211.52582.17612.16362.16491.09231.10011.49822.55282.72982.97343.4695
H103.46983.75854.23803.93832.14883.06262.44282.53791.09231.74322.11782.67592.39603.17733.6291
H112.99103.36253.95652.95832.14802.48682.49743.06021.10011.74322.13222.91993.17272.92893.9272
C121.49372.15612.14072.15172.55272.87793.47992.79151.49822.11782.13221.49352.14662.15222.1470
C132.55403.46222.69623.01563.89874.32884.68894.10942.55282.67592.91991.49351.08931.09711.0893
H143.47064.24233.66774.00204.10234.75994.72694.24312.72982.39603.17272.14661.08931.75721.7660
H152.96683.98203.15693.03374.41384.72215.12194.84032.97343.17732.92892.15221.09711.75721.7568
H162.74173.65582.43283.30774.65084.99935.53794.68553.46953.62913.92722.14701.08931.76601.7568

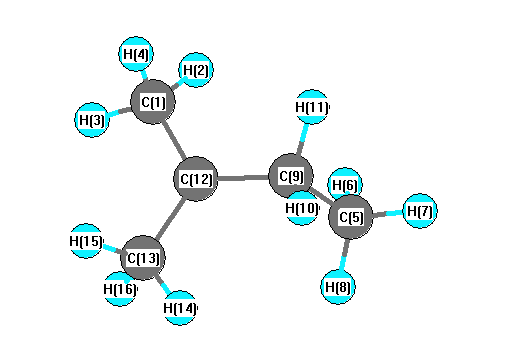
picture of 2-Methylbut-2-yl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 119.189 C1 C12 C13 117.517
H2 C1 H3 107.940 H2 C1 H4 106.916
H2 C1 C12 112.304 H3 C1 H4 107.051
H3 C1 C12 111.008 H4 C1 C12 111.367
C5 C9 H10 109.202 C5 C9 H11 108.687
C5 C9 C12 115.162 H6 C5 H7 107.597
H6 C5 H8 108.035 H6 C5 C9 111.657
H7 C5 H8 108.095 H7 C5 C9 110.614
H8 C5 C9 110.703 C9 C12 C13 117.140
H10 C9 H11 105.337 H10 C9 C12 108.664
H11 C9 C12 109.337 C12 C13 H14 111.463
C12 C13 H15 111.438 C12 C13 H16 111.495
H14 C13 H15 106.967 H14 C13 H16 108.313
H15 C13 H16 106.932
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability