Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -154.201060 |
| Energy at 298.15K | |
| HF Energy | -154.201060 |
| Nuclear repulsion energy | 74.854353 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3862 |
3686 |
10.13 |
|
|
|
| 2 |
A' |
3185 |
3040 |
9.05 |
|
|
|
| 3 |
A' |
2999 |
2862 |
68.02 |
|
|
|
| 4 |
A' |
1534 |
1464 |
1.15 |
|
|
|
| 5 |
A' |
1493 |
1425 |
5.65 |
|
|
|
| 6 |
A' |
1442 |
1376 |
6.41 |
|
|
|
| 7 |
A' |
1243 |
1186 |
58.55 |
|
|
|
| 8 |
A' |
1079 |
1030 |
104.37 |
|
|
|
| 9 |
A' |
1025 |
978 |
21.74 |
|
|
|
| 10 |
A' |
615 |
587 |
17.44 |
|
|
|
| 11 |
A' |
378 |
361 |
30.55 |
|
|
|
| 12 |
A" |
3296 |
3145 |
11.56 |
|
|
|
| 13 |
A" |
3035 |
2896 |
65.11 |
|
|
|
| 14 |
A" |
1291 |
1232 |
0.16 |
|
|
|
| 15 |
A" |
1178 |
1124 |
1.17 |
|
|
|
| 16 |
A" |
811 |
774 |
0.14 |
|
|
|
| 17 |
A" |
257 |
245 |
130.67 |
|
|
|
| 18 |
A" |
156i |
148i |
0.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14283.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 13630.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.112 |
-0.361 |
0.000 |
| C2 |
0.000 |
0.532 |
0.000 |
| C3 |
1.256 |
-0.260 |
0.000 |
| H4 |
-1.910 |
0.178 |
0.000 |
| H5 |
-0.025 |
1.185 |
0.887 |
| H6 |
-0.025 |
1.185 |
-0.887 |
| H7 |
1.661 |
-0.646 |
-0.929 |
| H8 |
1.661 |
-0.646 |
0.929 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4261 | 2.3704 | 0.9631 | 2.0874 | 2.0874 | 2.9385 | 2.9385 |
C2 | 1.4261 | | 1.4846 | 1.9426 | 1.1016 | 1.1016 | 2.2381 | 2.2381 | C3 | 2.3704 | 1.4846 | | 3.1964 | 2.1249 | 2.1249 | 1.0845 | 1.0845 | H4 | 0.9631 | 1.9426 | 3.1964 | | 2.3135 | 2.3135 | 3.7810 | 3.7810 | H5 | 2.0874 | 1.1016 | 2.1249 | 2.3135 | | 1.7735 | 3.0810 | 2.4896 | H6 | 2.0874 | 1.1016 | 2.1249 | 2.3135 | 1.7735 | | 2.4896 | 3.0810 | H7 | 2.9385 | 2.2381 | 1.0845 | 3.7810 | 3.0810 | 2.4896 | | 1.8578 | H8 | 2.9385 | 2.2381 | 1.0845 | 3.7810 | 2.4896 | 3.0810 | 1.8578 | |
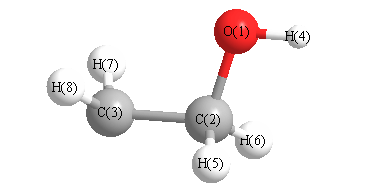 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.030 |
|
O1 |
C2 |
H5 |
110.684 |
| O1 |
C2 |
H6 |
110.684 |
|
C2 |
O1 |
H4 |
107.200 |
| C2 |
C3 |
H7 |
120.390 |
|
C2 |
C3 |
H8 |
120.390 |
| C3 |
C2 |
H5 |
109.604 |
|
C3 |
C2 |
H6 |
109.604 |
| H5 |
C2 |
H6 |
107.212 |
|
H7 |
C3 |
H8 |
117.856 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.542 |
|
|
|
| 2 |
C |
-0.020 |
|
|
|
| 3 |
C |
-0.251 |
|
|
|
| 4 |
H |
0.315 |
|
|
|
| 5 |
H |
0.111 |
|
|
|
| 6 |
H |
0.111 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.185 |
1.491 |
0.000 |
1.502 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.351 |
-2.787 |
0.000 |
| y |
-2.787 |
-19.731 |
0.000 |
| z |
0.000 |
0.000 |
-18.596 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.812 |
-2.787 |
0.000 |
| y |
-2.787 |
-2.257 |
0.000 |
| z |
0.000 |
0.000 |
-0.555 |
|
| Polar |
|---|
| 3z2-r2 | -1.110 |
|---|
| x2-y2 | 3.380 |
|---|
| xy | -2.787 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.234 |
-0.533 |
0.000 |
| y |
-0.533 |
3.236 |
0.000 |
| z |
0.000 |
0.000 |
3.446 |
<r2> (average value of r
2) Å
2
| <r2> |
50.029 |
| (<r2>)1/2 |
7.073 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -154.202905 |
| Energy at 298.15K | -154.207547 |
| HF Energy | -154.202905 |
| Nuclear repulsion energy | 75.197289 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3892 |
3714 |
19.65 |
|
|
|
| 2 |
A |
3315 |
3164 |
5.71 |
|
|
|
| 3 |
A |
3198 |
3052 |
4.99 |
|
|
|
| 4 |
A |
2969 |
2833 |
70.52 |
|
|
|
| 5 |
A |
2910 |
2777 |
72.60 |
|
|
|
| 6 |
A |
1511 |
1442 |
7.96 |
|
|
|
| 7 |
A |
1480 |
1412 |
5.94 |
|
|
|
| 8 |
A |
1428 |
1363 |
1.06 |
|
|
|
| 9 |
A |
1281 |
1222 |
73.99 |
|
|
|
| 10 |
A |
1230 |
1174 |
8.44 |
|
|
|
| 11 |
A |
1154 |
1101 |
50.19 |
|
|
|
| 12 |
A |
1079 |
1030 |
23.87 |
|
|
|
| 13 |
A |
977 |
932 |
11.07 |
|
|
|
| 14 |
A |
876 |
836 |
14.30 |
|
|
|
| 15 |
A |
454 |
433 |
25.41 |
|
|
|
| 16 |
A |
401 |
383 |
38.75 |
|
|
|
| 17 |
A |
278 |
265 |
98.86 |
|
|
|
| 18 |
A |
144 |
137 |
25.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14287.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 13634.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.102 |
-0.380 |
-0.058 |
| C2 |
-0.013 |
0.521 |
0.035 |
| C3 |
1.244 |
-0.253 |
-0.017 |
| H4 |
-1.902 |
0.125 |
0.108 |
| H5 |
-0.066 |
1.099 |
0.978 |
| H6 |
-0.032 |
1.264 |
-0.781 |
| H7 |
2.185 |
0.245 |
-0.220 |
| H8 |
1.241 |
-1.295 |
0.278 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4164 | 2.3495 | 0.9612 | 2.0826 | 2.0907 | 3.3492 | 2.5370 |
C2 | 1.4164 | | 1.4768 | 1.9314 | 1.1084 | 1.1040 | 2.2294 | 2.2198 | C3 | 2.3495 | 1.4768 | | 3.1711 | 2.1300 | 2.1248 | 1.0834 | 1.0831 | H4 | 0.9612 | 1.9314 | 3.1711 | | 2.2530 | 2.3630 | 4.1015 | 3.4529 | H5 | 2.0826 | 1.1084 | 2.1300 | 2.2530 | | 1.7675 | 2.6894 | 2.8165 | H6 | 2.0907 | 1.1040 | 2.1248 | 2.3630 | 1.7675 | | 2.5039 | 3.0484 | H7 | 3.3492 | 2.2294 | 1.0834 | 4.1015 | 2.6894 | 2.5039 | | 1.8735 | H8 | 2.5370 | 2.2198 | 1.0831 | 3.4529 | 2.8165 | 3.0484 | 1.8735 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.581 |
|
O1 |
C2 |
H5 |
110.558 |
| O1 |
C2 |
H6 |
111.498 |
|
C2 |
O1 |
H4 |
107.089 |
| C2 |
C3 |
H7 |
120.323 |
|
C2 |
C3 |
H8 |
119.467 |
| C3 |
C2 |
H5 |
110.147 |
|
C3 |
C2 |
H6 |
109.993 |
| H5 |
C2 |
H6 |
106.054 |
|
H7 |
C3 |
H8 |
119.710 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.546 |
|
|
|
| 2 |
C |
-0.008 |
|
|
|
| 3 |
C |
-0.268 |
|
|
|
| 4 |
H |
0.319 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
| 6 |
H |
0.115 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.319 |
1.635 |
0.448 |
1.725 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.612 |
-1.813 |
-0.559 |
| y |
-1.813 |
-18.948 |
-0.400 |
| z |
-0.559 |
-0.400 |
-20.029 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.876 |
-1.813 |
-0.559 |
| y |
-1.813 |
-1.128 |
-0.400 |
| z |
-0.559 |
-0.400 |
-2.749 |
|
| Polar |
|---|
| 3z2-r2 | -5.498 |
|---|
| x2-y2 | 3.336 |
|---|
| xy | -1.813 |
|---|
| xz | -0.559 |
|---|
| yz | -0.400 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.413 |
-0.197 |
-0.173 |
| y |
-0.197 |
3.782 |
-0.125 |
| z |
-0.173 |
-0.125 |
2.723 |
<r2> (average value of r
2) Å
2
| <r2> |
49.865 |
| (<r2>)1/2 |
7.061 |
