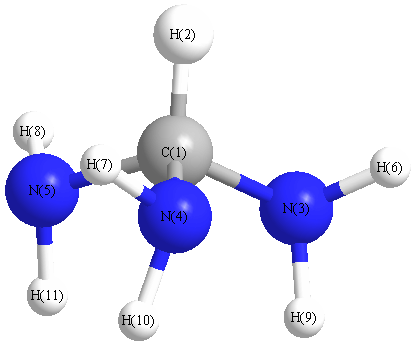
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3599 |
3435 |
0.24 |
|
|
|
| 2 |
A |
3498 |
3338 |
5.12 |
|
|
|
| 3 |
A |
3066 |
2926 |
38.18 |
|
|
|
| 4 |
A |
1654 |
1578 |
51.24 |
|
|
|
| 5 |
A |
1371 |
1309 |
1.67 |
|
|
|
| 6 |
A |
939 |
896 |
12.81 |
|
|
|
| 7 |
A |
884 |
844 |
0.54 |
|
|
|
| 8 |
A |
564 |
538 |
21.49 |
|
|
|
| 9 |
A |
316 |
301 |
74.10 |
|
|
|
| 10 |
E |
3600 |
3435 |
0.14 |
|
|
|
| 10 |
E |
3600 |
3435 |
0.14 |
|
|
|
| 11 |
E |
3502 |
3341 |
0.34 |
|
|
|
| 11 |
E |
3502 |
3341 |
0.34 |
|
|
|
| 12 |
E |
1667 |
1591 |
25.03 |
|
|
|
| 12 |
E |
1667 |
1591 |
25.03 |
|
|
|
| 13 |
E |
1426 |
1361 |
28.57 |
|
|
|
| 13 |
E |
1426 |
1361 |
28.58 |
|
|
|
| 14 |
E |
1236 |
1180 |
61.82 |
|
|
|
| 14 |
E |
1236 |
1180 |
61.83 |
|
|
|
| 15 |
E |
1062 |
1013 |
41.81 |
|
|
|
| 15 |
E |
1062 |
1013 |
41.81 |
|
|
|
| 16 |
E |
910 |
869 |
239.14 |
|
|
|
| 16 |
E |
910 |
869 |
239.14 |
|
|
|
| 17 |
E |
446 |
425 |
39.88 |
|
|
|
| 17 |
E |
446 |
425 |
39.88 |
|
|
|
| 18 |
E |
291 |
278 |
21.72 |
|
|
|
| 18 |
E |
291 |
278 |
21.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22085.0 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 21075.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.186 |
|
|
|
| 2 |
H |
0.142 |
|
|
|
| 3 |
N |
-0.607 |
|
|
|
| 4 |
N |
-0.607 |
|
|
|
| 5 |
N |
-0.607 |
|
|
|
| 6 |
H |
0.263 |
|
|
|
| 7 |
H |
0.263 |
|
|
|
| 8 |
H |
0.263 |
|
|
|
| 9 |
H |
0.234 |
|
|
|
| 10 |
H |
0.234 |
|
|
|
| 11 |
H |
0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.756 |
1.756 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.040 |
0.000 |
0.000 |
| y |
0.000 |
-27.040 |
0.000 |
| z |
0.000 |
0.000 |
-21.937 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.552 |
0.000 |
0.000 |
| y |
0.000 |
-2.552 |
0.000 |
| z |
0.000 |
0.000 |
5.103 |
|
| Polar |
|---|
| 3z2-r2 | 10.206 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.963 |
0.000 |
0.000 |
| y |
0.000 |
4.963 |
0.000 |
| z |
0.000 |
0.000 |
4.885 |
<r2> (average value of r
2) Å
2
| <r2> |
81.502 |
| (<r2>)1/2 |
9.028 |