return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH7N3 (triaminomethane)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-205.993299
Energy at 298.15K-206.003573
HF Energy-205.289997
Nuclear repulsion energy137.873189
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3647 3419 0.56      
2 A 3546 3325 5.02      
3 A 3140 2944 37.04      
4 A 1700 1594 48.80      
5 A 1409 1321 1.59      
6 A 985 924 11.19      
7 A 904 848 1.03      
8 A 566 531 23.82      
9 A 314 294 81.61      
10 E 3646 3419 0.08      
10 E 3646 3419 0.08      
11 E 3549 3327 0.00      
11 E 3549 3327 0.00      
12 E 1711 1605 24.24      
13 E 1471 1379 30.76      
14 E 1266 1187 61.18      
15 E 1094 1025 36.36      
16 E 962 902 221.31      
16 E 962 902 221.31      
17 E 454 426 39.89      
17 E 454 426 39.89      
18 E 300 281 20.13      
18 E 300 281 20.13      

Unscaled Zero Point Vibrational Energy (zpe) 22557.0 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 21149.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.29210 0.29210 0.16802

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.368
H2 0.000 0.000 1.460
N3 0.000 1.396 -0.056
N4 1.209 -0.698 -0.056
N5 -1.209 -0.698 -0.056
H6 0.871 1.822 0.243
H7 1.142 -1.666 0.243
H8 -2.013 -0.156 0.243
H9 0.008 1.430 -1.072
H10 1.235 -0.722 -1.072
H11 -1.242 -0.708 -1.072

Atom - Atom Distances (Å)
  C1 H2 N3 N4 N5 H6 H7 H8 H9 H10 H11
C11.09221.45871.45871.45872.02312.02312.02312.02972.02972.0297
H21.09222.06092.06092.06092.35752.35752.35752.90852.90852.9085
N31.45872.06092.41732.41731.01523.28082.55941.01692.65322.6462
N41.45872.06092.41732.41732.55941.01533.28082.64621.01692.6532
N51.45872.06092.41732.41733.28082.55941.01522.65322.64621.0169
H62.02312.35751.01522.55943.28083.49753.49751.62212.88643.5496
H72.02312.35753.28081.01532.55943.49753.49753.54961.62212.8864
H82.02312.35752.55943.28081.01523.49753.49752.88643.54961.6221
H92.02972.90851.01692.64622.65321.62213.54962.88642.47682.4768
H102.02972.90852.65321.01692.64622.88641.62213.54962.47682.4767
H112.02972.90852.64622.65321.01693.54962.88641.62212.47682.4767

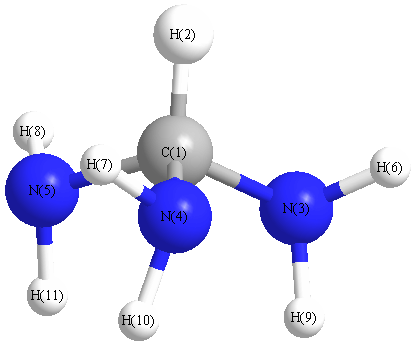
picture of triaminomethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 108.395 C1 N3 H9 108.841
C1 N4 H7 108.395 C1 N4 H10 108.841
C1 N5 H8 108.395 C1 N5 H11 108.841
H2 C1 N3 106.907 H2 C1 N4 106.907
H2 C1 N5 106.907 N3 C1 N4 111.909
N3 C1 N5 111.909 N4 C1 N5 111.909
H6 N3 H9 105.925 H7 N4 H10 105.925
H8 N5 H11 105.925
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability