Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -217.748580 |
| Energy at 298.15K | -217.756506 |
| HF Energy | -217.122840 |
| Nuclear repulsion energy | 128.292293 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3236 |
3024 |
22.25 |
|
|
|
| 2 |
A' |
3149 |
2943 |
27.64 |
|
|
|
| 3 |
A' |
3133 |
2927 |
22.62 |
|
|
|
| 4 |
A' |
3125 |
2920 |
27.13 |
|
|
|
| 5 |
A' |
1589 |
1485 |
1.08 |
|
|
|
| 6 |
A' |
1571 |
1468 |
5.06 |
|
|
|
| 7 |
A' |
1556 |
1454 |
0.27 |
|
|
|
| 8 |
A' |
1490 |
1392 |
19.83 |
|
|
|
| 9 |
A' |
1471 |
1375 |
2.10 |
|
|
|
| 10 |
A' |
1375 |
1285 |
2.20 |
|
|
|
| 11 |
A' |
1178 |
1101 |
1.75 |
|
|
|
| 12 |
A' |
1117 |
1044 |
71.73 |
|
|
|
| 13 |
A' |
1084 |
1013 |
18.31 |
|
|
|
| 14 |
A' |
933 |
872 |
7.99 |
|
|
|
| 15 |
A' |
460 |
430 |
5.86 |
|
|
|
| 16 |
A' |
273 |
255 |
3.47 |
|
|
|
| 17 |
A" |
3228 |
3016 |
57.43 |
|
|
|
| 18 |
A" |
3207 |
2997 |
10.73 |
|
|
|
| 19 |
A" |
3184 |
2975 |
18.98 |
|
|
|
| 20 |
A" |
1565 |
1462 |
6.44 |
|
|
|
| 21 |
A" |
1360 |
1271 |
0.21 |
|
|
|
| 22 |
A" |
1312 |
1226 |
0.14 |
|
|
|
| 23 |
A" |
1237 |
1155 |
1.76 |
|
|
|
| 24 |
A" |
924 |
863 |
1.84 |
|
|
|
| 25 |
A" |
788 |
736 |
1.13 |
|
|
|
| 26 |
A" |
245 |
229 |
0.01 |
|
|
|
| 27 |
A" |
140 |
131 |
2.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21964.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 20523.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.117 |
-0.782 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
-1.454 |
1.176 |
0.000 |
| F4 |
1.463 |
-1.152 |
0.000 |
| H5 |
-0.352 |
-1.213 |
0.886 |
| H6 |
-0.352 |
-1.213 |
-0.886 |
| H7 |
0.520 |
1.109 |
-0.876 |
| H8 |
0.520 |
1.109 |
0.876 |
| H9 |
-1.524 |
2.260 |
0.000 |
| H10 |
-1.980 |
0.808 |
-0.880 |
| H11 |
-1.980 |
0.808 |
0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5098 | 2.5105 | 1.3956 | 1.0910 | 1.0910 | 2.1227 | 2.1227 | 3.4571 | 2.7751 | 2.7751 |
C2 | 1.5098 | | 1.5229 | 2.3785 | 2.1576 | 2.1576 | 1.0894 | 1.0894 | 2.1648 | 2.1680 | 2.1680 | C3 | 2.5105 | 1.5229 | | 3.7320 | 2.7752 | 2.7752 | 2.1607 | 2.1607 | 1.0870 | 1.0889 | 1.0889 | F4 | 1.3956 | 2.3785 | 3.7320 | | 2.0208 | 2.0208 | 2.6019 | 2.6019 | 4.5353 | 4.0582 | 4.0582 | H5 | 1.0910 | 2.1576 | 2.7752 | 2.0208 | | 1.7714 | 3.0420 | 2.4798 | 3.7708 | 3.1384 | 2.5945 | H6 | 1.0910 | 2.1576 | 2.7752 | 2.0208 | 1.7714 | | 2.4798 | 3.0420 | 3.7708 | 2.5945 | 3.1384 | H7 | 2.1227 | 1.0894 | 2.1607 | 2.6019 | 3.0420 | 2.4798 | | 1.7527 | 2.5044 | 2.5173 | 3.0695 | H8 | 2.1227 | 1.0894 | 2.1607 | 2.6019 | 2.4798 | 3.0420 | 1.7527 | | 2.5044 | 3.0695 | 2.5173 | H9 | 3.4571 | 2.1648 | 1.0870 | 4.5353 | 3.7708 | 3.7708 | 2.5044 | 2.5044 | | 1.7579 | 1.7579 | H10 | 2.7751 | 2.1680 | 1.0889 | 4.0582 | 3.1384 | 2.5945 | 2.5173 | 3.0695 | 1.7579 | | 1.7602 | H11 | 2.7751 | 2.1680 | 1.0889 | 4.0582 | 2.5945 | 3.1384 | 3.0695 | 2.5173 | 1.7579 | 1.7602 | |
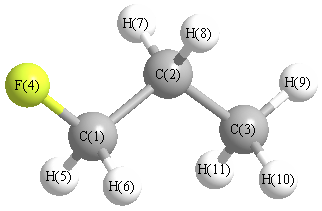 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.745 |
|
C1 |
C2 |
H7 |
108.415 |
| C1 |
C2 |
H8 |
108.415 |
|
C2 |
C1 |
F4 |
109.834 |
| C2 |
C1 |
H5 |
111.088 |
|
C2 |
C1 |
H6 |
111.088 |
| C2 |
C3 |
H9 |
110.987 |
|
C2 |
C3 |
H10 |
111.130 |
| C2 |
C3 |
H11 |
111.130 |
|
C3 |
C2 |
H7 |
110.507 |
| C3 |
C2 |
H8 |
110.507 |
|
F4 |
C1 |
H5 |
108.091 |
| F4 |
C1 |
H6 |
108.091 |
|
H5 |
C1 |
H6 |
108.547 |
| H7 |
C2 |
H8 |
107.100 |
|
H9 |
C3 |
H10 |
107.790 |
| H9 |
C3 |
H11 |
107.790 |
|
H10 |
C3 |
H11 |
107.861 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -217.749454 |
| Energy at 298.15K | |
| HF Energy | -217.123205 |
| Nuclear repulsion energy | 130.963505 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3250 |
3037 |
17.84 |
|
|
|
| 2 |
A |
3230 |
3018 |
34.89 |
|
|
|
| 3 |
A |
3201 |
2991 |
44.57 |
|
|
|
| 4 |
A |
3189 |
2980 |
17.97 |
|
|
|
| 5 |
A |
3138 |
2932 |
14.40 |
|
|
|
| 6 |
A |
3136 |
2930 |
27.78 |
|
|
|
| 7 |
A |
3128 |
2923 |
33.74 |
|
|
|
| 8 |
A |
1583 |
1479 |
0.34 |
|
|
|
| 9 |
A |
1573 |
1470 |
6.46 |
|
|
|
| 10 |
A |
1559 |
1457 |
5.82 |
|
|
|
| 11 |
A |
1538 |
1438 |
1.85 |
|
|
|
| 12 |
A |
1482 |
1385 |
14.94 |
|
|
|
| 13 |
A |
1467 |
1371 |
6.54 |
|
|
|
| 14 |
A |
1427 |
1333 |
1.01 |
|
|
|
| 15 |
A |
1346 |
1257 |
0.78 |
|
|
|
| 16 |
A |
1311 |
1225 |
0.72 |
|
|
|
| 17 |
A |
1221 |
1141 |
2.05 |
|
|
|
| 18 |
A |
1163 |
1087 |
11.06 |
|
|
|
| 19 |
A |
1135 |
1060 |
39.29 |
|
|
|
| 20 |
A |
1018 |
951 |
36.13 |
|
|
|
| 21 |
A |
952 |
890 |
2.91 |
|
|
|
| 22 |
A |
914 |
854 |
3.94 |
|
|
|
| 23 |
A |
795 |
743 |
0.40 |
|
|
|
| 24 |
A |
493 |
461 |
3.55 |
|
|
|
| 25 |
A |
327 |
306 |
1.14 |
|
|
|
| 26 |
A |
239 |
223 |
2.25 |
|
|
|
| 27 |
A |
152 |
142 |
2.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21983.7 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 20541.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.795 |
0.524 |
0.291 |
| C2 |
-0.595 |
0.665 |
-0.279 |
| C3 |
-1.501 |
-0.490 |
0.120 |
| F4 |
1.366 |
-0.667 |
-0.169 |
| H5 |
1.443 |
1.345 |
-0.016 |
| H6 |
0.771 |
0.481 |
1.381 |
| H7 |
-0.519 |
0.726 |
-1.365 |
| H8 |
-1.008 |
1.615 |
0.065 |
| H9 |
-2.491 |
-0.385 |
-0.318 |
| H10 |
-1.075 |
-1.432 |
-0.215 |
| H11 |
-1.614 |
-0.537 |
1.202 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5099 | 2.5160 | 1.3985 | 1.0899 | 1.0911 | 2.1236 | 2.1205 | 3.4634 | 2.7526 | 2.7861 |
C2 | 1.5099 | | 1.5215 | 2.3744 | 2.1647 | 2.1582 | 1.0900 | 1.0915 | 2.1673 | 2.1521 | 2.1631 | C3 | 2.5160 | 1.5215 | | 2.8872 | 3.4717 | 2.7741 | 2.1560 | 2.1630 | 1.0874 | 1.0863 | 1.0891 | F4 | 1.3985 | 2.3744 | 2.8872 | | 2.0196 | 2.0183 | 2.6321 | 3.3023 | 3.8703 | 2.5584 | 3.2832 | H5 | 1.0899 | 2.1647 | 3.4717 | 2.0196 | | 1.7745 | 2.4599 | 2.4677 | 4.3079 | 3.7533 | 3.7912 | H6 | 1.0911 | 2.1582 | 2.7741 | 2.0183 | 1.7745 | | 3.0435 | 2.4869 | 3.7783 | 3.1005 | 2.5999 | H7 | 2.1236 | 1.0900 | 2.1560 | 2.6321 | 2.4599 | 3.0435 | | 1.7536 | 2.4938 | 2.5073 | 3.0635 | H8 | 2.1205 | 1.0915 | 2.1630 | 3.3023 | 2.4677 | 2.4869 | 1.7536 | | 2.5190 | 3.0604 | 2.5085 | H9 | 3.4634 | 2.1673 | 1.0874 | 3.8703 | 4.3079 | 3.7783 | 2.4938 | 2.5190 | | 1.7636 | 1.7611 | H10 | 2.7526 | 2.1521 | 1.0863 | 2.5584 | 3.7533 | 3.1005 | 2.5073 | 3.0604 | 1.7636 | | 1.7605 | H11 | 2.7861 | 2.1631 | 1.0891 | 3.2832 | 3.7912 | 2.5999 | 3.0635 | 2.5085 | 1.7611 | 1.7605 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.199 |
|
C1 |
C2 |
H7 |
108.460 |
| C1 |
C2 |
H8 |
108.131 |
|
C2 |
C1 |
F4 |
109.393 |
| C2 |
C1 |
H5 |
111.730 |
|
C2 |
C1 |
H6 |
111.127 |
| C2 |
C3 |
H9 |
111.265 |
|
C2 |
C3 |
H10 |
110.111 |
| C2 |
C3 |
H11 |
110.819 |
|
C3 |
C2 |
H7 |
110.206 |
| C3 |
C2 |
H8 |
110.670 |
|
F4 |
C1 |
H5 |
107.862 |
| F4 |
C1 |
H6 |
107.690 |
|
H5 |
C1 |
H6 |
108.902 |
| H7 |
C2 |
H8 |
106.999 |
|
H9 |
C3 |
H10 |
108.459 |
| H9 |
C3 |
H11 |
108.027 |
|
H10 |
C3 |
H11 |
108.057 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
