Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -218.123496 |
| Energy at 298.15K | -218.131355 |
| HF Energy | -218.123496 |
| Nuclear repulsion energy | 128.231691 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3011 |
24.57 |
|
|
|
| 2 |
A' |
3081 |
2936 |
36.00 |
|
|
|
| 3 |
A' |
3070 |
2926 |
20.72 |
|
|
|
| 4 |
A' |
3055 |
2912 |
32.53 |
|
|
|
| 5 |
A' |
1553 |
1480 |
2.37 |
|
|
|
| 6 |
A' |
1535 |
1463 |
6.19 |
|
|
|
| 7 |
A' |
1521 |
1449 |
0.39 |
|
|
|
| 8 |
A' |
1460 |
1392 |
20.26 |
|
|
|
| 9 |
A' |
1440 |
1373 |
3.31 |
|
|
|
| 10 |
A' |
1348 |
1285 |
1.46 |
|
|
|
| 11 |
A' |
1159 |
1105 |
6.15 |
|
|
|
| 12 |
A' |
1117 |
1064 |
93.21 |
|
|
|
| 13 |
A' |
1070 |
1020 |
2.57 |
|
|
|
| 14 |
A' |
917 |
874 |
9.48 |
|
|
|
| 15 |
A' |
456 |
434 |
5.84 |
|
|
|
| 16 |
A' |
266 |
253 |
2.86 |
|
|
|
| 17 |
A" |
3149 |
3001 |
64.58 |
|
|
|
| 18 |
A" |
3125 |
2978 |
10.22 |
|
|
|
| 19 |
A" |
3097 |
2952 |
22.71 |
|
|
|
| 20 |
A" |
1529 |
1458 |
8.76 |
|
|
|
| 21 |
A" |
1332 |
1270 |
0.24 |
|
|
|
| 22 |
A" |
1285 |
1225 |
0.31 |
|
|
|
| 23 |
A" |
1210 |
1153 |
1.16 |
|
|
|
| 24 |
A" |
904 |
861 |
2.16 |
|
|
|
| 25 |
A" |
773 |
737 |
2.41 |
|
|
|
| 26 |
A" |
238 |
227 |
0.00 |
|
|
|
| 27 |
A" |
132 |
126 |
2.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21490.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20482.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.126 |
-0.786 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
-1.455 |
1.176 |
0.000 |
| F4 |
1.459 |
-1.151 |
0.000 |
| H5 |
-0.350 |
-1.220 |
0.890 |
| H6 |
-0.350 |
-1.220 |
-0.890 |
| H7 |
0.523 |
1.115 |
-0.880 |
| H8 |
0.523 |
1.115 |
0.880 |
| H9 |
-1.525 |
2.268 |
0.000 |
| H10 |
-1.988 |
0.810 |
-0.885 |
| H11 |
-1.988 |
0.810 |
0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5134 | 2.5194 | 1.3816 | 1.0984 | 1.0984 | 2.1321 | 2.1321 | 3.4714 | 2.7926 | 2.7926 |
C2 | 1.5134 | | 1.5239 | 2.3742 | 2.1650 | 2.1650 | 1.0968 | 1.0968 | 2.1712 | 2.1781 | 2.1781 | C3 | 2.5194 | 1.5239 | | 3.7286 | 2.7847 | 2.7847 | 2.1658 | 2.1658 | 1.0943 | 1.0964 | 1.0964 | F4 | 1.3816 | 2.3742 | 3.7286 | | 2.0163 | 2.0163 | 2.6050 | 2.6050 | 4.5376 | 4.0629 | 4.0629 | H5 | 1.0984 | 2.1650 | 2.7847 | 2.0163 | | 1.7793 | 3.0575 | 2.4931 | 3.7868 | 3.1551 | 2.6086 | H6 | 1.0984 | 2.1650 | 2.7847 | 2.0163 | 1.7793 | | 2.4931 | 3.0575 | 3.7868 | 2.6086 | 3.1551 | H7 | 2.1321 | 1.0968 | 2.1658 | 2.6050 | 3.0575 | 2.4931 | | 1.7608 | 2.5095 | 2.5298 | 3.0849 | H8 | 2.1321 | 1.0968 | 2.1658 | 2.6050 | 2.4931 | 3.0575 | 1.7608 | | 2.5095 | 3.0849 | 2.5298 | H9 | 3.4714 | 2.1712 | 1.0943 | 4.5376 | 3.7868 | 3.7868 | 2.5095 | 2.5095 | | 1.7678 | 1.7678 | H10 | 2.7926 | 2.1781 | 1.0964 | 4.0629 | 3.1551 | 2.6086 | 2.5298 | 3.0849 | 1.7678 | | 1.7702 | H11 | 2.7926 | 2.1781 | 1.0964 | 4.0629 | 2.6086 | 3.1551 | 3.0849 | 2.5298 | 1.7678 | 1.7702 | |
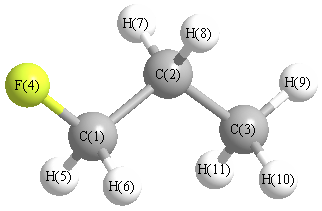 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.093 |
|
C1 |
C2 |
H7 |
108.491 |
| C1 |
C2 |
H8 |
108.491 |
|
C2 |
C1 |
F4 |
110.110 |
| C2 |
C1 |
H5 |
110.985 |
|
C2 |
C1 |
H6 |
110.985 |
| C2 |
C3 |
H9 |
110.980 |
|
C2 |
C3 |
H10 |
111.405 |
| C2 |
C3 |
H11 |
111.405 |
|
C3 |
C2 |
H7 |
110.407 |
| C3 |
C2 |
H8 |
110.407 |
|
F4 |
C1 |
H5 |
108.242 |
| F4 |
C1 |
H6 |
108.242 |
|
H5 |
C1 |
H6 |
108.181 |
| H7 |
C2 |
H8 |
106.776 |
|
H9 |
C3 |
H10 |
107.601 |
| H9 |
C3 |
H11 |
107.601 |
|
H10 |
C3 |
H11 |
107.662 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.021 |
|
|
|
| 2 |
C |
-0.364 |
|
|
|
| 3 |
C |
-0.538 |
|
|
|
| 4 |
F |
-0.319 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.182 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.594 |
0.727 |
0.000 |
1.752 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.584 |
1.983 |
0.000 |
| y |
1.983 |
-25.504 |
0.000 |
| z |
0.000 |
0.000 |
-24.366 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.649 |
1.983 |
0.000 |
| y |
1.983 |
0.471 |
0.000 |
| z |
0.000 |
0.000 |
2.178 |
|
| Polar |
|---|
| 3z2-r2 | 4.357 |
|---|
| x2-y2 | -2.080 |
|---|
| xy | 1.983 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.069 |
-0.221 |
0.000 |
| y |
-0.221 |
5.075 |
0.000 |
| z |
0.000 |
0.000 |
4.725 |
<r2> (average value of r
2) Å
2
| <r2> |
102.764 |
| (<r2>)1/2 |
10.137 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -218.124052 |
| Energy at 298.15K | |
| HF Energy | -218.124052 |
| Nuclear repulsion energy | 130.770800 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3022 |
20.94 |
|
|
|
| 2 |
A |
3152 |
3005 |
39.62 |
|
|
|
| 3 |
A |
3120 |
2974 |
42.89 |
|
|
|
| 4 |
A |
3109 |
2963 |
32.49 |
|
|
|
| 5 |
A |
3075 |
2931 |
19.66 |
|
|
|
| 6 |
A |
3072 |
2928 |
18.16 |
|
|
|
| 7 |
A |
3060 |
2917 |
41.30 |
|
|
|
| 8 |
A |
1547 |
1474 |
0.80 |
|
|
|
| 9 |
A |
1537 |
1465 |
9.24 |
|
|
|
| 10 |
A |
1523 |
1452 |
7.99 |
|
|
|
| 11 |
A |
1504 |
1434 |
2.57 |
|
|
|
| 12 |
A |
1455 |
1387 |
16.33 |
|
|
|
| 13 |
A |
1436 |
1369 |
6.61 |
|
|
|
| 14 |
A |
1399 |
1333 |
0.70 |
|
|
|
| 15 |
A |
1321 |
1259 |
1.26 |
|
|
|
| 16 |
A |
1290 |
1230 |
0.93 |
|
|
|
| 17 |
A |
1195 |
1139 |
1.43 |
|
|
|
| 18 |
A |
1149 |
1095 |
27.87 |
|
|
|
| 19 |
A |
1123 |
1070 |
36.61 |
|
|
|
| 20 |
A |
1010 |
962 |
33.71 |
|
|
|
| 21 |
A |
933 |
889 |
3.22 |
|
|
|
| 22 |
A |
899 |
857 |
3.69 |
|
|
|
| 23 |
A |
781 |
744 |
0.72 |
|
|
|
| 24 |
A |
485 |
462 |
3.05 |
|
|
|
| 25 |
A |
318 |
303 |
1.16 |
|
|
|
| 26 |
A |
225 |
214 |
1.82 |
|
|
|
| 27 |
A |
148 |
141 |
2.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21516.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20507.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.802 |
0.512 |
0.290 |
| C2 |
-0.593 |
0.664 |
-0.276 |
| C3 |
-1.513 |
-0.484 |
0.118 |
| F4 |
1.373 |
-0.663 |
-0.170 |
| H5 |
1.449 |
1.345 |
-0.011 |
| H6 |
0.778 |
0.468 |
1.388 |
| H7 |
-0.519 |
0.730 |
-1.369 |
| H8 |
-1.000 |
1.623 |
0.071 |
| H9 |
-2.505 |
-0.373 |
-0.332 |
| H10 |
-1.094 |
-1.439 |
-0.213 |
| H11 |
-1.641 |
-0.533 |
1.206 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5129 | 2.5260 | 1.3847 | 1.0973 | 1.0984 | 2.1321 | 2.1280 | 3.4790 | 2.7659 | 2.8104 |
C2 | 1.5129 | | 1.5230 | 2.3738 | 2.1686 | 2.1646 | 1.0973 | 1.0982 | 2.1755 | 2.1622 | 2.1743 | C3 | 2.5260 | 1.5230 | | 2.9060 | 3.4836 | 2.7869 | 2.1615 | 2.1692 | 1.0946 | 1.0940 | 1.0966 | F4 | 1.3847 | 2.3738 | 2.9060 | | 2.0159 | 2.0144 | 2.6376 | 3.3033 | 3.8920 | 2.5861 | 3.3160 | H5 | 1.0973 | 2.1686 | 3.4836 | 2.0159 | | 1.7821 | 2.4690 | 2.4653 | 4.3226 | 3.7756 | 3.8152 | H6 | 1.0984 | 2.1646 | 2.7869 | 2.0144 | 1.7821 | | 3.0577 | 2.4957 | 3.7999 | 3.1140 | 2.6242 | H7 | 2.1321 | 1.0973 | 2.1615 | 2.6376 | 2.4690 | 3.0577 | | 1.7611 | 2.4967 | 2.5238 | 3.0795 | H8 | 2.1280 | 1.0982 | 2.1692 | 3.3033 | 2.4653 | 2.4957 | 1.7611 | | 2.5319 | 3.0762 | 2.5195 | H9 | 3.4790 | 2.1755 | 1.0946 | 3.8920 | 4.3226 | 3.7999 | 2.4967 | 2.5319 | | 1.7726 | 1.7710 | H10 | 2.7659 | 2.1622 | 1.0940 | 2.5861 | 3.7756 | 3.1140 | 2.5238 | 3.0762 | 1.7726 | | 1.7699 | H11 | 2.8104 | 2.1743 | 1.0966 | 3.3160 | 3.8152 | 2.6242 | 3.0795 | 2.5195 | 1.7710 | 1.7699 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.614 |
|
C1 |
C2 |
H7 |
108.485 |
| C1 |
C2 |
H8 |
108.114 |
|
C2 |
C1 |
F4 |
109.934 |
| C2 |
C1 |
H5 |
111.367 |
|
C2 |
C1 |
H6 |
110.976 |
| C2 |
C3 |
H9 |
111.379 |
|
C2 |
C3 |
H10 |
110.350 |
| C2 |
C3 |
H11 |
111.154 |
|
C3 |
C2 |
H7 |
110.096 |
| C3 |
C2 |
H8 |
110.650 |
|
F4 |
C1 |
H5 |
108.062 |
| F4 |
C1 |
H6 |
107.875 |
|
H5 |
C1 |
H6 |
108.512 |
| H7 |
C2 |
H8 |
106.671 |
|
H9 |
C3 |
H10 |
108.178 |
| H9 |
C3 |
H11 |
107.847 |
|
H10 |
C3 |
H11 |
107.796 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.012 |
|
|
|
| 2 |
C |
-0.363 |
|
|
|
| 3 |
C |
-0.525 |
|
|
|
| 4 |
F |
-0.318 |
|
|
|
| 5 |
H |
0.157 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
| 7 |
H |
0.181 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.192 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.850 |
1.335 |
0.411 |
1.635 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.400 |
1.562 |
0.573 |
| y |
1.562 |
-25.013 |
0.056 |
| z |
0.573 |
0.056 |
-24.478 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.655 |
1.562 |
0.573 |
| y |
1.562 |
0.426 |
0.056 |
| z |
0.573 |
0.056 |
1.229 |
|
| Polar |
|---|
| 3z2-r2 | 2.457 |
|---|
| x2-y2 | -1.387 |
|---|
| xy | 1.562 |
|---|
| xz | 0.573 |
|---|
| yz | 0.056 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.224 |
0.227 |
0.036 |
| y |
0.227 |
4.875 |
-0.025 |
| z |
0.036 |
-0.025 |
4.774 |
<r2> (average value of r
2) Å
2
| <r2> |
90.588 |
| (<r2>)1/2 |
9.518 |
