return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-330.549189
Energy at 298.15K-330.554608
HF Energy-329.562424
Nuclear repulsion energy269.163764
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3297 3081 0.11      
2 A1 3286 3070 14.80      
3 A1 3264 3050 0.13      
4 A1 1685 1575 44.67      
5 A1 1562 1460 75.62      
6 A1 1292 1207 79.85      
7 A1 1202 1124 7.18      
8 A1 1058 989 3.59      
9 A1 1028 961 0.24      
10 A1 833 778 20.00      
11 A1 524 490 3.38      
12 A2 921 860 0.00      
13 A2 831 776 0.00      
14 A2 410 383 0.00      
15 B1 919 859 0.49      
16 B1 871 814 3.23      
17 B1 753 704 85.63      
18 B1 533 498 0.80      
19 B1 480 448 4.07      
20 B1 238 222 0.08      
21 B2 3296 3080 4.24      
22 B2 3273 3058 9.03      
23 B2 1685 1574 6.25      
24 B2 1516 1417 1.36      
25 B2 1472 1376 0.57      
26 B2 1348 1260 0.22      
27 B2 1207 1128 0.20      
28 B2 1112 1039 6.41      
29 B2 627 586 0.07      
30 B2 404 378 2.38      

Unscaled Zero Point Vibrational Energy (zpe) 20463.2 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 19120.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.18950 0.08539 0.05887

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.284
C2 0.000 0.000 0.928
C3 0.000 1.216 0.259
C4 0.000 -1.216 0.259
C5 0.000 1.207 -1.135
C6 0.000 -1.207 -1.135
C7 0.000 0.000 -1.834
H8 0.000 2.136 0.826
H9 0.000 -2.136 0.826
H10 0.000 2.146 -1.672
H11 0.000 -2.146 -1.672
H12 0.000 0.000 -2.915

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.35612.36212.36213.62603.62604.11792.58592.58594.50084.50085.1987
C21.35611.38801.38802.39032.39032.76182.13802.13803.37133.37133.8426
C32.36211.38802.43241.39432.79602.42071.08013.39942.14363.87733.3989
C42.36211.38802.43242.79601.39432.42073.39941.08013.87732.14363.3989
C53.62602.39031.39432.79602.41471.39502.16963.87571.08133.39582.1505
C63.62602.39032.79601.39432.41471.39503.87572.16963.39581.08132.1505
C74.11792.76182.42072.42071.39501.39503.41103.41102.15172.15171.0809
H82.58592.13801.08013.39942.16963.87573.41104.27122.49844.95694.3073
H92.58592.13803.39941.08013.87572.16963.41104.27124.95692.49844.3073
H104.50083.37132.14363.87731.08133.39582.15172.49844.95694.29142.4794
H114.50083.37133.87732.14363.39581.08132.15174.95692.49844.29142.4794
H125.19873.84263.39893.39892.15052.15051.08094.30734.30732.47942.4794

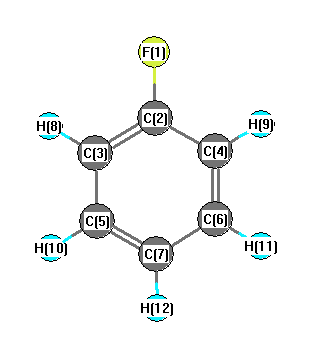
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 118.806 F1 C2 C4 118.806
C2 C3 C5 118.442 C2 C3 H8 119.540
C2 C4 C6 118.442 C2 C4 H9 119.540
C3 C2 C4 122.389 C3 C5 C7 120.424
C3 C5 H10 119.434 C4 C6 C7 120.424
C4 C6 H11 119.434 C5 C3 H8 122.018
C5 C7 C6 119.880 C5 C7 H12 120.060
C6 C4 H9 122.018 C6 C7 H12 120.060
C7 C5 H10 120.142 C7 C6 H11 120.142
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability