Jump to
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -245.141224 |
| Energy at 298.15K | |
| HF Energy | -245.141224 |
| Nuclear repulsion energy | 123.027030 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3776 |
3620 |
74.55 |
|
|
|
| 2 |
A' |
3775 |
3619 |
48.63 |
|
|
|
| 3 |
A' |
3630 |
3480 |
47.72 |
|
|
|
| 4 |
A' |
1854 |
1777 |
461.46 |
|
|
|
| 5 |
A' |
1626 |
1559 |
128.47 |
|
|
|
| 6 |
A' |
1441 |
1382 |
131.13 |
|
|
|
| 7 |
A' |
1227 |
1177 |
197.39 |
|
|
|
| 8 |
A' |
1072 |
1027 |
37.58 |
|
|
|
| 9 |
A' |
955 |
916 |
40.34 |
|
|
|
| 10 |
A' |
569 |
546 |
36.07 |
|
|
|
| 11 |
A' |
473 |
453 |
7.39 |
|
|
|
| 12 |
A" |
762 |
731 |
30.91 |
|
|
|
| 13 |
A" |
596 |
572 |
62.43 |
|
|
|
| 14 |
A" |
479 |
459 |
69.53 |
|
|
|
| 15 |
A" |
216i |
207i |
259.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11009.4 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 10554.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.130 |
0.000 |
| O2 |
-0.071 |
1.348 |
0.000 |
| N3 |
1.144 |
-0.602 |
0.000 |
| O4 |
-1.092 |
-0.692 |
0.000 |
| H5 |
2.023 |
-0.113 |
0.000 |
| H6 |
1.123 |
-1.609 |
0.000 |
| H7 |
-1.852 |
-0.088 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2194 | 1.3587 | 1.3671 | 2.0373 | 2.0702 | 1.8647 |
O2 | 1.2194 | | 2.2978 | 2.2813 | 2.5531 | 3.1885 | 2.2874 | N3 | 1.3587 | 2.2978 | | 2.2376 | 1.0056 | 1.0066 | 3.0398 | O4 | 1.3671 | 2.2813 | 2.2376 | | 3.1677 | 2.3966 | 0.9713 | H5 | 2.0373 | 2.5531 | 1.0056 | 3.1677 | | 1.7454 | 3.8746 | H6 | 2.0702 | 3.1885 | 1.0066 | 2.3966 | 1.7454 | | 3.3410 | H7 | 1.8647 | 2.2874 | 3.0398 | 0.9713 | 3.8746 | 3.3410 | |
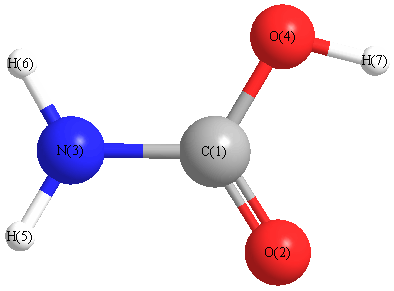 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.251 |
|
C1 |
N3 |
H6 |
121.432 |
| C1 |
O4 |
H7 |
104.492 |
|
O2 |
C1 |
N3 |
125.982 |
| O2 |
C1 |
O4 |
123.667 |
|
N3 |
C1 |
O4 |
110.350 |
| H5 |
N3 |
H6 |
120.317 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.725 |
|
|
|
| 2 |
O |
-0.508 |
|
|
|
| 3 |
N |
-0.647 |
|
|
|
| 4 |
O |
-0.519 |
|
|
|
| 5 |
H |
0.306 |
|
|
|
| 6 |
H |
0.303 |
|
|
|
| 7 |
H |
0.341 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.796 |
-2.241 |
0.000 |
2.378 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.238 |
-0.333 |
0.000 |
| y |
-0.333 |
4.097 |
0.000 |
| z |
0.000 |
0.000 |
1.731 |
<r2> (average value of r
2) Å
2
| <r2> |
64.419 |
| (<r2>)1/2 |
8.026 |
Jump to
S1C1
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -245.141326 |
| Energy at 298.15K | -245.146041 |
| HF Energy | -245.141326 |
| Nuclear repulsion energy | 122.983489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3779 |
3623 |
63.80 |
|
|
|
| 2 |
A |
3749 |
3594 |
50.59 |
|
|
|
| 3 |
A |
3611 |
3461 |
39.59 |
|
|
|
| 4 |
A |
1855 |
1778 |
445.82 |
|
|
|
| 5 |
A |
1631 |
1563 |
115.73 |
|
|
|
| 6 |
A |
1439 |
1380 |
133.51 |
|
|
|
| 7 |
A |
1231 |
1180 |
173.83 |
|
|
|
| 8 |
A |
1081 |
1036 |
61.20 |
|
|
|
| 9 |
A |
956 |
917 |
39.36 |
|
|
|
| 10 |
A |
762 |
730 |
40.50 |
|
|
|
| 11 |
A |
590 |
566 |
82.56 |
|
|
|
| 12 |
A |
569 |
545 |
38.44 |
|
|
|
| 13 |
A |
491 |
470 |
32.20 |
|
|
|
| 14 |
A |
458 |
439 |
22.41 |
|
|
|
| 15 |
A |
300 |
288 |
273.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11249.7 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 10785.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.124 |
-0.001 |
| O2 |
-0.479 |
1.261 |
0.006 |
| N3 |
1.277 |
-0.225 |
-0.048 |
| O4 |
-0.828 |
-0.992 |
0.003 |
| H5 |
1.945 |
0.508 |
0.126 |
| H6 |
1.546 |
-1.177 |
0.148 |
| H7 |
-1.738 |
-0.651 |
0.003 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2187 | 1.3635 | 1.3668 | 2.0254 | 2.0570 | 1.8664 |
O2 | 1.2187 | | 2.3013 | 2.2805 | 2.5404 | 3.1726 | 2.2897 | N3 | 1.3635 | 2.3013 | | 2.2412 | 1.0072 | 1.0082 | 3.0450 | O4 | 1.3668 | 2.2805 | 2.2412 | | 3.1552 | 2.3859 | 0.9712 | H5 | 2.0254 | 2.5404 | 1.0072 | 3.1552 | | 1.7319 | 3.8624 | H6 | 2.0570 | 3.1726 | 1.0082 | 2.3859 | 1.7319 | | 3.3286 | H7 | 1.8664 | 2.2897 | 3.0450 | 0.9712 | 3.8624 | 3.3286 | |
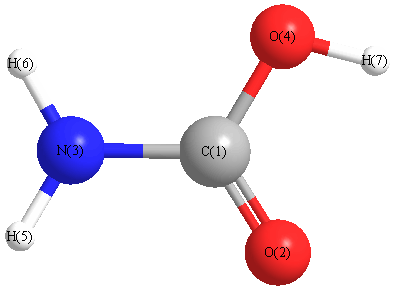 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.568 |
|
C1 |
N3 |
H6 |
119.539 |
| C1 |
O4 |
H7 |
104.658 |
|
O2 |
C1 |
N3 |
125.957 |
| O2 |
C1 |
O4 |
123.681 |
|
N3 |
C1 |
O4 |
110.340 |
| H5 |
N3 |
H6 |
118.491 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.716 |
|
|
|
| 2 |
O |
-0.504 |
|
|
|
| 3 |
N |
-0.640 |
|
|
|
| 4 |
O |
-0.516 |
|
|
|
| 5 |
H |
0.303 |
|
|
|
| 6 |
H |
0.300 |
|
|
|
| 7 |
H |
0.341 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.308 |
-1.838 |
0.618 |
2.339 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.420 |
-0.221 |
0.012 |
| y |
-0.221 |
3.935 |
-0.003 |
| z |
0.012 |
-0.003 |
1.763 |
<r2> (average value of r
2) Å
2
| <r2> |
64.454 |
| (<r2>)1/2 |
8.028 |
