Jump to
S1C2
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -357.534775 |
| Energy at 298.15K | -357.539698 |
| HF Energy | -356.554382 |
| Nuclear repulsion energy | 231.794926 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3843 |
3618 |
95.01 |
|
|
|
| 2 |
A' |
3842 |
3617 |
58.33 |
|
|
|
| 3 |
A' |
3698 |
3481 |
74.74 |
|
|
|
| 4 |
A' |
1869 |
1760 |
9.17 |
|
|
|
| 5 |
A' |
1838 |
1731 |
535.81 |
|
|
|
| 6 |
A' |
1659 |
1561 |
112.89 |
|
|
|
| 7 |
A' |
1481 |
1394 |
13.81 |
|
|
|
| 8 |
A' |
1358 |
1279 |
64.44 |
|
|
|
| 9 |
A' |
1221 |
1149 |
301.39 |
|
|
|
| 10 |
A' |
1124 |
1058 |
3.26 |
|
|
|
| 11 |
A' |
799 |
752 |
6.48 |
|
|
|
| 12 |
A' |
614 |
578 |
79.12 |
|
|
|
| 13 |
A' |
538 |
507 |
0.20 |
|
|
|
| 14 |
A' |
423 |
398 |
4.07 |
|
|
|
| 15 |
A' |
276 |
260 |
13.74 |
|
|
|
| 16 |
A" |
806 |
758 |
7.13 |
|
|
|
| 17 |
A" |
679 |
640 |
153.83 |
|
|
|
| 18 |
A" |
632 |
595 |
12.64 |
|
|
|
| 19 |
A" |
430 |
404 |
3.86 |
|
|
|
| 20 |
A" |
264 |
248 |
271.31 |
|
|
|
| 21 |
A" |
44 |
42 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13718.5 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 12914.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.753 |
0.000 |
| C2 |
-0.052 |
-0.786 |
0.000 |
| O3 |
-1.101 |
-1.410 |
0.000 |
| O4 |
1.040 |
1.383 |
0.000 |
| O5 |
-1.222 |
1.297 |
0.000 |
| N6 |
1.195 |
-1.314 |
0.000 |
| H7 |
1.300 |
-2.312 |
0.000 |
| H8 |
1.999 |
-0.711 |
0.000 |
| H9 |
-1.092 |
2.258 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5398 | 2.4266 | 1.2168 | 1.3384 | 2.3872 | 3.3289 | 2.4776 | 1.8594 |
C2 | 1.5398 | | 1.2200 | 2.4293 | 2.3900 | 1.3546 | 2.0386 | 2.0525 | 3.2166 | O3 | 2.4266 | 1.2200 | | 3.5197 | 2.7103 | 2.2983 | 2.5644 | 3.1775 | 3.6678 | O4 | 1.2168 | 2.4293 | 3.5197 | | 2.2645 | 2.7015 | 3.7045 | 2.3035 | 2.3045 | O5 | 1.3384 | 2.3900 | 2.7103 | 2.2645 | | 3.5588 | 4.4035 | 3.7964 | 0.9690 | N6 | 2.3872 | 1.3546 | 2.2983 | 2.7015 | 3.5588 | | 1.0038 | 1.0042 | 4.2410 | H7 | 3.3289 | 2.0386 | 2.5644 | 3.7045 | 4.4035 | 1.0038 | | 1.7468 | 5.1577 | H8 | 2.4776 | 2.0525 | 3.1775 | 2.3035 | 3.7964 | 1.0042 | 1.7468 | | 4.2857 | H9 | 1.8594 | 3.2166 | 3.6678 | 2.3045 | 0.9690 | 4.2410 | 5.1577 | 4.2857 | |
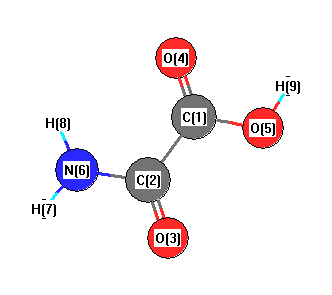 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.688 |
|
C1 |
C2 |
N6 |
110.967 |
| C1 |
O5 |
H9 |
106.275 |
|
C2 |
C1 |
O4 |
123.170 |
| C2 |
C1 |
O5 |
112.084 |
|
C2 |
N6 |
H7 |
118.876 |
| C2 |
N6 |
H8 |
120.229 |
|
O3 |
C2 |
N6 |
126.345 |
| O4 |
C1 |
O5 |
124.745 |
|
H7 |
N6 |
H8 |
120.896 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -357.540013 |
| Energy at 298.15K | -357.545239 |
| Nuclear repulsion energy | 233.151646 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3832 |
3608 |
89.05 |
|
|
|
| 2 |
A' |
3733 |
3514 |
129.58 |
|
|
|
| 3 |
A' |
3688 |
3472 |
82.90 |
|
|
|
| 4 |
A' |
1900 |
1789 |
154.66 |
|
|
|
| 5 |
A' |
1831 |
1724 |
351.14 |
|
|
|
| 6 |
A' |
1660 |
1563 |
69.64 |
|
|
|
| 7 |
A' |
1496 |
1408 |
156.78 |
|
|
|
| 8 |
A' |
1380 |
1299 |
421.36 |
|
|
|
| 9 |
A' |
1245 |
1172 |
10.77 |
|
|
|
| 10 |
A' |
1128 |
1062 |
2.42 |
|
|
|
| 11 |
A' |
822 |
774 |
10.63 |
|
|
|
| 12 |
A' |
632 |
595 |
15.84 |
|
|
|
| 13 |
A' |
555 |
522 |
2.40 |
|
|
|
| 14 |
A' |
416 |
391 |
6.33 |
|
|
|
| 15 |
A' |
282 |
266 |
39.57 |
|
|
|
| 16 |
A" |
788 |
742 |
3.69 |
|
|
|
| 17 |
A" |
729 |
686 |
121.93 |
|
|
|
| 18 |
A" |
662 |
623 |
6.61 |
|
|
|
| 19 |
A" |
453 |
427 |
92.43 |
|
|
|
| 20 |
A" |
349 |
329 |
207.02 |
|
|
|
| 21 |
A" |
107 |
101 |
2.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13843.2 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 13032.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.009 |
-0.793 |
0.000 |
| C2 |
0.000 |
0.748 |
0.000 |
| O3 |
-1.075 |
1.348 |
0.000 |
| O4 |
1.027 |
-1.448 |
0.000 |
| O5 |
-1.227 |
-1.298 |
0.000 |
| N6 |
1.229 |
1.288 |
0.000 |
| H7 |
1.337 |
2.287 |
0.000 |
| H8 |
2.031 |
0.681 |
0.000 |
| H9 |
-1.829 |
-0.531 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5408 | 2.3986 | 1.2115 | 1.3349 | 2.4123 | 3.3534 | 2.5022 | 1.8558 |
C2 | 1.5408 | | 1.2306 | 2.4249 | 2.3857 | 1.3427 | 2.0381 | 2.0321 | 2.2314 | O3 | 2.3986 | 1.2306 | | 3.4979 | 2.6496 | 2.3050 | 2.5882 | 3.1765 | 2.0237 | O4 | 1.2115 | 2.4249 | 3.4979 | | 2.2594 | 2.7437 | 3.7477 | 2.3539 | 2.9997 | O5 | 1.3349 | 2.3857 | 2.6496 | 2.2594 | | 3.5665 | 4.4070 | 3.8118 | 0.9748 | N6 | 2.4123 | 1.3427 | 2.3050 | 2.7437 | 3.5665 | | 1.0044 | 1.0054 | 3.5579 | H7 | 3.3534 | 2.0381 | 2.5882 | 3.7477 | 4.4070 | 1.0044 | | 1.7492 | 4.2376 | H8 | 2.5022 | 2.0321 | 3.1765 | 2.3539 | 3.8118 | 1.0054 | 1.7492 | | 4.0452 | H9 | 1.8558 | 2.2314 | 2.0237 | 2.9997 | 0.9748 | 3.5579 | 4.2376 | 4.0452 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.461 |
|
C1 |
C2 |
N6 |
113.382 |
| C1 |
O5 |
H9 |
105.867 |
|
C2 |
C1 |
O4 |
123.091 |
| C2 |
C1 |
O5 |
111.916 |
|
C2 |
N6 |
H7 |
119.849 |
| C2 |
N6 |
H8 |
119.159 |
|
O3 |
C2 |
N6 |
127.157 |
| O4 |
C1 |
O5 |
124.993 |
|
H7 |
N6 |
H8 |
120.992 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
