Jump to
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -357.519504 |
| Energy at 298.15K | -357.524452 |
| HF Energy | -356.555829 |
| Nuclear repulsion energy | 232.344317 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3874 |
3632 |
79.07 |
|
|
|
| 2 |
A' |
3861 |
3620 |
81.42 |
|
|
|
| 3 |
A' |
3716 |
3484 |
79.71 |
|
|
|
| 4 |
A' |
1909 |
1790 |
7.82 |
|
|
|
| 5 |
A' |
1887 |
1770 |
566.45 |
|
|
|
| 6 |
A' |
1669 |
1565 |
119.96 |
|
|
|
| 7 |
A' |
1495 |
1402 |
17.46 |
|
|
|
| 8 |
A' |
1368 |
1283 |
73.24 |
|
|
|
| 9 |
A' |
1237 |
1160 |
315.60 |
|
|
|
| 10 |
A' |
1134 |
1064 |
3.73 |
|
|
|
| 11 |
A' |
809 |
759 |
7.57 |
|
|
|
| 12 |
A' |
622 |
583 |
80.27 |
|
|
|
| 13 |
A' |
545 |
511 |
0.16 |
|
|
|
| 14 |
A' |
428 |
401 |
3.87 |
|
|
|
| 15 |
A' |
280 |
262 |
13.87 |
|
|
|
| 16 |
A" |
817 |
766 |
6.47 |
|
|
|
| 17 |
A" |
678 |
636 |
158.55 |
|
|
|
| 18 |
A" |
628 |
589 |
12.58 |
|
|
|
| 19 |
A" |
434 |
407 |
2.67 |
|
|
|
| 20 |
A" |
259 |
242 |
277.37 |
|
|
|
| 21 |
A" |
45 |
42 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13848.7 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 12984.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.752 |
0.000 |
| C2 |
-0.052 |
-0.785 |
0.000 |
| O3 |
-1.098 |
-1.403 |
0.000 |
| O4 |
1.038 |
1.378 |
0.000 |
| O5 |
-1.218 |
1.298 |
0.000 |
| N6 |
1.192 |
-1.316 |
0.000 |
| H7 |
1.293 |
-2.314 |
0.000 |
| H8 |
1.998 |
-0.718 |
0.000 |
| H9 |
-1.092 |
2.257 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5376 | 2.4183 | 1.2117 | 1.3350 | 2.3869 | 3.3274 | 2.4802 | 1.8588 |
C2 | 1.5376 | | 1.2150 | 2.4216 | 2.3867 | 1.3529 | 2.0367 | 2.0508 | 3.2141 | O3 | 2.4183 | 1.2150 | | 3.5060 | 2.7028 | 2.2922 | 2.5595 | 3.1708 | 3.6591 | O4 | 1.2117 | 2.4216 | 3.5060 | | 2.2574 | 2.6981 | 3.7004 | 2.3052 | 2.3033 | O5 | 1.3350 | 2.3867 | 2.7028 | 2.2574 | | 3.5554 | 4.3990 | 3.7954 | 0.9673 | N6 | 2.3869 | 1.3529 | 2.2922 | 2.6981 | 3.5554 | | 1.0030 | 1.0031 | 4.2400 | H7 | 3.3274 | 2.0367 | 2.5595 | 3.7004 | 4.3990 | 1.0030 | | 1.7443 | 5.1552 | H8 | 2.4802 | 2.0508 | 3.1708 | 2.3052 | 3.7954 | 1.0031 | 1.7443 | | 4.2885 | H9 | 1.8588 | 3.2141 | 3.6591 | 2.3033 | 0.9673 | 4.2400 | 5.1552 | 4.2885 | |
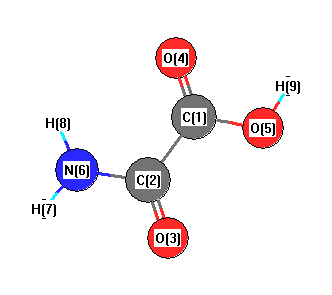 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.499 |
|
C1 |
C2 |
N6 |
111.172 |
| C1 |
O5 |
H9 |
106.589 |
|
C2 |
C1 |
O4 |
123.038 |
| C2 |
C1 |
O5 |
112.183 |
|
C2 |
N6 |
H7 |
118.902 |
| C2 |
N6 |
H8 |
120.300 |
|
O3 |
C2 |
N6 |
126.328 |
| O4 |
C1 |
O5 |
124.779 |
|
H7 |
N6 |
H8 |
120.798 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -357.524665 |
| Energy at 298.15K | -357.529923 |
| Nuclear repulsion energy | 233.673354 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3849 |
3609 |
92.13 |
|
|
|
| 2 |
A' |
3776 |
3541 |
129.78 |
|
|
|
| 3 |
A' |
3705 |
3474 |
88.92 |
|
|
|
| 4 |
A' |
1939 |
1818 |
179.49 |
|
|
|
| 5 |
A' |
1875 |
1758 |
366.15 |
|
|
|
| 6 |
A' |
1672 |
1568 |
78.41 |
|
|
|
| 7 |
A' |
1509 |
1415 |
161.20 |
|
|
|
| 8 |
A' |
1389 |
1302 |
445.11 |
|
|
|
| 9 |
A' |
1260 |
1182 |
7.44 |
|
|
|
| 10 |
A' |
1139 |
1068 |
3.00 |
|
|
|
| 11 |
A' |
833 |
781 |
11.06 |
|
|
|
| 12 |
A' |
640 |
600 |
16.88 |
|
|
|
| 13 |
A' |
561 |
526 |
2.49 |
|
|
|
| 14 |
A' |
420 |
394 |
6.39 |
|
|
|
| 15 |
A' |
286 |
268 |
39.46 |
|
|
|
| 16 |
A" |
799 |
749 |
1.67 |
|
|
|
| 17 |
A" |
722 |
677 |
127.16 |
|
|
|
| 18 |
A" |
661 |
619 |
7.80 |
|
|
|
| 19 |
A" |
459 |
430 |
89.00 |
|
|
|
| 20 |
A" |
351 |
329 |
215.39 |
|
|
|
| 21 |
A" |
107 |
101 |
2.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13975.8 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 13103.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.007 |
-0.791 |
0.000 |
| C2 |
0.000 |
0.748 |
0.000 |
| O3 |
-1.070 |
1.344 |
0.000 |
| O4 |
1.023 |
-1.443 |
0.000 |
| O5 |
-1.224 |
-1.299 |
0.000 |
| N6 |
1.228 |
1.288 |
0.000 |
| H7 |
1.334 |
2.285 |
0.000 |
| H8 |
2.030 |
0.683 |
0.000 |
| H9 |
-1.831 |
-0.539 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5387 | 2.3913 | 1.2069 | 1.3317 | 2.4104 | 3.3504 | 2.5031 | 1.8552 |
C2 | 1.5387 | | 1.2251 | 2.4174 | 2.3845 | 1.3411 | 2.0358 | 2.0309 | 2.2376 | O3 | 2.3913 | 1.2251 | | 3.4851 | 2.6471 | 2.2985 | 2.5820 | 3.1697 | 2.0305 | O4 | 1.2069 | 2.4174 | 3.4851 | | 2.2515 | 2.7379 | 3.7411 | 2.3524 | 2.9936 | O5 | 1.3317 | 2.3845 | 2.6471 | 2.2515 | | 3.5636 | 4.4034 | 3.8100 | 0.9726 | N6 | 2.4104 | 1.3411 | 2.2985 | 2.7379 | 3.5636 | | 1.0036 | 1.0044 | 3.5623 | H7 | 3.3504 | 2.0358 | 2.5820 | 3.7411 | 4.4034 | 1.0036 | | 1.7468 | 4.2418 | H8 | 2.5031 | 2.0309 | 3.1697 | 2.3524 | 3.8100 | 1.0044 | 1.7468 | | 4.0496 | H9 | 1.8552 | 2.2376 | 2.0305 | 2.9936 | 0.9726 | 3.5623 | 4.2418 | 4.0496 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.387 |
|
C1 |
C2 |
N6 |
113.474 |
| C1 |
O5 |
H9 |
106.199 |
|
C2 |
C1 |
O4 |
122.944 |
| C2 |
C1 |
O5 |
112.148 |
|
C2 |
N6 |
H7 |
119.822 |
| C2 |
N6 |
H8 |
119.275 |
|
O3 |
C2 |
N6 |
127.139 |
| O4 |
C1 |
O5 |
124.908 |
|
H7 |
N6 |
H8 |
120.903 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
