return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H10 (Norbornene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-271.886355
Energy at 298.15K-271.898632
HF Energy-270.876643
Nuclear repulsion energy309.203043
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3303 3086 15.99      
2 A' 3217 3006 41.82      
3 A' 3213 3003 25.16      
4 A' 3198 2988 6.96      
5 A' 3150 2943 40.89      
6 A' 3135 2929 30.92      
7 A' 1645 1537 0.64      
8 A' 1567 1464 0.34      
9 A' 1541 1440 2.01      
10 A' 1372 1282 0.37      
11 A' 1352 1264 0.94      
12 A' 1226 1146 0.23      
13 A' 1179 1102 4.70      
14 A' 1140 1065 0.69      
15 A' 1069 999 1.18      
16 A' 1012 945 0.19      
17 A' 990 925 0.30      
18 A' 951 889 0.63      
19 A' 919 859 4.25      
20 A' 844 789 0.38      
21 A' 799 747 3.50      
22 A' 737 689 36.84      
23 A' 483 451 0.78      
24 A' 386 361 1.95      
25 A" 3278 3063 5.97      
26 A" 3202 2992 0.04      
27 A" 3195 2985 59.29      
28 A" 3139 2933 22.60      
29 A" 1543 1442 0.36      
30 A" 1408 1316 6.53      
31 A" 1343 1255 1.90      
32 A" 1324 1237 0.10      
33 A" 1315 1228 0.25      
34 A" 1272 1188 0.49      
35 A" 1226 1146 0.09      
36 A" 1171 1095 0.18      
37 A" 1076 1005 0.54      
38 A" 994 929 0.00      
39 A" 979 915 0.10      
40 A" 928 867 4.47      
41 A" 870 813 2.46      
42 A" 822 768 1.08      
43 A" 683 638 0.46      
44 A" 486 454 0.15      
45 A" 250 234 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 34465.8 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 32204.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.13232 0.11572 0.10142

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.120 0.790 0.000
C2 0.222 0.249 1.122
C3 0.222 0.249 -1.122
C4 -1.119 0.780 0.673
C5 -1.119 0.780 -0.673
C6 0.222 -1.267 0.775
C7 0.222 -1.267 -0.775
H8 1.183 1.877 0.000
H9 2.122 0.356 0.000
H10 0.511 0.477 2.146
H11 0.511 0.477 -2.146
H12 -1.955 0.987 1.326
H13 -1.955 0.987 -1.326
H14 1.119 -1.747 1.167
H15 1.119 -1.747 -1.167
H16 -0.641 -1.776 1.199
H17 -0.641 -1.776 -1.199

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.53551.53552.33852.33852.37502.37501.08871.09162.25242.25243.35473.35472.79282.79283.33593.3359
C21.53552.24351.51022.30261.55572.42872.19802.20881.08783.28792.30763.35762.18913.16702.20343.1993
C31.53552.24352.30261.51022.42871.55572.19802.20883.28791.08783.35762.30763.16702.18913.19932.2034
C42.33851.51022.30261.34672.44922.84372.63783.33712.21693.27021.08042.17673.41133.84462.65283.2042
C52.33852.30261.51021.34672.84372.44922.63783.33713.27022.21692.17671.08043.84463.41133.20422.6528
C62.37501.55572.42872.44922.84371.55043.37812.61612.23683.41413.18163.77271.09022.19281.08832.2142
C72.37502.42871.55572.84372.44921.55043.37812.61613.41412.23683.77273.18162.19281.09022.21421.0883
H81.08872.19802.19802.63782.63783.37813.37811.78762.64882.64883.52123.52123.80803.80804.25614.2561
H91.09162.20882.20883.33713.33712.61612.61611.78762.68592.68594.33324.33322.60582.60583.69043.6904
H102.25241.08783.28792.21693.27022.23683.41412.64882.68594.29112.64794.28832.50454.03632.70184.1942
H112.25243.28791.08783.27022.21693.41412.23682.64882.68594.29114.28832.64794.03632.50454.19422.7018
H123.35472.30763.35761.08042.17673.18163.77273.52124.33322.64794.28832.65154.11674.81023.06193.9666
H133.35473.35762.30762.17671.08043.77273.18163.52124.33324.28832.64792.65154.81024.11673.96663.0619
H142.79282.18913.16703.41133.84461.09022.19283.80802.60582.50454.03634.11674.81022.33471.76102.9496
H152.79283.16702.18913.84463.41132.19281.09023.80802.60584.03632.50454.81024.11672.33472.94961.7610
H163.33592.20343.19932.65283.20421.08832.21424.25613.69042.70184.19423.06193.96661.76102.94962.3982
H173.33593.19932.20343.20422.65282.21421.08834.25613.69044.19422.70183.96663.06192.94961.76102.3982

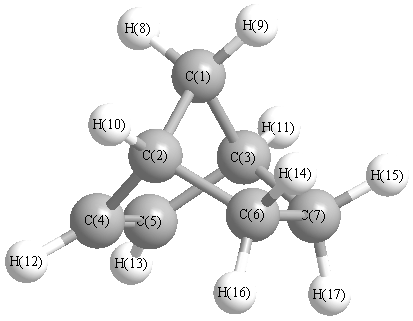
picture of Norbornene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 100.309 C1 C2 C6 100.404
C1 C2 H10 117.304 C1 C3 C5 100.309
C1 C3 C7 100.404 C1 C3 H11 117.304
C2 C1 C3 93.865 C2 C1 H8 112.664
C2 C1 H9 113.364 C2 C4 C5 107.272
C2 C4 H12 125.108 C2 C6 C7 102.871
C2 C6 H14 110.419 C2 C6 H16 111.667
C3 C1 H8 112.664 C3 C1 H9 113.364
C3 C5 C4 107.272 C3 C5 H13 125.108
C3 C7 C6 102.871 C3 C7 H15 110.419
C3 C7 H17 111.667 C4 C2 C6 106.031
C4 C2 H10 116.207 C4 C5 H13 127.142
C5 C3 C7 106.031 C5 C3 H11 116.207
C5 C4 H12 127.142 C6 C2 H10 114.428
C6 C7 H15 111.084 C6 C7 H17 112.925
C7 C3 H11 114.428 C7 C6 H14 111.084
C7 C6 H16 112.925 H8 C1 H9 110.145
H14 C6 H16 107.867 H15 C7 H17 107.867
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability