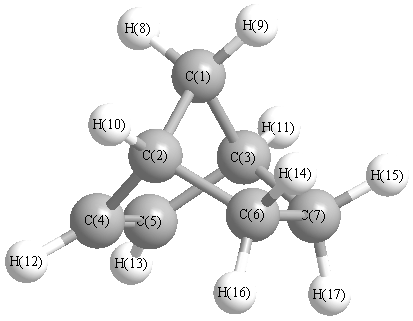
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3251 |
3102 |
18.09 |
|
|
|
| 2 |
A' |
3150 |
3006 |
36.31 |
|
|
|
| 3 |
A' |
3147 |
3003 |
38.82 |
|
|
|
| 4 |
A' |
3142 |
2998 |
7.85 |
|
|
|
| 5 |
A' |
3091 |
2950 |
48.31 |
|
|
|
| 6 |
A' |
3076 |
2936 |
36.23 |
|
|
|
| 7 |
A' |
1676 |
1599 |
1.17 |
|
|
|
| 8 |
A' |
1523 |
1454 |
0.77 |
|
|
|
| 9 |
A' |
1496 |
1428 |
3.15 |
|
|
|
| 10 |
A' |
1341 |
1279 |
0.45 |
|
|
|
| 11 |
A' |
1327 |
1267 |
0.88 |
|
|
|
| 12 |
A' |
1204 |
1149 |
0.29 |
|
|
|
| 13 |
A' |
1157 |
1104 |
5.46 |
|
|
|
| 14 |
A' |
1122 |
1071 |
1.07 |
|
|
|
| 15 |
A' |
1048 |
1000 |
1.44 |
|
|
|
| 16 |
A' |
992 |
947 |
0.32 |
|
|
|
| 17 |
A' |
971 |
927 |
0.35 |
|
|
|
| 18 |
A' |
933 |
890 |
0.78 |
|
|
|
| 19 |
A' |
903 |
861 |
5.59 |
|
|
|
| 20 |
A' |
827 |
789 |
0.71 |
|
|
|
| 21 |
A' |
787 |
751 |
5.17 |
|
|
|
| 22 |
A' |
734 |
701 |
36.40 |
|
|
|
| 23 |
A' |
479 |
457 |
0.90 |
|
|
|
| 24 |
A' |
380 |
363 |
2.01 |
|
|
|
| 25 |
A" |
3225 |
3077 |
7.80 |
|
|
|
| 26 |
A" |
3139 |
2996 |
66.50 |
|
|
|
| 27 |
A" |
3132 |
2989 |
3.83 |
|
|
|
| 28 |
A" |
3079 |
2938 |
26.09 |
|
|
|
| 29 |
A" |
1496 |
1428 |
0.71 |
|
|
|
| 30 |
A" |
1382 |
1319 |
8.11 |
|
|
|
| 31 |
A" |
1317 |
1257 |
2.56 |
|
|
|
| 32 |
A" |
1294 |
1235 |
0.24 |
|
|
|
| 33 |
A" |
1286 |
1228 |
0.44 |
|
|
|
| 34 |
A" |
1246 |
1189 |
0.49 |
|
|
|
| 35 |
A" |
1208 |
1153 |
0.18 |
|
|
|
| 36 |
A" |
1147 |
1095 |
0.34 |
|
|
|
| 37 |
A" |
1057 |
1009 |
0.88 |
|
|
|
| 38 |
A" |
976 |
931 |
0.01 |
|
|
|
| 39 |
A" |
963 |
919 |
0.38 |
|
|
|
| 40 |
A" |
939 |
896 |
4.75 |
|
|
|
| 41 |
A" |
857 |
818 |
3.47 |
|
|
|
| 42 |
A" |
812 |
775 |
1.62 |
|
|
|
| 43 |
A" |
679 |
648 |
0.56 |
|
|
|
| 44 |
A" |
488 |
466 |
0.20 |
|
|
|
| 45 |
A" |
254 |
242 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33866.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 32318.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.213 |
|
|
|
| 2 |
C |
-0.180 |
|
|
|
| 3 |
C |
-0.180 |
|
|
|
| 4 |
C |
-0.084 |
|
|
|
| 5 |
C |
-0.084 |
|
|
|
| 6 |
C |
-0.228 |
|
|
|
| 7 |
C |
-0.228 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
| 13 |
H |
0.114 |
|
|
|
| 14 |
H |
0.125 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.127 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.077 |
-0.262 |
0.000 |
0.273 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.776 |
-0.368 |
0.000 |
| y |
-0.368 |
-44.311 |
0.000 |
| z |
0.000 |
0.000 |
-41.568 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.164 |
-0.368 |
0.000 |
| y |
-0.368 |
-2.639 |
0.000 |
| z |
0.000 |
0.000 |
1.475 |
|
| Polar |
|---|
| 3z2-r2 | 2.951 |
|---|
| x2-y2 | 2.535 |
|---|
| xy | -0.368 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.639 |
-0.663 |
0.000 |
| y |
-0.663 |
8.987 |
0.000 |
| z |
0.000 |
0.000 |
10.456 |
<r2> (average value of r
2) Å
2
| <r2> |
163.833 |
| (<r2>)1/2 |
12.800 |