Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -192.903931 |
| Energy at 298.15K | -192.911122 |
| HF Energy | -192.903931 |
| Nuclear repulsion energy | 128.564274 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3115 |
2972 |
23.85 |
|
|
|
| 2 |
A1 |
3051 |
2911 |
1.87 |
|
|
|
| 3 |
A1 |
1561 |
1490 |
0.02 |
|
|
|
| 4 |
A1 |
1501 |
1432 |
2.56 |
|
|
|
| 5 |
A1 |
1399 |
1335 |
3.86 |
|
|
|
| 6 |
A1 |
1082 |
1032 |
8.40 |
|
|
|
| 7 |
A1 |
966 |
921 |
27.20 |
|
|
|
| 8 |
A1 |
818 |
780 |
3.39 |
|
|
|
| 9 |
A2 |
3090 |
2948 |
0.00 |
|
|
|
| 10 |
A2 |
1243 |
1186 |
0.00 |
|
|
|
| 11 |
A2 |
1176 |
1122 |
0.00 |
|
|
|
| 12 |
A2 |
845 |
806 |
0.00 |
|
|
|
| 13 |
B1 |
3175 |
3030 |
26.86 |
|
|
|
| 14 |
B1 |
3089 |
2948 |
90.32 |
|
|
|
| 15 |
B1 |
1204 |
1149 |
0.92 |
|
|
|
| 16 |
B1 |
1158 |
1105 |
1.82 |
|
|
|
| 17 |
B1 |
770 |
735 |
0.40 |
|
|
|
| 18 |
B1 |
35 |
33 |
2.61 |
|
|
|
| 19 |
B2 |
3039 |
2900 |
159.55 |
|
|
|
| 20 |
B2 |
1529 |
1459 |
3.96 |
|
|
|
| 21 |
B2 |
1322 |
1262 |
0.14 |
|
|
|
| 22 |
B2 |
1272 |
1214 |
10.24 |
|
|
|
| 23 |
B2 |
1090 |
1040 |
107.11 |
|
|
|
| 24 |
B2 |
972 |
928 |
4.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19248.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 18369.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.066 |
| C2 |
0.000 |
0.000 |
-1.073 |
| C3 |
0.000 |
1.029 |
0.066 |
| C4 |
0.000 |
-1.029 |
0.066 |
| H5 |
0.890 |
0.000 |
-1.705 |
| H6 |
-0.890 |
0.000 |
-1.705 |
| H7 |
0.891 |
1.666 |
0.131 |
| H8 |
-0.891 |
1.666 |
0.131 |
| H9 |
-0.891 |
-1.666 |
0.131 |
| H10 |
0.891 |
-1.666 |
0.131 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1391 | 1.4350 | 1.4350 | 2.9107 | 2.9107 | 2.1080 | 2.1080 | 2.1080 | 2.1080 |
C2 | 2.1391 | | 1.5345 | 1.5345 | 1.0916 | 1.0916 | 2.2401 | 2.2401 | 2.2401 | 2.2401 | C3 | 1.4350 | 1.5345 | | 2.0573 | 2.2329 | 2.2329 | 1.0976 | 1.0976 | 2.8386 | 2.8386 | C4 | 1.4350 | 1.5345 | 2.0573 | | 2.2329 | 2.2329 | 2.8386 | 2.8386 | 1.0976 | 1.0976 | H5 | 2.9107 | 1.0916 | 2.2329 | 2.2329 | | 1.7798 | 2.4789 | 3.0526 | 3.0526 | 2.4789 | H6 | 2.9107 | 1.0916 | 2.2329 | 2.2329 | 1.7798 | | 3.0526 | 2.4789 | 2.4789 | 3.0526 | H7 | 2.1080 | 2.2401 | 1.0976 | 2.8386 | 2.4789 | 3.0526 | | 1.7829 | 3.7783 | 3.3311 | H8 | 2.1080 | 2.2401 | 1.0976 | 2.8386 | 3.0526 | 2.4789 | 1.7829 | | 3.3311 | 3.7783 | H9 | 2.1080 | 2.2401 | 2.8386 | 1.0976 | 3.0526 | 2.4789 | 3.7783 | 3.3311 | | 1.7829 | H10 | 2.1080 | 2.2401 | 2.8386 | 1.0976 | 2.4789 | 3.0526 | 3.3311 | 3.7783 | 1.7829 | |
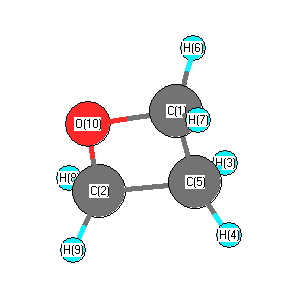 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.109 |
|
O1 |
C3 |
H7 |
112.002 |
| O1 |
C3 |
H8 |
112.002 |
|
O1 |
C4 |
C2 |
92.109 |
| O1 |
C4 |
H9 |
112.002 |
|
O1 |
C4 |
H10 |
112.002 |
| C2 |
C3 |
H7 |
115.660 |
|
C2 |
C3 |
H8 |
115.660 |
| C2 |
C4 |
H9 |
115.660 |
|
C2 |
C4 |
H10 |
115.660 |
| C3 |
O1 |
C4 |
91.591 |
|
C3 |
C2 |
C4 |
84.190 |
| C3 |
C2 |
H5 |
115.453 |
|
C3 |
C2 |
H6 |
115.453 |
| C4 |
C2 |
H5 |
115.453 |
|
C4 |
C2 |
H6 |
115.453 |
| H5 |
C2 |
H6 |
109.215 |
|
H7 |
C3 |
H8 |
108.630 |
| H9 |
C4 |
H10 |
108.630 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.518 |
|
|
|
| 2 |
C |
-0.353 |
|
|
|
| 3 |
C |
0.051 |
|
|
|
| 4 |
C |
0.051 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.142 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.121 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.972 |
1.972 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.952 |
0.000 |
0.000 |
| y |
0.000 |
-22.656 |
0.000 |
| z |
0.000 |
0.000 |
-27.343 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.048 |
0.000 |
0.000 |
| y |
0.000 |
2.991 |
0.000 |
| z |
0.000 |
0.000 |
-4.039 |
|
| Polar |
|---|
| 3z2-r2 | -8.079 |
|---|
| x2-y2 | -1.295 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.693 |
0.000 |
0.000 |
| y |
0.000 |
5.611 |
0.000 |
| z |
0.000 |
0.000 |
4.500 |
<r2> (average value of r
2) Å
2
| <r2> |
65.890 |
| (<r2>)1/2 |
8.117 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -192.903931 |
| Energy at 298.15K | -192.911123 |
| HF Energy | -192.903931 |
| Nuclear repulsion energy | 128.562600 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3175 |
3030 |
26.89 |
|
|
|
| 2 |
A' |
3115 |
2972 |
23.86 |
|
|
|
| 3 |
A' |
3089 |
2948 |
90.28 |
|
|
|
| 4 |
A' |
3051 |
2911 |
1.87 |
|
|
|
| 5 |
A' |
1561 |
1490 |
0.02 |
|
|
|
| 6 |
A' |
1501 |
1432 |
2.56 |
|
|
|
| 7 |
A' |
1399 |
1335 |
3.86 |
|
|
|
| 8 |
A' |
1204 |
1149 |
0.93 |
|
|
|
| 9 |
A' |
1158 |
1105 |
1.82 |
|
|
|
| 10 |
A' |
1082 |
1032 |
8.40 |
|
|
|
| 11 |
A' |
966 |
921 |
27.20 |
|
|
|
| 12 |
A' |
818 |
780 |
3.39 |
|
|
|
| 13 |
A' |
770 |
735 |
0.40 |
|
|
|
| 14 |
A' |
35 |
34 |
2.60 |
|
|
|
| 15 |
A" |
3090 |
2948 |
0.00 |
|
|
|
| 16 |
A" |
3039 |
2900 |
159.50 |
|
|
|
| 17 |
A" |
1529 |
1459 |
3.96 |
|
|
|
| 18 |
A" |
1322 |
1261 |
0.14 |
|
|
|
| 19 |
A" |
1272 |
1214 |
10.22 |
|
|
|
| 20 |
A" |
1243 |
1186 |
0.00 |
|
|
|
| 21 |
A" |
1176 |
1122 |
0.00 |
|
|
|
| 22 |
A" |
1090 |
1040 |
107.11 |
|
|
|
| 23 |
A" |
972 |
928 |
4.12 |
|
|
|
| 24 |
A" |
845 |
806 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19248.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 18369.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.001 |
-1.066 |
0.000 |
| C2 |
0.000 |
1.073 |
0.000 |
| C3 |
0.000 |
-0.066 |
1.029 |
| C4 |
0.000 |
-0.066 |
-1.029 |
| H5 |
0.890 |
1.705 |
0.000 |
| H6 |
-0.890 |
1.705 |
0.000 |
| H7 |
0.892 |
-0.131 |
1.665 |
| H8 |
-0.891 |
-0.131 |
1.666 |
| H9 |
0.892 |
-0.131 |
-1.665 |
| H10 |
-0.891 |
-0.131 |
-1.666 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1390 | 1.4350 | 1.4350 | 2.9111 | 2.9102 | 2.1081 | 2.1081 | 2.1081 | 2.1081 |
C2 | 2.1390 | | 1.5345 | 1.5345 | 1.0916 | 1.0916 | 2.2399 | 2.2402 | 2.2399 | 2.2402 | C3 | 1.4350 | 1.5345 | | 2.0575 | 2.2331 | 2.2329 | 1.0976 | 1.0975 | 2.8384 | 2.8392 | C4 | 1.4350 | 1.5345 | 2.0575 | | 2.2331 | 2.2329 | 2.8384 | 2.8392 | 1.0976 | 1.0975 | H5 | 2.9111 | 1.0916 | 2.2331 | 2.2331 | | 1.7797 | 2.4788 | 3.0525 | 2.4788 | 3.0525 | H6 | 2.9102 | 1.0916 | 2.2329 | 2.2329 | 1.7797 | | 3.0526 | 2.4790 | 3.0526 | 2.4790 | H7 | 2.1081 | 2.2399 | 1.0976 | 2.8384 | 2.4788 | 3.0526 | | 1.7830 | 3.3301 | 3.7784 | H8 | 2.1081 | 2.2402 | 1.0975 | 2.8392 | 3.0525 | 2.4790 | 1.7830 | | 3.7784 | 3.3324 | H9 | 2.1081 | 2.2399 | 2.8384 | 1.0976 | 2.4788 | 3.0526 | 3.3301 | 3.7784 | | 1.7830 | H10 | 2.1081 | 2.2402 | 2.8392 | 1.0975 | 3.0525 | 2.4790 | 3.7784 | 3.3324 | 1.7830 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.101 |
|
O1 |
C3 |
H7 |
112.006 |
| O1 |
C3 |
H8 |
112.004 |
|
O1 |
C4 |
C2 |
92.101 |
| O1 |
C4 |
H9 |
112.006 |
|
O1 |
C4 |
H10 |
112.004 |
| C2 |
C3 |
H7 |
115.648 |
|
C2 |
C3 |
H8 |
115.669 |
| C2 |
C4 |
H9 |
115.648 |
|
C2 |
C4 |
H10 |
115.669 |
| C3 |
O1 |
C4 |
91.601 |
|
C3 |
C2 |
C4 |
84.197 |
| C3 |
C2 |
H5 |
115.459 |
|
C3 |
C2 |
H6 |
115.448 |
| C4 |
C2 |
H5 |
115.459 |
|
C4 |
C2 |
H6 |
115.448 |
| H5 |
C2 |
H6 |
109.209 |
|
H7 |
C3 |
H8 |
108.634 |
| H9 |
C4 |
H10 |
108.634 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.518 |
|
|
|
| 2 |
C |
-0.353 |
|
|
|
| 3 |
C |
0.051 |
|
|
|
| 4 |
C |
0.051 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.142 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.121 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.002 |
1.972 |
0.000 |
1.972 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.952 |
-0.002 |
0.000 |
| y |
-0.002 |
-27.343 |
0.000 |
| z |
0.000 |
0.000 |
-22.656 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.048 |
-0.002 |
0.000 |
| y |
-0.002 |
-4.040 |
0.000 |
| z |
0.000 |
0.000 |
2.992 |
|
| Polar |
|---|
| 3z2-r2 | 5.984 |
|---|
| x2-y2 | 3.392 |
|---|
| xy | -0.002 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.693 |
0.000 |
0.000 |
| y |
0.000 |
4.499 |
0.000 |
| z |
0.000 |
0.000 |
5.612 |
<r2> (average value of r
2) Å
2
| <r2> |
65.891 |
| (<r2>)1/2 |
8.117 |
