return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O2 (1,3-Propanediol)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-268.803316
Energy at 298.15K-268.813437
Nuclear repulsion energy195.484637
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3900 3644 28.83      
2 A 3842 3590 126.72      
3 A 3203 2993 24.27      
4 A 3183 2975 45.88      
5 A 3145 2939 60.39      
6 A 3131 2926 9.23      
7 A 3088 2885 53.92      
8 A 3060 2860 65.74      
9 A 1587 1483 2.18      
10 A 1572 1469 0.07      
11 A 1523 1423 12.25      
12 A 1498 1400 49.47      
13 A 1490 1392 45.37      
14 A 1438 1344 2.34      
15 A 1419 1326 0.20      
16 A 1342 1254 23.34      
17 A 1313 1226 3.91      
18 A 1255 1173 12.89      
19 A 1226 1145 56.87      
20 A 1168 1092 30.40      
21 A 1135 1060 8.68      
22 A 1125 1052 122.05      
23 A 995 929 1.97      
24 A 944 882 2.32      
25 A 937 875 11.25      
26 A 850 794 9.68      
27 A 564 527 164.90      
28 A 518 484 3.05      
29 A 397 371 4.16      
30 A 340 318 21.94      
31 A 289 270 76.68      
32 A 226 211 2.86      
33 A 109 102 1.09      

Unscaled Zero Point Vibrational Energy (zpe) 25905.9 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 24206.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.25711 0.13336 0.09710

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.020 1.038 -0.347
C2 1.285 0.468 0.167
C3 -1.243 0.409 0.310
H4 -0.086 0.886 -1.424
H5 -0.014 2.115 -0.163
O6 1.280 -0.943 -0.084
O7 -1.492 -0.911 -0.143
H8 2.059 -1.327 0.332
H9 -0.642 -1.367 -0.081
H10 2.135 0.941 -0.332
H11 1.372 0.663 1.241
H12 -1.122 0.444 1.401
H13 -2.133 0.987 0.066

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 O6 O7 H8 H9 H10 H11 H12 H13
C11.51451.52391.08981.09202.38492.45123.22242.49912.15732.14532.14992.1536
C21.51452.53242.14162.12331.43363.11621.96232.67271.09331.09472.70473.4588
C31.52392.53242.13842.15472.88941.41753.73071.91563.47892.78731.09821.0890
H41.08982.14162.13841.76262.64712.61683.54772.68152.47553.04613.04132.5345
H51.09202.12332.15471.76263.32163.36754.04883.53932.45422.44982.54242.4116
O62.38491.43362.88942.64713.32162.77300.96351.96812.08442.08403.14623.9241
O72.45123.11621.41752.61683.36752.77303.60720.96714.07703.54892.08712.0139
H83.22241.96233.73073.54774.04880.96353.60722.73282.36542.29283.79454.7961
H92.49912.67271.91562.68153.53931.96810.96712.73283.61993.15032.38902.7906
H102.15731.09333.47892.47552.45422.08444.07702.36543.61991.77003.72224.2868
H112.14531.09472.78733.04612.44982.08403.54892.29283.15031.77002.50863.7111
H122.14992.70471.09823.04132.54243.14622.08713.79452.38903.72222.50861.7602
H132.15363.45881.08902.53452.41163.92412.01394.79612.79064.28683.71111.7602

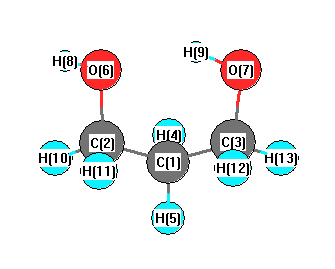
picture of 1,3-Propanediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O6 107.954 C1 C2 H10 110.599
C1 C2 H11 109.564 C1 C3 O7 112.836
C1 C3 H12 109.076 C1 C3 H13 109.901
C2 C1 C3 112.917 C2 C1 H4 109.559
C2 C1 H5 107.999 C2 O6 H8 108.294
C3 C1 H4 108.661 C3 C1 H5 109.809
C3 O7 H9 105.329 H4 C1 H5 107.770
O6 C2 H10 110.426 O6 C2 H11 110.311
O7 C3 H12 111.489 O7 C3 H13 106.179
H10 C2 H11 107.992 H12 C3 H13 107.173
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability