return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-306.793223
Energy at 298.15K-306.804625
Nuclear repulsion energy265.471367
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3239 3026 32.03      
2 A' 3217 3006 26.75      
3 A' 3206 2996 15.91      
4 A' 3140 2934 16.68      
5 A' 3073 2872 90.49      
6 A' 3025 2827 71.85      
7 A' 1574 1471 5.19      
8 A' 1558 1455 0.27      
9 A' 1524 1424 4.46      
10 A' 1452 1357 25.66      
11 A' 1352 1263 4.19      
12 A' 1240 1159 51.73      
13 A' 1215 1135 122.65      
14 A' 1142 1067 16.02      
15 A' 1030 962 37.70      
16 A' 949 887 7.09      
17 A' 871 814 12.40      
18 A' 658 615 5.81      
19 A' 493 460 0.85      
20 A' 454 424 10.52      
21 A' 278 259 2.22      
22 A" 3209 2998 36.60      
23 A" 3067 2865 18.80      
24 A" 1555 1453 1.66      
25 A" 1481 1384 19.03      
26 A" 1430 1336 0.22      
27 A" 1405 1312 0.01      
28 A" 1364 1274 1.68      
29 A" 1292 1207 28.49      
30 A" 1258 1175 1.07      
31 A" 1123 1050 58.12      
32 A" 1063 994 70.60      
33 A" 942 880 18.16      
34 A" 926 865 2.33      
35 A" 464 434 7.67      
36 A" 276 258 2.12      

Unscaled Zero Point Vibrational Energy (zpe) 27771.2 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 25949.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.16787 0.16081 0.09253

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.631 -1.193 0.000
O2 0.016 -0.765 1.174
O3 0.016 -0.765 -1.174
C4 0.016 0.661 1.233
C5 0.016 0.661 -1.233
C6 0.697 1.233 0.000
H7 -0.602 -2.278 0.000
H8 -1.671 -0.829 0.000
H9 0.532 0.929 2.152
H10 -1.017 1.028 1.294
H11 0.532 0.929 -2.152
H12 -1.017 1.028 -1.294
H13 1.744 0.934 0.000
H14 0.645 2.323 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.40741.40742.31932.31932.76631.08491.10153.23872.59963.23872.59963.18883.7409
O21.40742.34801.42762.79802.41562.01212.05612.02322.07283.76813.22092.69293.3631
O31.40742.34802.79801.42762.41562.01212.05613.76813.22092.02322.07282.69293.3631
C42.31931.42762.79802.46621.52043.24672.56661.08721.09793.43452.75492.14032.1628
C52.31932.79801.42762.46621.52043.24672.56663.43452.75491.08721.09792.14032.1628
C62.76632.41562.41561.52041.52043.74363.14012.17952.15782.17952.15781.08871.0912
H71.08492.01212.01213.24673.24673.74361.80034.02523.57434.02523.57433.97754.7669
H81.10152.05612.05612.56662.56663.14011.80033.54602.35613.54602.35613.84323.9114
H93.23872.02323.76811.08723.43452.17954.02523.54601.77314.30373.77952.46992.5662
H102.59962.07283.22091.09792.75492.15783.57432.35611.77313.77952.58863.05082.4729
H113.23873.76812.02323.43451.08722.17954.02523.54604.30373.77951.77312.46992.5662
H122.59963.22092.07282.75491.09792.15783.57432.35613.77952.58861.77313.05082.4729
H133.18882.69292.69292.14032.14031.08873.97753.84322.46993.05082.46993.05081.7712
H143.74093.36313.36312.16282.16281.09124.76693.91142.56622.47292.56622.47291.7712

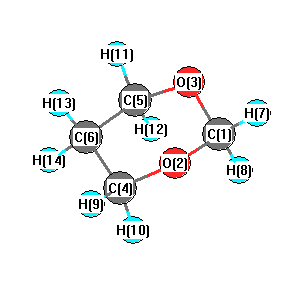
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 109.792 C1 O3 C5 109.792
O2 C1 O3 113.059 O2 C1 H7 106.965
O2 C1 H8 109.476 O2 C4 C6 110.012
O2 C4 H9 106.345 O2 C4 H10 109.628
O3 C1 H7 106.965 O3 C1 H8 109.476
O3 C5 C6 110.012 O3 C5 H11 106.345
O3 C5 H12 109.628 C4 C6 C5 108.400
C4 C6 H13 109.116 C4 C6 H14 110.748
C5 C6 H13 109.116 C5 C6 H14 110.748
C6 C4 H9 112.343 C6 C4 H10 109.949
C6 C5 H11 112.343 C6 C5 H12 109.949
H7 C1 H8 110.860 H9 C4 H10 108.474
H11 C5 H12 108.474 H13 C6 H14 108.683
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability