return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-306.819133
Energy at 298.15K-306.830523
Nuclear repulsion energy264.919247
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3214 3026 36.10      
2 A' 3186 2999 33.97      
3 A' 3174 2988 21.32      
4 A' 3116 2933 20.65      
5 A' 3054 2875 103.81      
6 A' 3014 2838 64.69      
7 A' 1576 1484 4.38      
8 A' 1559 1468 0.46      
9 A' 1528 1439 3.68      
10 A' 1456 1370 30.27      
11 A' 1351 1272 4.78      
12 A' 1247 1174 61.53      
13 A' 1220 1148 119.35      
14 A' 1142 1075 13.83      
15 A' 1031 971 37.70      
16 A' 946 890 7.79      
17 A' 870 819 12.36      
18 A' 662 623 6.06      
19 A' 494 465 0.71      
20 A' 448 421 11.08      
21 A' 272 256 2.10      
22 A" 3181 2995 44.22      
23 A" 3046 2868 19.56      
24 A" 1557 1466 1.20      
25 A" 1485 1398 22.91      
26 A" 1433 1349 0.04      
27 A" 1406 1324 0.00      
28 A" 1365 1285 1.69      
29 A" 1292 1216 30.01      
30 A" 1258 1184 1.25      
31 A" 1134 1067 72.05      
32 A" 1066 1004 57.99      
33 A" 943 888 14.93      
34 A" 925 871 0.71      
35 A" 466 439 7.91      
36 A" 272 256 2.32      

Unscaled Zero Point Vibrational Energy (zpe) 27693.2 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 26070.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.16699 0.16026 0.09202

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.629 -1.201 0.000
O2 0.016 -0.765 1.173
O3 0.016 -0.765 -1.173
C4 0.016 0.662 1.239
C5 0.016 0.662 -1.239
C6 0.693 1.241 0.000
H7 -0.593 -2.287 0.000
H8 -1.673 -0.847 0.000
H9 0.539 0.928 2.156
H10 -1.018 1.029 1.310
H11 0.539 0.928 -2.156
H12 -1.018 1.029 -1.310
H13 1.746 0.954 0.000
H14 0.630 2.333 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.40821.40822.32812.32812.77701.08641.10243.24752.61483.24752.61483.20663.7511
O21.40822.34651.42842.80272.42092.01542.05862.02652.07523.77183.23322.70603.3691
O31.40822.34652.80271.42842.42092.01542.05863.77183.23322.02652.07522.70603.3691
C42.32811.42842.80272.47811.52623.25542.58151.08901.09953.44582.77502.14732.1690
C52.32812.80271.42842.47811.52623.25542.58153.44582.77501.08901.09952.14732.1690
C62.77702.42092.42091.52621.52623.75493.15562.18452.16502.18452.16501.09111.0933
H71.08642.01542.01543.25543.25543.75491.80024.03283.58994.03283.58993.99614.7784
H81.10242.05862.05862.58152.58153.15561.80023.56272.37943.56272.37943.86383.9257
H93.24752.02653.77181.08903.44582.18454.03283.56271.77514.31293.80122.47112.5753
H102.61482.07523.23321.09952.77502.16503.58992.37941.77513.80122.61973.05892.4760
H113.24753.77182.02653.44581.08902.18454.03283.56274.31293.80121.77512.47112.5753
H122.61483.23322.07522.77501.09952.16503.58992.37943.80122.61971.77513.05892.4760
H133.20662.70602.70602.14732.14731.09113.99613.86382.47113.05892.47113.05891.7737
H143.75113.36913.36912.16902.16901.09334.77843.92572.57532.47602.57532.47601.7737

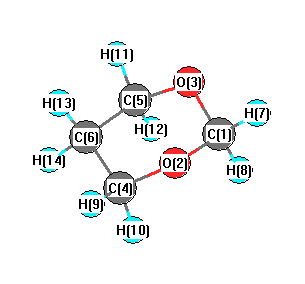
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.319 C1 O3 C5 110.319
O2 C1 O3 112.854 O2 C1 H7 107.079
O2 C1 H8 109.560 O2 C4 C6 110.001
O2 C4 H9 106.439 O2 C4 H10 109.666
O3 C1 H7 107.079 O3 C1 H8 109.560
O3 C5 C6 110.001 O3 C5 H11 106.439
O3 C5 H12 109.666 C4 C6 C5 108.559
C4 C6 H13 109.128 C4 C6 H14 110.708
C5 C6 H13 109.128 C5 C6 H14 110.708
C6 C4 H9 112.222 C6 C4 H10 110.023
C6 C5 H11 112.222 C6 C5 H12 110.023
H7 C1 H8 110.662 H9 C4 H10 108.402
H11 C5 H12 108.402 H13 C6 H14 108.581
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability